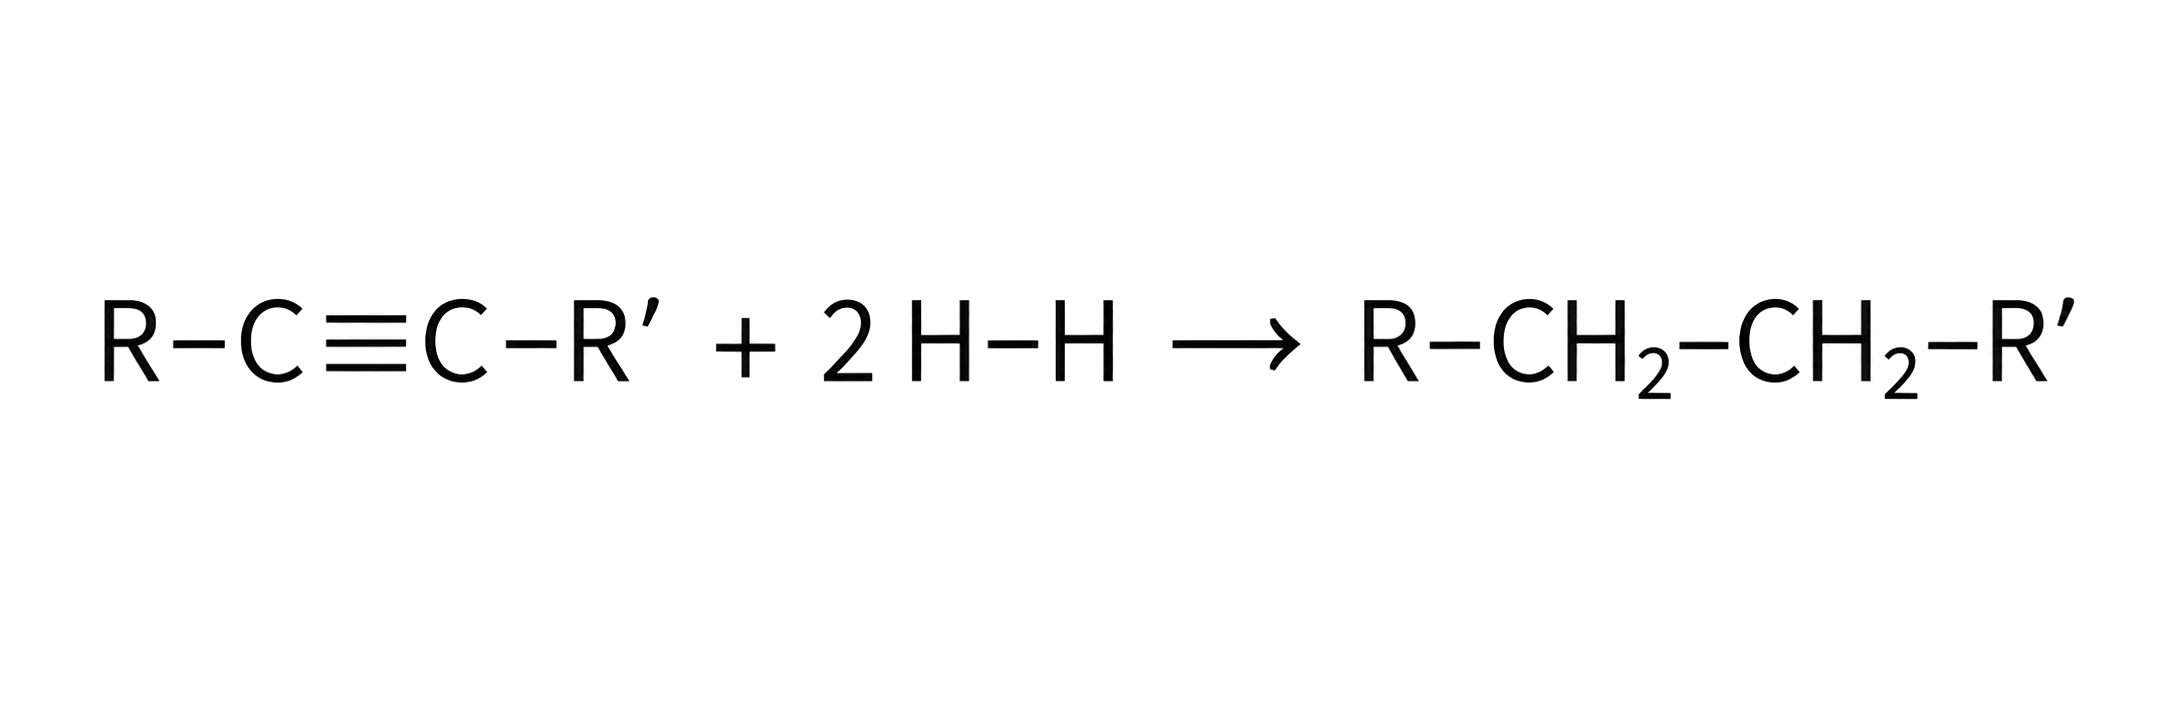Reactivity 3. What are the mechanisms of chemical change?

Syllabus
First assessment 2025
Section
—
Level
HL

3.1 Proton transfer reactions

Syllabus
First assessment 2025
Topic
3.1
Level
HL

Brønsted–Lowry Acids and Bases

A Brønsted–Lowry acid donates H+ and a Brønsted–Lowry base accepts H+. An alkali is a base that is soluble in water.
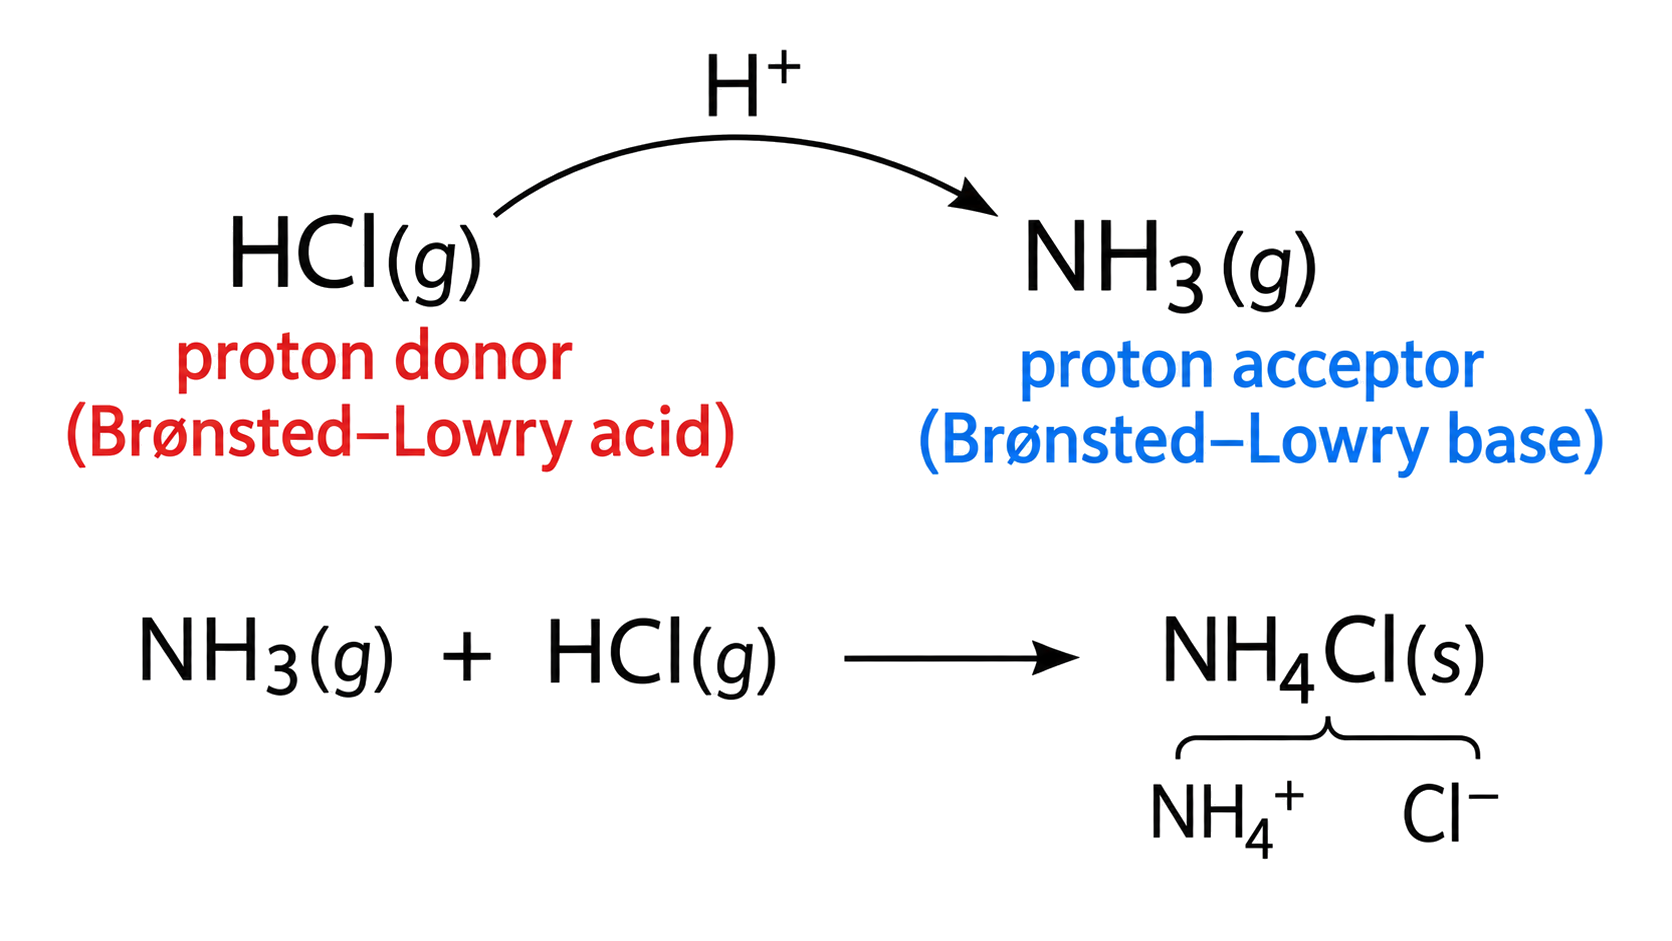
HCl(g) is the proton donor and Brønsted–Lowry acid; NH3(g) is the proton acceptor and Brønsted–Lowry base; the proton-transfer cue runs from HCl to NH3; the overall product is unambiguously NH4Cl(s), composed of NH4+ and Cl−.

Follow the proton: the species losing it is the acid and the species gaining it is the base.

Pair species that differ by exactly one H⁺ to identify conjugate acid–base pairs. Charge alone does not decide the role: in NH₄⁺ + H₂O ⇌ NH₃ + H₃O⁺, NH₄⁺ is the proton donor and water is the acceptor.

Assigning Acid and Base Roles

[ 2 ]

Describe whether ammonia acts as a Brønsted-Lowry acid or base in its reaction with water. Include an equation in your answer.

Conjugate Acid–Base Pairs

A conjugate base is what remains after an acid donates one proton. A conjugate acid is formed when a base accepts one proton; the pair differs by exactly one H+.

Remove H+ to find the conjugate base or add H+ to find the conjugate acid, then check the charge changes by one unit.

NH₄⁺/NH₃ and H₂CO₃/HCO₃⁻ are conjugate pairs because each pair differs by one H⁺. Removing H⁺ lowers charge by one; adding H⁺ raises it by one. Do not pair species merely because they occur on opposite sides of an equation—trace the specific proton transfer.

Deducing Conjugate Formulae

[ 2 ]

A solution of nitrous acid contains two conjugate acid-base pairs.

State the formulas of the conjugate acid and conjugate base in each pair.

Conjugate acid:
Conjugate base:
Conjugate acid:
Conjugate base:

Amphiprotic Species

An amphiprotic species can donate H+ in one reaction and accept H+ in another.
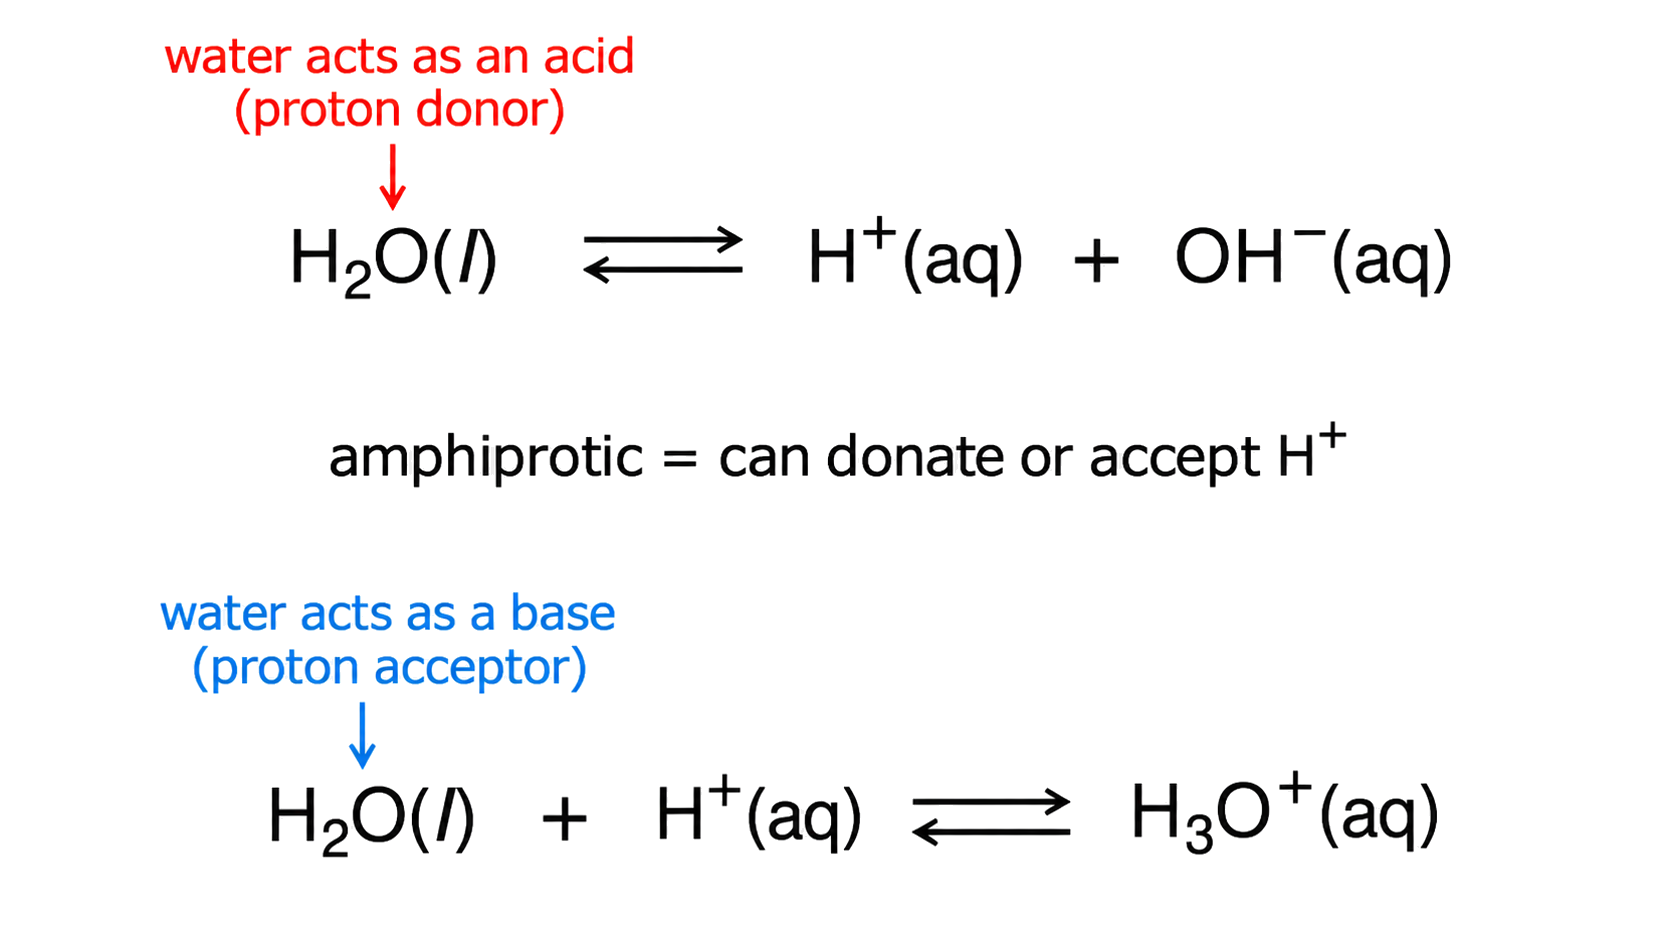
water donates a proton in H2O(l) ⇌ H+(aq) + OH−(aq); water accepts a proton in H2O(l) + H+(aq) ⇌ H3O+(aq); acid means proton donor and base means proton acceptor; amphiprotic is stated as the ability to donate or accept H+.

Write one equation in which the species becomes its conjugate base and another in which it becomes its conjugate acid.

For HCO₃⁻, donation gives CO₃²⁻ whereas acceptance gives H₂CO₃. Showing both reactions is the evidence for amphiprotic behaviour; one acid–base equation alone is insufficient.

Showing Amphiprotic Behaviour

[ 2 ]

Formulate two equations to show the amphiprotic nature of H2PO4−\mathrm{H}_{2} \mathrm{PO}_{4}^{-}.

pH and Hydrogen-Ion Concentration

pH=−log10[H+];[H+]=10(−pH)pH = −log10[H+]; [H+] = 10^(−pH)

pH is logarithmic: a one-unit change represents a tenfold concentration change. Universal indicator gives a colour range; a pH probe gives an instrumental pH measurement.

For [H⁺] = 2.0 × 10⁻³ mol dm⁻³, pH = 2.70; the leading 2 makes the answer non-integer. A colour indicator estimates a range, whereas a calibrated probe supports a numerical measurement.

Calculating pH and [H+]

[ 1 ]

A solution has a pH of 3.0 . What is the hydrogen ion concentration in the solution in moldm−3\mathrm{mol} \mathrm{dm}^{-3} ?

The Ion Product of Water

Kw=[H+][OH−]Kw = [H+][OH−]

Solution Ion comparison
acidic [H+] > [OH−]
neutral [H+] = [OH−]
basic [H+] < [OH−]

At 25 °C, Kw = 1.0 × 10⁻¹⁴, so a neutral solution has [H⁺] = [OH⁻] = 1.0 × 10⁻⁷ mol dm⁻³. Neutrality always means equal ion concentrations; neutral pH is not necessarily 7 when temperature changes.

At a fixed temperature, Kw is constant, so [OH-] = Kw/[H+]: a higher [H+] means a lower [OH-]. For example, at pH 9.3 and 25 C, [OH-] = 2.0 x 10^-5 mol dm^-3. Classify a solution from the ion comparison; do not assume neutral pH is 7 at every temperature.

Classifying Solutions with Kw

[ 1 ]

Calculate the concentration of hydroxide ions in an ammonia solution with pH=9.3. Use sections 1 and 2 of the data booklet.

Strong and Weak Acids and Bases

A strong acid or base ionizes completely in aqueous solution; a weak acid or base ionizes only partially. The equilibrium favours the weaker conjugate species.
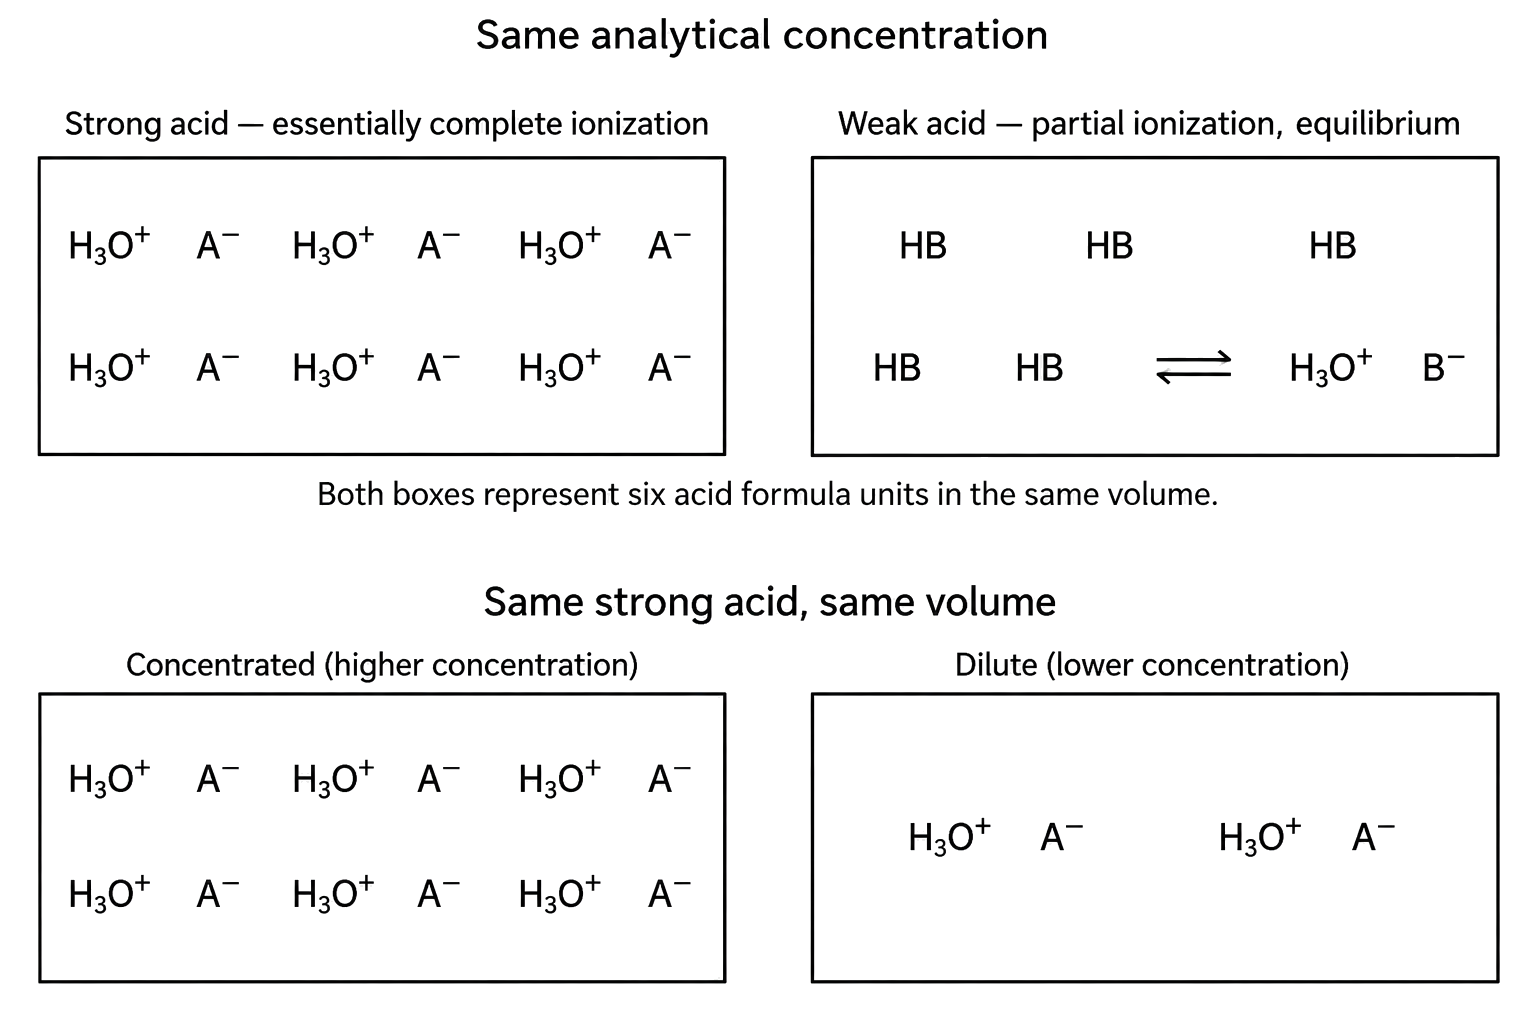
same-concentration strong acid shows six H3O+/A− pairs and no undissociated acid; same-concentration weak acid shows five HB plus one H3O+ and one B−, totalling six acid formula units; same-strength concentration comparison uses six versus two ion pairs in equal volumes; strength and concentration are explicitly independent.

Strength is the extent of ionization, whereas concentration is the amount of solute per volume. A concentrated weak acid can be more acidic than a dilute strong acid.

Represent a strong acid with essentially complete ionization and a weak acid with an equilibrium containing substantial undissociated acid. Strength is an equilibrium property, while concentration is an initial amount per volume; pH depends on both, so strength alone cannot rank arbitrary solutions.

Distinguishing Strength from Concentration

[ 2 ]

Explain the difference in pH .

Neutralization Reactions

Acids neutralize metal oxides and hydroxides to form salt and water. Carbonates and hydrogencarbonates also produce carbon dioxide when the reaction requires it; balance all formulae and coefficients.

Identify the parent acid and parent base of a salt by tracing its anion and cation back to the neutralization reactants.

Balance proton capacity as well as atoms: H₂SO₄ + 2NaOH → Na₂SO₄ + 2H₂O, while an acid–carbonate reaction also releases CO₂. To identify parents of Na₂SO₄, trace SO₄²⁻ to the acid and Na⁺ to the base rather than treating the salt name as a reaction equation.

Writing Neutralization Equations

[ 1 ]

Write two equations showing how these antacids neutralize excess hydrochloric acid.

Magnesium carbonate:

Aluminium hydroxide:

Strong-Acid–Strong-Base Titration Curves

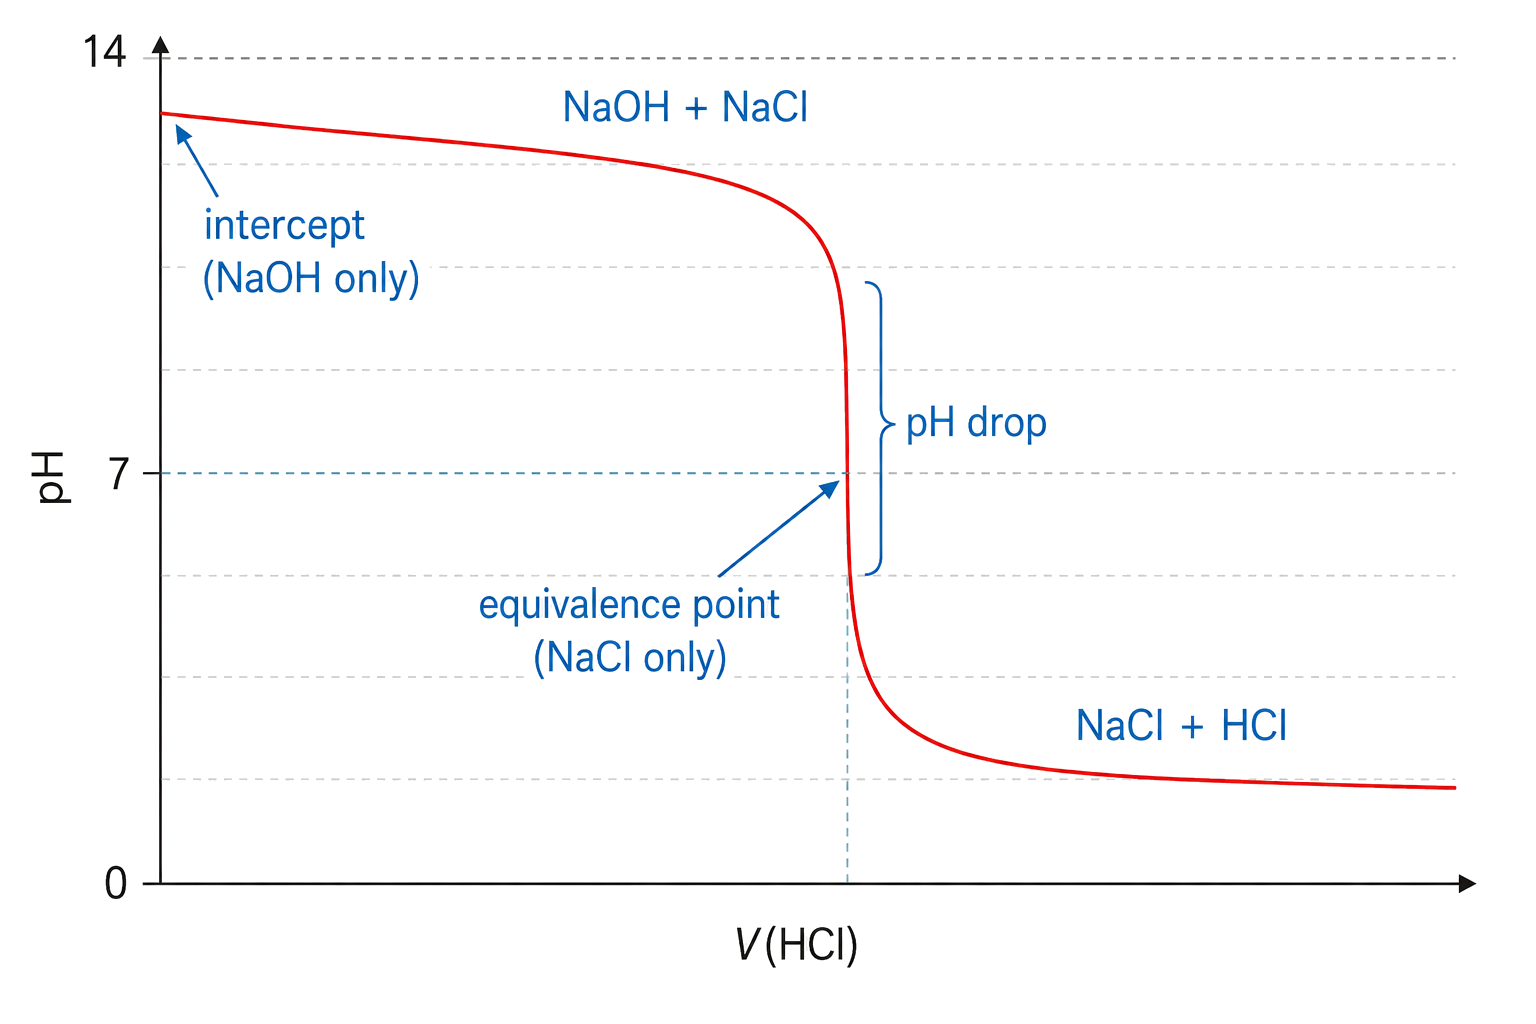
The equivalence point is where stoichiometric amounts of analyte and titrant have reacted. A monoprotic strong-acid–strong-base curve has a steep neutral region centred at the equivalence point.
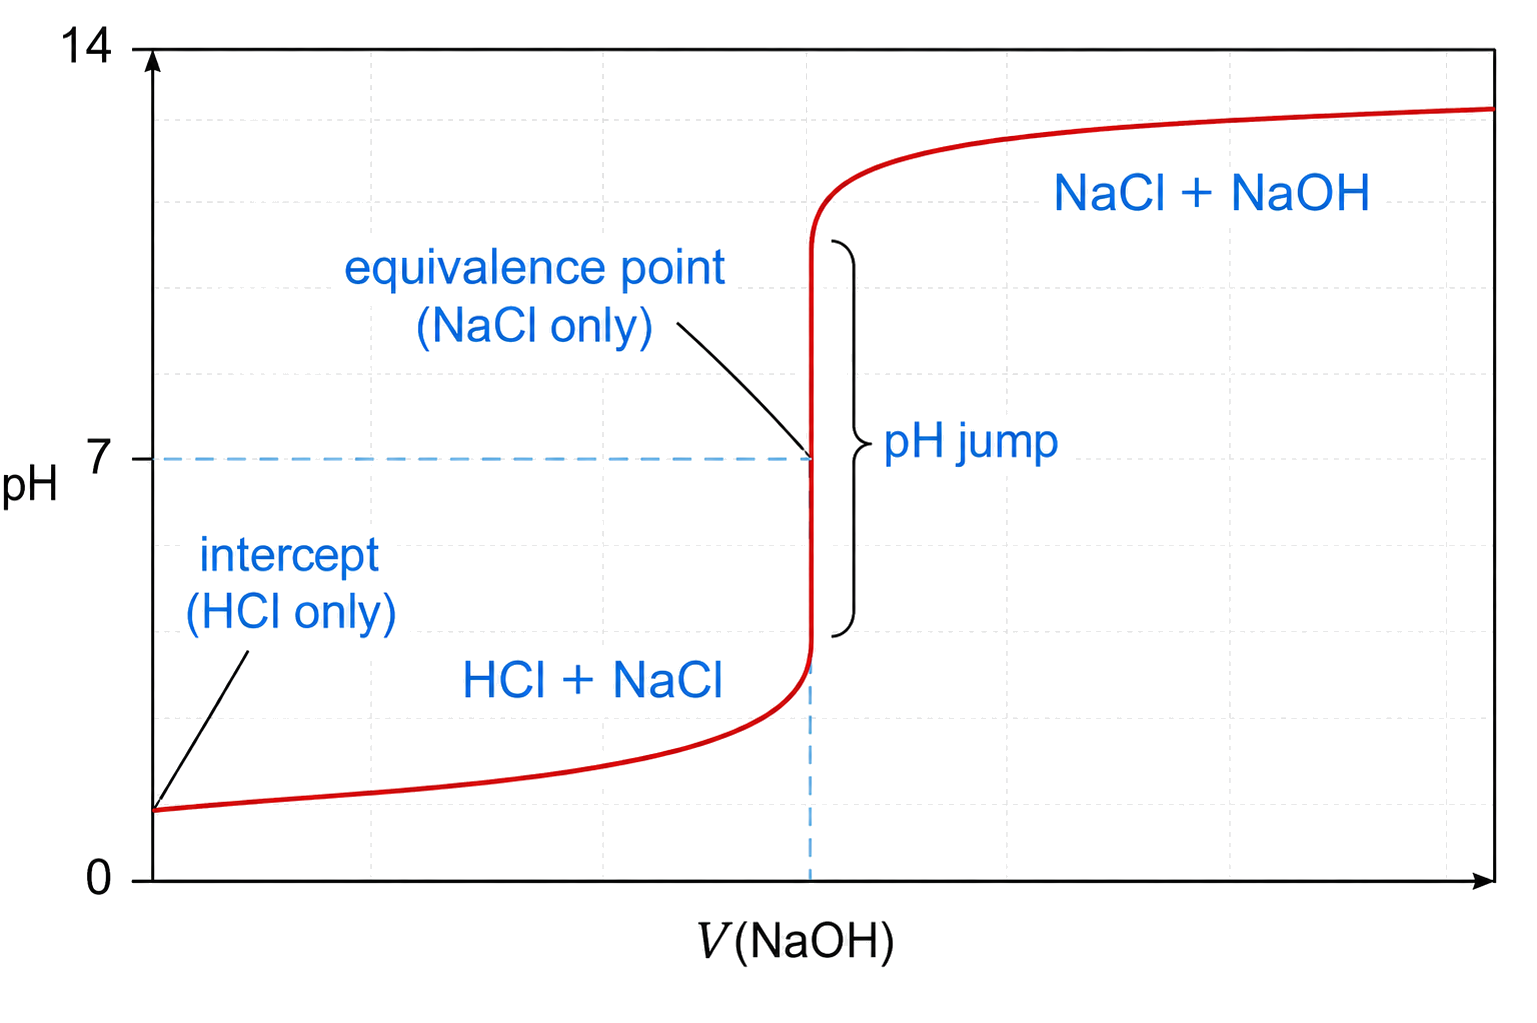
HCl is the strong-acid analyte and NaOH is the titrant; the curve rises from low pH and the equivalence point is on pH 7; HCl plus NaCl appears before equivalence and NaCl plus NaOH after equivalence; the steep region is identified as a pH jump without inventing a volume.
NaOH is the strong-base analyte and HCl is the titrant; the curve falls from high pH and the equivalence point is on pH 7; NaOH plus NaCl appears before equivalence and NaCl plus HCl after equivalence; the steep region is identified as a pH drop without inventing a volume.

Read the initial pH, steep intercept region and final plateau; curve direction depends on whether acid or base is added.

For a strong acid titrated with strong base at 25 °C, calculate the initial pH from excess acid, locate equivalence from stoichiometric moles, and place the steep section around pH 7. Equivalence is a mole condition; it is not the same as equal solution volumes unless concentrations and stoichiometry make it so.

Interpreting a Strong Titration Curve

[ 1 ]

Which graph would be obtained by adding 0.10moldm−3HCl(aq)0.10 \mathrm{moldm}^{-3} \mathrm{HCl}(\mathrm{aq}) to 25 cm325 \mathrm{~cm}^{3} of 0.10moldm−3NaOH(aq)0.10 \mathrm{moldm}^{-3} \mathrm{NaOH}(\mathrm{aq}) ?

pOH and Ion Concentrations

HL only

pOH=−log10[OH−];[OH−]=10(−pOH);pH+pOH=14at25°CpOH = −log10[OH−]; [OH−] = 10^(−pOH); pH + pOH = 14 at 25 °C

Move between pH and pOH, then between the logarithm and concentration. Keep the 25 °C condition attached to pH+pOH=14.

A reliable route is [OH⁻] → pOH → pH → [H⁺], or the reverse, with each logarithm shown. Use pH + pOH = pKw; replacing pKw by 14 is valid only at 25 °C.

Worked pOH example: for 0.025 mol dm−30.025\,\mathrm{mol\,dm^{-3}} KOH(aq)\ce{KOH(aq)}, complete dissociation gives [OHX−]=0.025 mol dm−3[\ce{OH^-}]=0.025\,\mathrm{mol\,dm^{-3}}. Therefore pOH=−log⁡10(0.025)=1.60\mathrm{pOH}=-\log_{10}(0.025)=1.60. The low pOH is consistent with a basic solution; at 298 K, pH=14.00−1.60=12.40\mathrm{pH}=14.00-1.60=12.40.

Interconverting pH and pOH

HL only

[ 1 ]

What is the concentration of OH−(aq)\mathrm{OH}^{-}(\mathrm{aq}), in moldm−3\mathrm{mol} \mathrm{dm}^{-3}, in a solution at 298.15 K with a pH of 4.50 ?

pH=−log⁡10[H+][H+]=10−pHKw=[H+][OH−]Kw=1.00×10⁻¹⁴ mol2dm−6\mathrm{pH}=-\log _{10}\left[\mathrm{H}^{+}\right] \quad\left[\mathrm{H}^{+}\right]=10^{-\mathrm{pH}} \quad K_{\mathrm{w}}=\left[\mathrm{H}^{+}\right]\left[\mathrm{OH}^{-}\right] \quad K_{\mathrm{w}}=1.00 \times 10^{-14} \mathrm{~mol}^{2} \mathrm{dm}^{-6}

Ka, Kb, pKa and pKb

HL only

Ka=[A−][H3O+]/[HA];Kb=[BH+][OH−]/[B]Ka = [A−][H3O+] / [HA]; Kb = [BH+][OH−] / [B]

Ka and Kb measure dissociation extent. pKa = −log Ka, so a lower pKa indicates a stronger acid; use the corresponding comparison for bases and pKb.

Large Ka and small pKa both indicate the stronger acid; large Kb and small pKb indicate the stronger base. Strength describes extent of ionization, whereas concentration describes amount per volume—dilute and weak are not synonyms.

Interpreting Ka and pKa

HL only

[ 1 ]

State the KaK_{\mathrm{a}} expression for ethanoic acid.

Conjugate Acid–Base Constants

HL only

Ka×Kb=KwKa × Kb = Kw

For a conjugate pair, calculate the missing constant by dividing Kw by the known Ka or Kb, keeping the pair direction consistent.

Match the constants to one conjugate pair: Ka(HA) × Kb(A⁻) = Kw. A stronger acid therefore has a weaker conjugate base at the same temperature; do not multiply constants belonging to unrelated species.

Worked conjugate-constant example at 298 K: methylamine has pKb=3.34\mathrm{p}K_b=3.34, so for its conjugate acid CHX3NHX3X+\ce{CH3NH3+}, pKa=pKw−pKb=14.00−3.34=10.66\mathrm{p}K_a=\mathrm{p}K_w-\mathrm{p}K_b=14.00-3.34=10.66. Equivalently, KaKb=KwK_aK_b=K_w. This relationship applies only to a conjugate acid–base pair at the same temperature.

Calculating a Conjugate Constant

HL only

[ 1 ]

Calculate the KbK_{b} of the conjugate base of ethanoic acid using sections 2 and 21 of the data booklet.

Salt Hydrolysis

HL only

Trace each salt ion to its parent acid or base. A conjugate base from a weak acid can hydrolyse water to produce OH− and an alkaline solution; a conjugate acid from a weak base can produce H3O+.
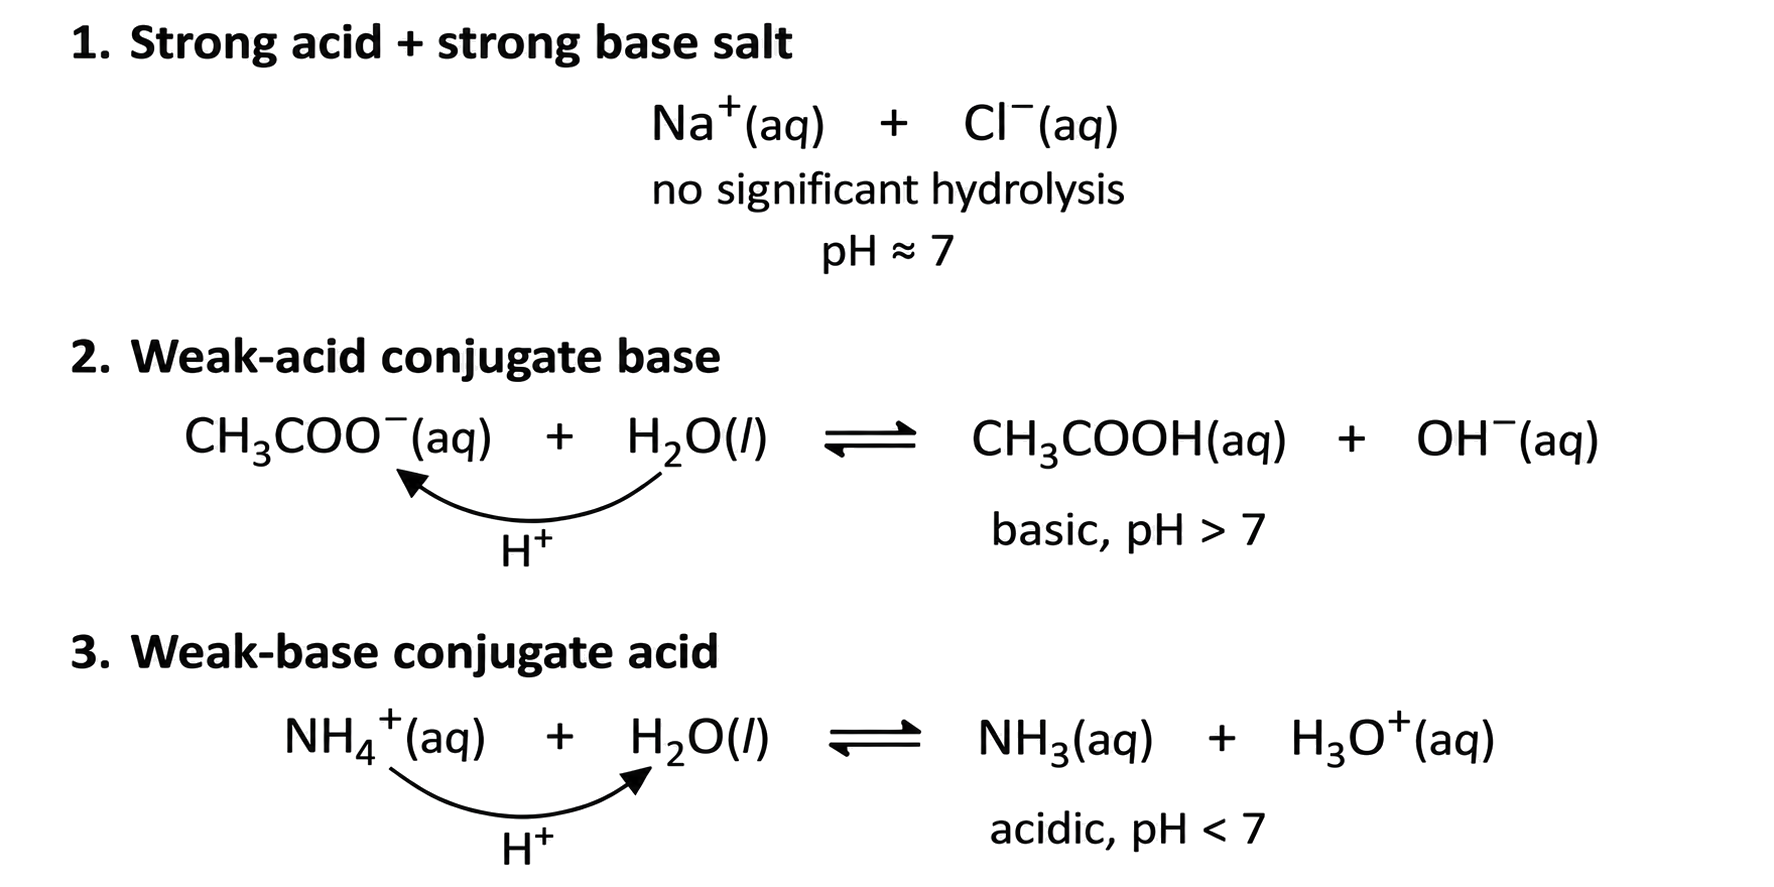
Na+ and Cl− show no significant hydrolysis and pH approximately 7; CH3COO− accepts H+ from water and forms OH−, so pH > 7; NH4+ donates H+ to water and forms H3O+, so pH < 7; all charges, phases and equilibrium arrows are correct.

A−+H2O⇌HA+OH−A− + H2O ⇌ HA + OH−

Spectator ions from strong parents do not control pH. NH₄Cl is acidic because NH₄⁺ donates a proton to water, while a carbonate salt is basic because CO₃²⁻ accepts one; write the hydrolysing ion, not the intact salt, in the equilibrium.

Predicting Salt-Solution pH

HL only

[ 1 ]

Explain, with reference to acid-base equilibria, why the sodium benzoate solution formed has a pH>7.

Weak-Acid and Weak-Base Titration Curves

HL only

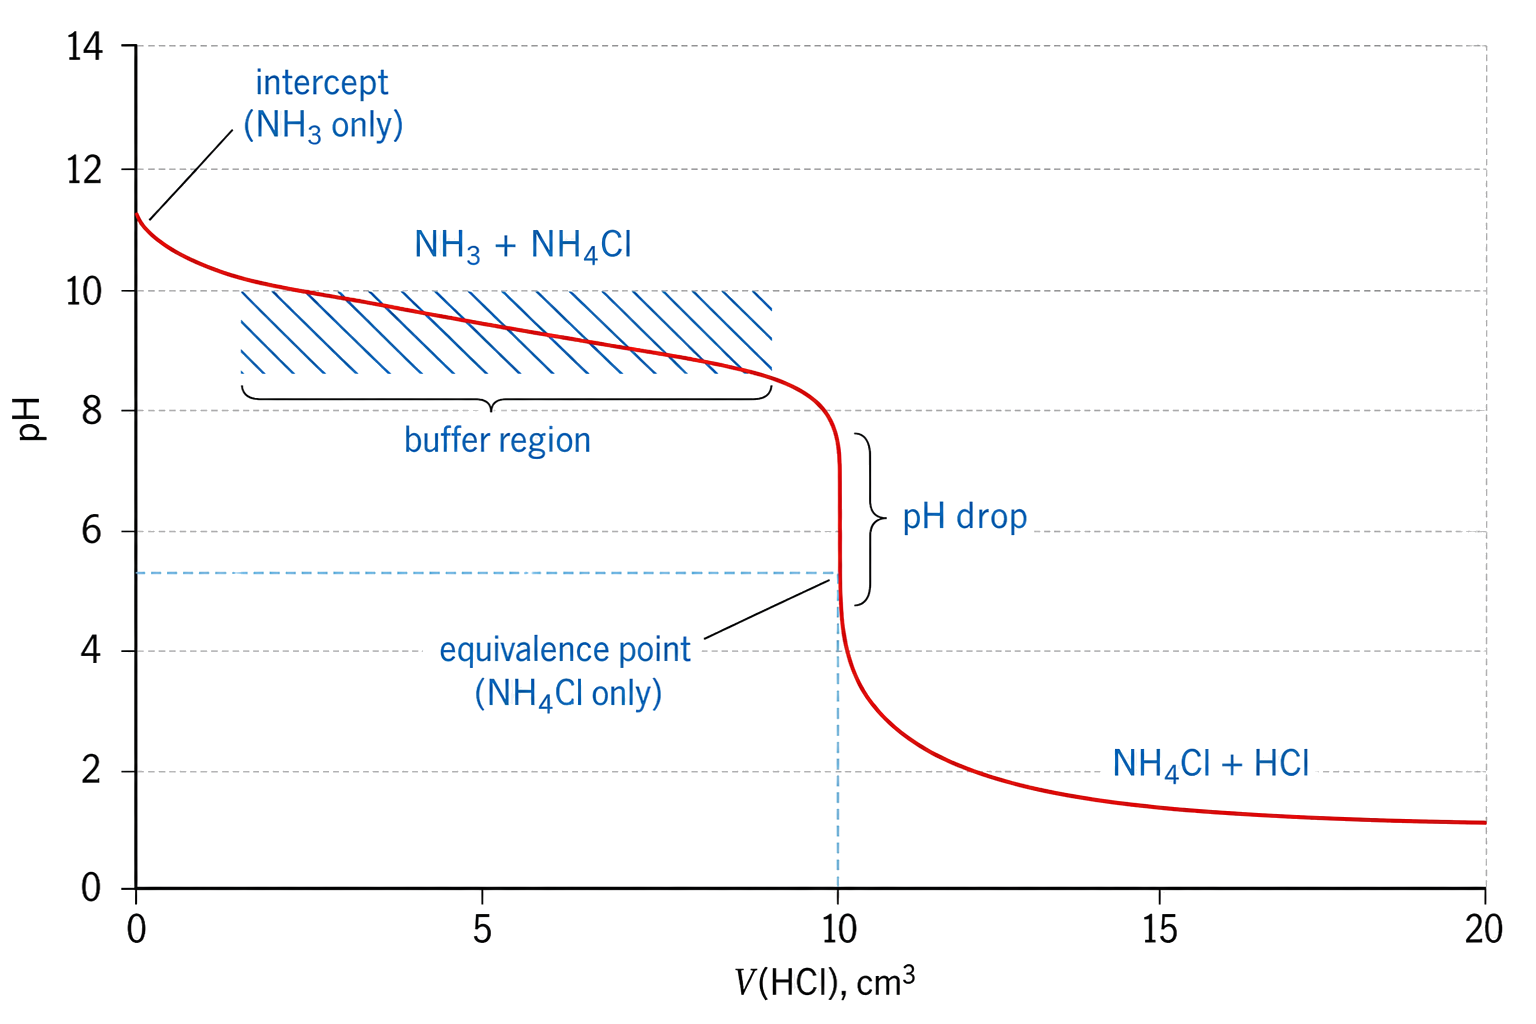
Compare all four strong/weak combinations by starting pH, buffer region, equivalence-point pH and steep-section position. For a weak acid titrated with strong base, half-equivalence gives pH = pKa and the equivalence solution is basic. For a weak base titrated with strong acid, half-equivalence gives pOH = pKb (then pH = pKw − pOH) and the equivalence solution is acidic. Weak–weak curves often lack a sufficiently steep indicator region.
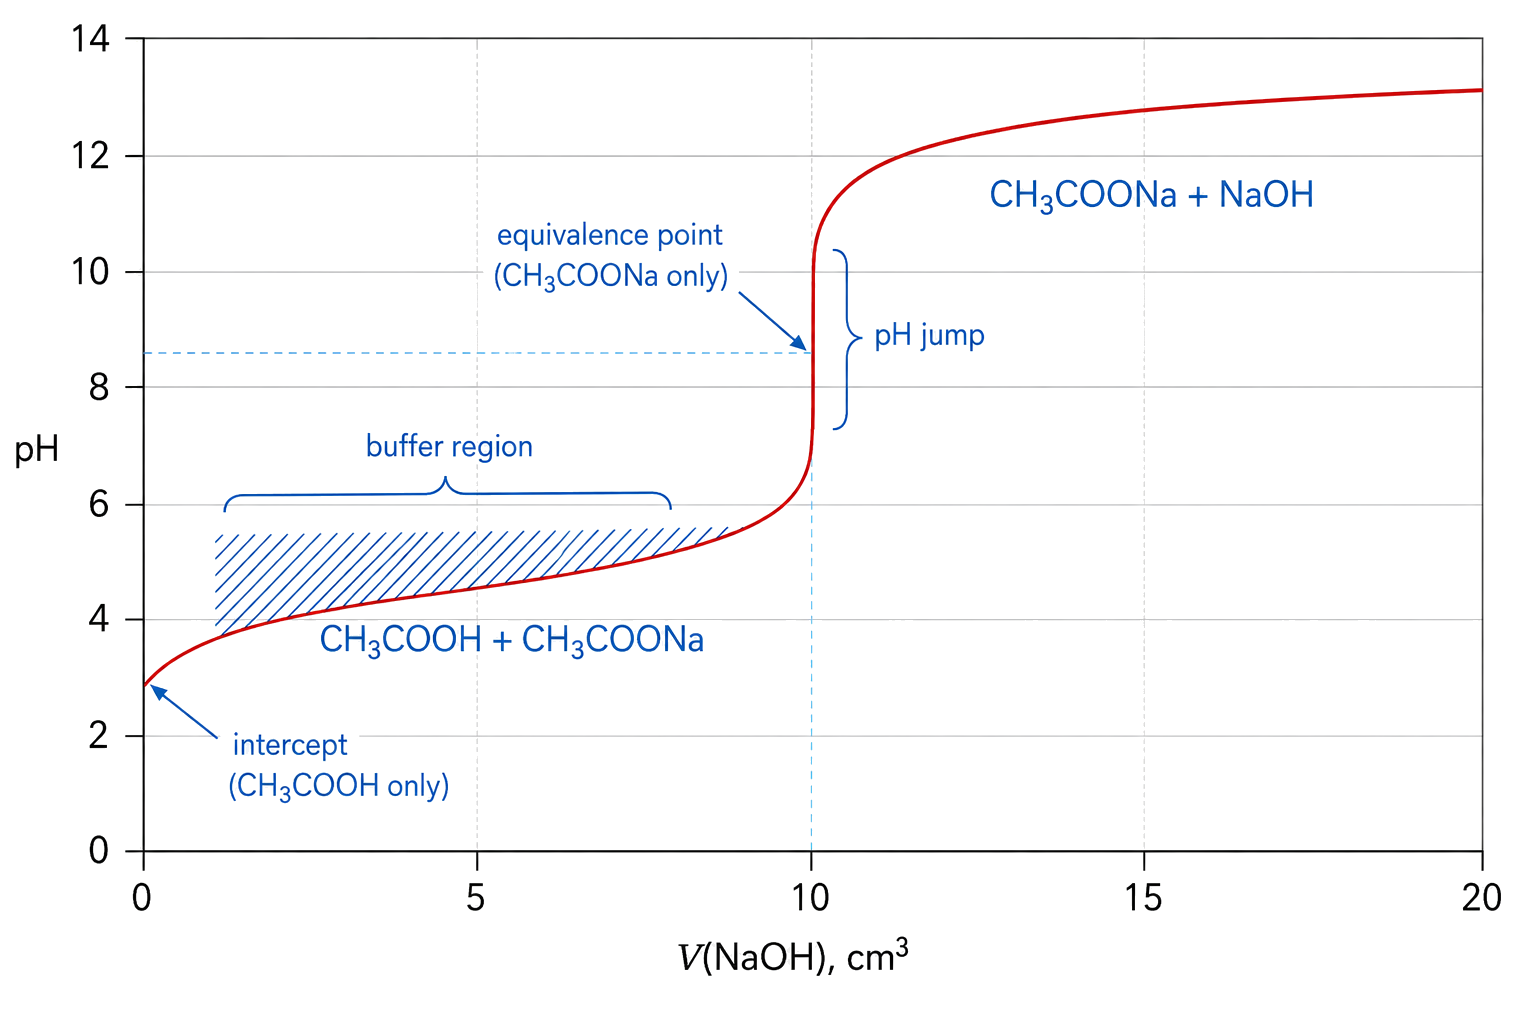
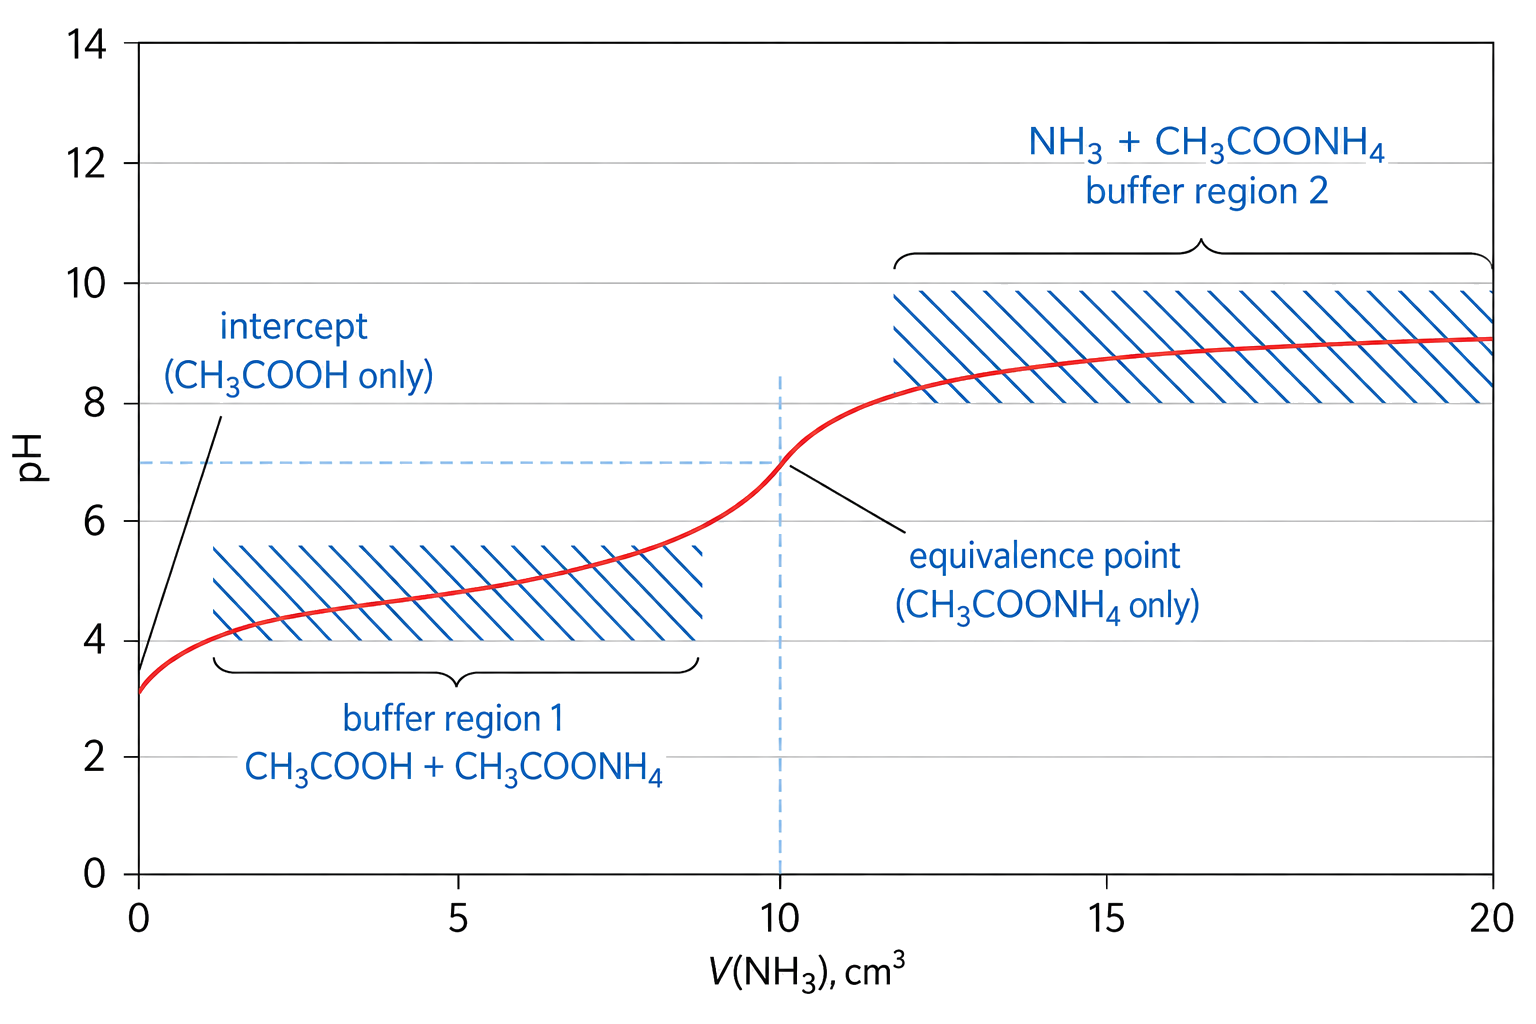
CH3COOH is titrated with NaOH and the curve rises from about pH 3; the pre-equivalence buffer region and CH3COOH plus CH3COONa composition are preserved; equivalence occurs at 10 cm3 above pH 7 with CH3COONa only; post-equivalence composition is CH3COONa plus NaOH and the curve approaches high pH.
NH3 is titrated with HCl and the curve falls from about pH 11; the pre-equivalence buffer region and NH3 plus NH4Cl composition are preserved; equivalence occurs at 10 cm3 below pH 7 with NH4Cl only; post-equivalence composition is NH4Cl plus HCl and the curve approaches low pH.
CH3COOH is titrated with NH3 and the equivalence point is near pH 7 at 10 cm3; the curve changes gradually and does not falsely show a sharp pH jump; buffer region 1 contains CH3COOH plus CH3COONH4 before equivalence; buffer region 2 contains NH3 plus CH3COONH4 after equivalence.

A buffer region contains appreciable weak acid and conjugate base, so the pH changes relatively slowly as titrant is added.

A weak acid–strong base curve starts at a higher pH than an equally concentrated strong acid, contains a buffer region, has pH = pKa at half-equivalence, and has an alkaline equivalence point from conjugate-base hydrolysis. Weak–weak titrations often lack a sufficiently steep jump for a simple indicator endpoint.

Reading Weak Titration Curves

HL only

[ 1 ]

Annotate the graph to find the pKa\mathrm{p} K_{\mathrm{a}} of benzoic acid.

Acid–Base Indicators

HL only

HInd+H2O⇌H3O++Ind−HInd + H2O ⇌ H3O+ + Ind−

HInd and Ind− have different colours. Changing pH shifts their ratio; the visible transition occurs around pH ≈ pKa. Universal indicator is a mixture of indicators with different transition ranges.

Added acid shifts HInd ⇌ H⁺ + Ind⁻ toward the HInd colour; added base favours Ind⁻. The visible transition spans a range around pKa because both colours must change in relative abundance. Universal indicator combines several such equilibria and is not one substance with every colour.

Explaining Indicator Colour Change

HL only

[ 2 ]

Explain how the indicator HInd, that is a weak acid, shows changes in pH using an equation.

Choosing an Indicator

HL only

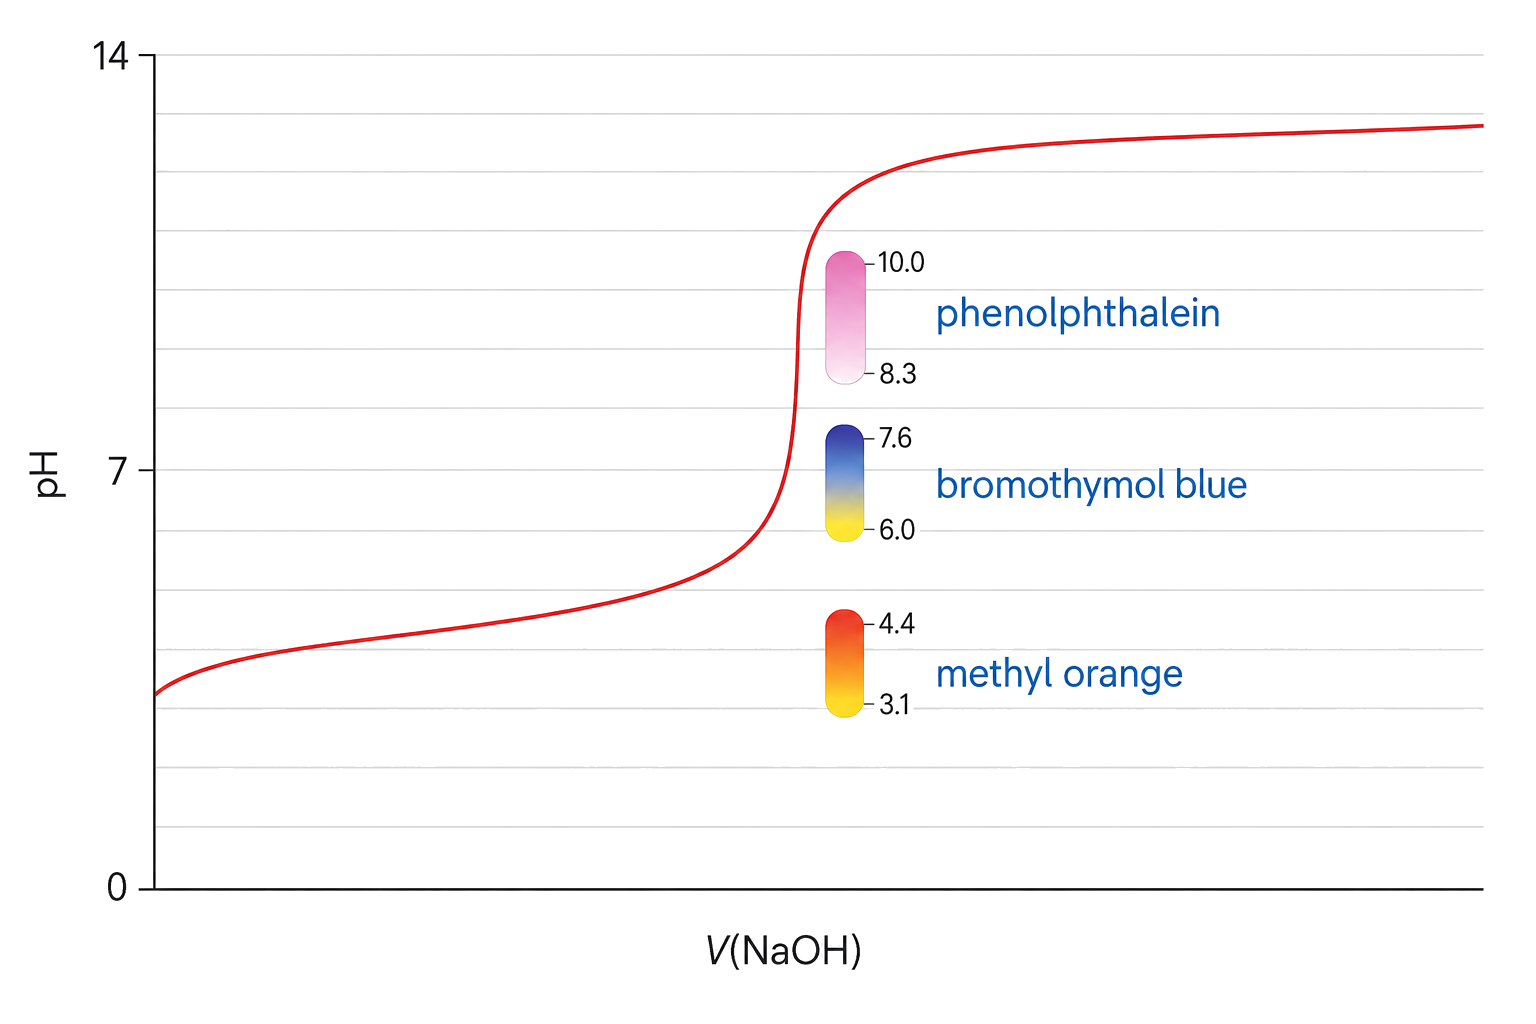
Choose an indicator whose endpoint transition range lies within the steep pH change around the equivalence point. Use salt identity to predict whether that equivalence pH is acidic, neutral or alkaline.
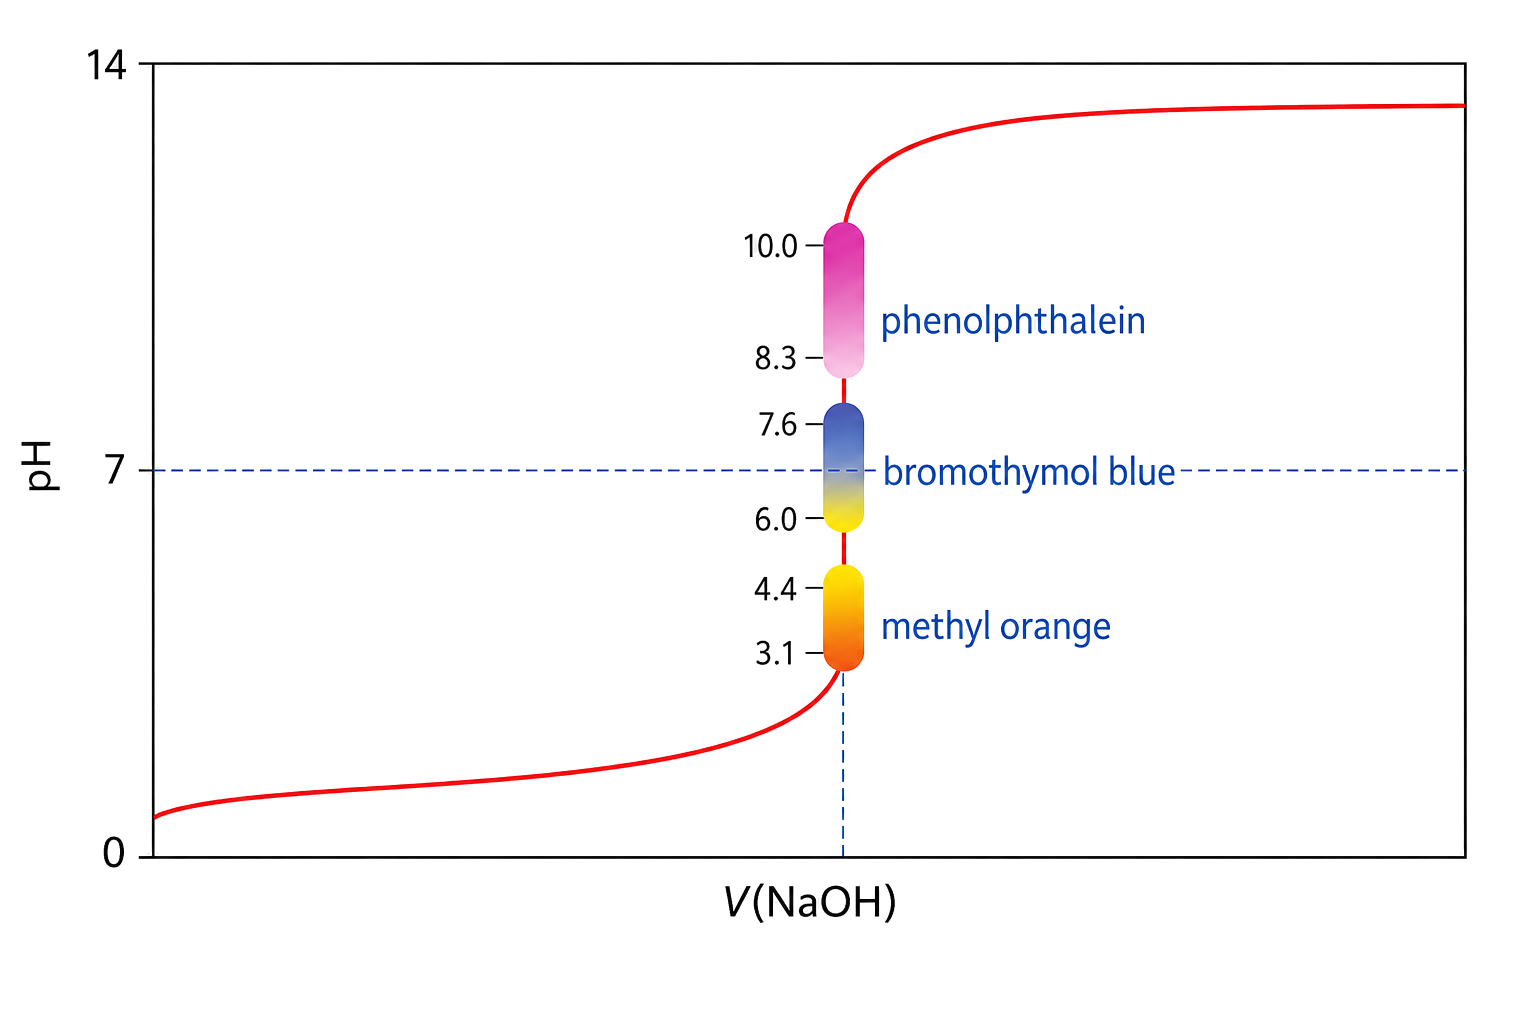
the strong-acid strong-base curve has a large pH jump centered at pH 7; methyl orange spans pH 3.1 to 4.4, bromothymol blue 6.0 to 7.6, and phenolphthalein 8.3 to 10.0; all three indicator ranges overlap the steep region; the pH axis contains only correct 0, 7, and 14 master labels.
the weak-acid strong-base curve starts near pH 3 and has equivalence above pH 7; phenolphthalein lies within the steep region and is shown as the best range; bromothymol blue overlaps only the lower edge of the steep region; methyl orange lies before the steep region and is not implied to be suitable.

The equivalence point is the stoichiometric condition; the endpoint is the observed indicator colour change. A good indicator makes them coincide closely.

Overlay the indicator transition range on the titration curve and require the whole visible change to fall inside the steep region. A weak acid–strong base equivalence is alkaline and favours an alkaline-range indicator; a strong acid–weak base equivalence is acidic. Endpoint proximity, not a memorized indicator name alone, is the criterion.

Matching Indicator Range to Equivalence pH

HL only

[ 1 ]

What is the best indicator to use in the titration of phenylamine with nitric acid?

Buffer Solutions

HL only

An acidic buffer contains a weak acid and its conjugate base; a basic buffer contains a weak base and its conjugate acid. The pair resists pH change when small amounts of strong acid or base are added.
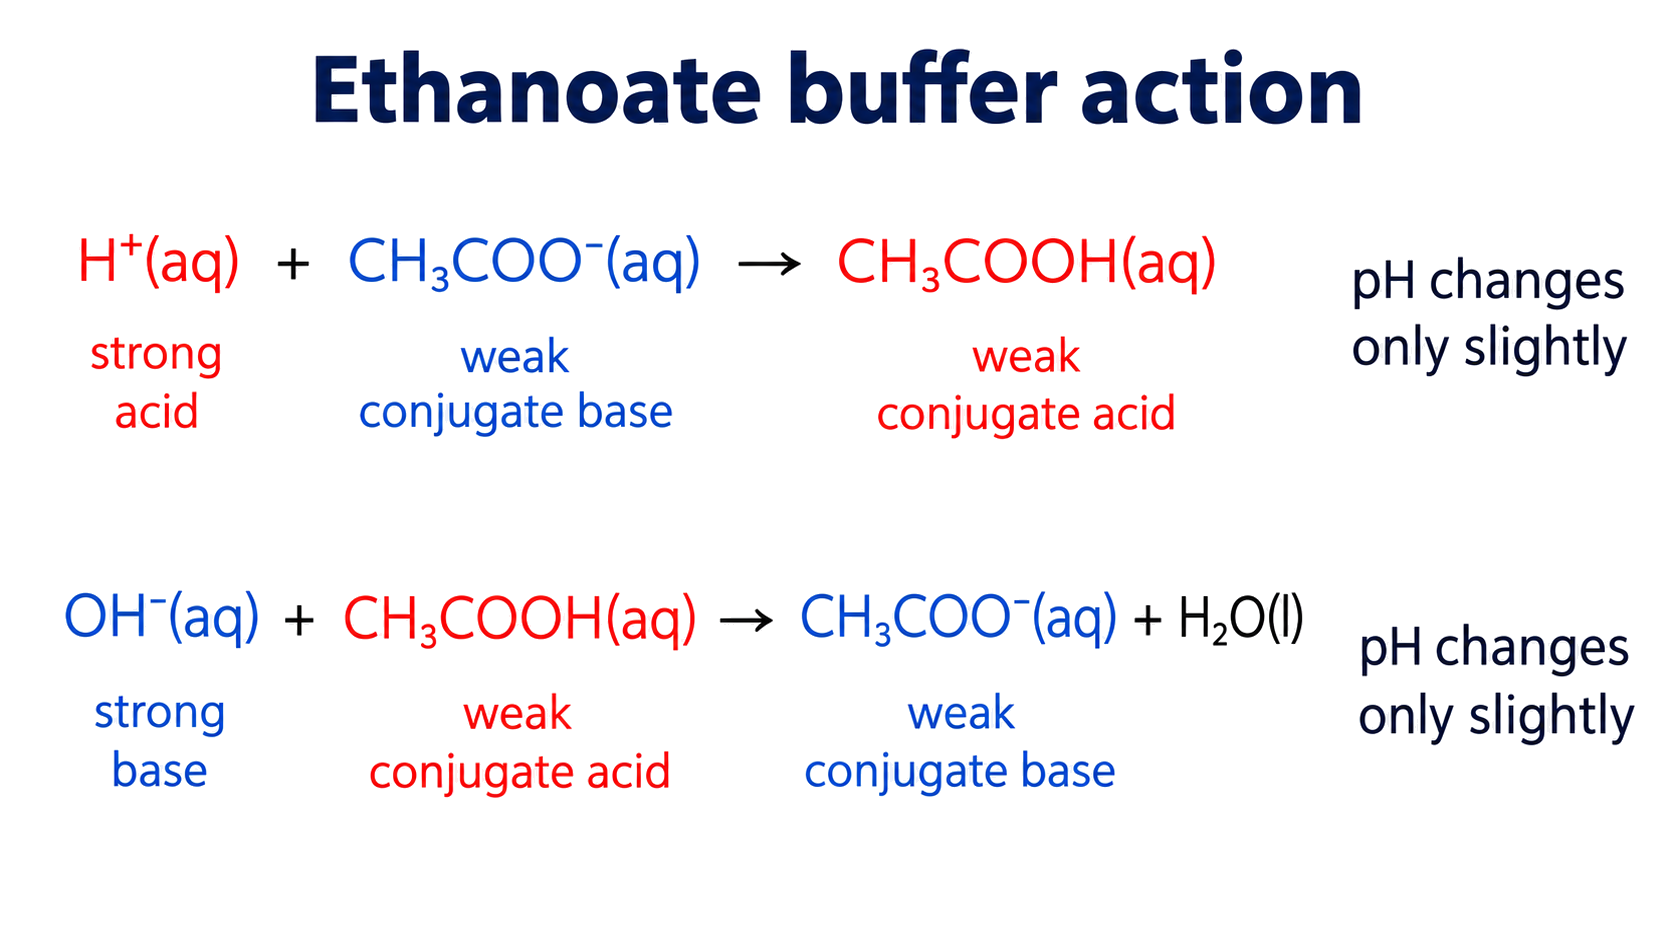
added H+ is consumed by CH3COO− to form CH3COOH; added OH− is consumed by CH3COOH to form CH3COO− and H2O; strong and weak conjugate roles are assigned correctly; the pH is described as changing only slightly, not as absolutely unchanged.

The conjugate base consumes added H+; the weak acid equilibrium supplies H+ when added OH− removes it. In both cases the conjugate equilibrium shifts to oppose the change.

In an ethanoic acid/ethanoate buffer, CH₃COO⁻ consumes added H⁺ and CH₃COOH consumes added OH⁻, so the conjugate ratio changes only slightly. A buffer resists small additions but has finite capacity; once one component is nearly exhausted, the pH can change sharply.

Explaining Buffer Action

HL only

[ 2 ]

Write equations to show the action of the buffer solution when small amounts of a strong acid or a strong base are added.

Addition of strong acid:
Addition of strong base:

Buffer pH and Composition

HL only

pH≈pKa+log10([A−]/[HA])pH ≈ pKa + log10([A−]/[HA])

Buffer pH depends on pKa and the conjugate-to-parent ratio. Dilution changes both concentrations by the same factor, so the ratio and pH remain approximately constant.

When a small amount of acid is added, A⁻ removes it to form HA; added base is removed by HA to form A⁻. Dilution leaves their ratio nearly unchanged but reduces buffer capacity, so resistance to a large addition is not unchanged.

basicbuffer:pOH≈pKb+log10([BH+]/[B]);thenpH=pKw−pOHbasic buffer: pOH ≈ pKb + log₁₀([BH⁺]/[B]); then pH = pKw − pOH

Worked buffer example: an ethanoate buffer contains 0.100 mol dm−30.100\,\mathrm{mol\,dm^{-3}} CHX3COOH\ce{CH3COOH} and 0.200 mol dm−30.200\,\mathrm{mol\,dm^{-3}} CHX3COOX−\ce{CH3COO^-}, with pKa=4.76\mathrm{p}K_a=4.76. Substitution gives pH=pKa+log⁡10([AX−]/[HA])=4.76+log⁡10(0.200/0.100)=5.06\mathrm{pH}=\mathrm{p}K_a+\log_{10}([\ce{A^-}]/[\ce{HA}])=4.76+\log_{10}(0.200/0.100)=5.06. The pH is above pKa\mathrm{p}K_a because the conjugate base is more concentrated than the acid.

Calculating Buffer Composition

HL only

[ 3 ]

This 1.00 moldm−31.00 \mathrm{~mol} \mathrm{dm}^{-3} solution of nitrous acid was used to prepare a buffer with pH 3.00 .

Calculate the concentration of the conjugate base of nitrous acid required to make this buffer. The pKa\mathrm{pK}_{\mathrm{a}} of nitrous acid is 3.25.

Concentration of conjugate base:

Proton Transfer Reactions Summary

Retrieve the route: track proton transfer and conjugates, calculate pH and Kw, distinguish strength, balance neutralization, read titration curves, use Ka/Kb and hydrolysis, select indicators, and explain and calculate buffer behaviour.

Check donor versus acceptor, one-proton differences, logarithm direction, ion comparison, strength versus concentration, equivalence versus endpoint, pKa landmarks, conjugate equations and dilution ratios.

3.2 Electron transfer reactions

Syllabus
First assessment 2025
Topic
3.2
Level
HL

Oxidation and Reduction

Oxidation is loss of electrons and an increase in oxidation state; reduction is gain of electrons and a decrease. The oxidizing agent is reduced, and the reducing agent is oxidized.

Use the oxidation-state rules and total charge to identify which species changed and which agent caused the change.

In Zn + Cu²⁺ → Zn²⁺ + Cu, Zn rises from 0 to +2 and is oxidized, so it is the reducing agent; Cu²⁺ falls from +2 to 0 and is reduced, so it is the oxidizing agent. Name agents from what happens to them, not from the process they cause in the other species.

Identifying Redox Agents

[ 2 ]

Identify the oxidising and reducing agents, and the species oxidised and reduced, in the forward reaction.

CO(g)\mathbf{C O}(\mathbf{g})H2O(g)\mathbf{H}_{\mathbf{2}} \mathbf{O}(\mathbf{g})
oxidising or reducing agent?
species oxidised or reduced?

Redox Half-Equations

Separate oxidation and reduction, balance atoms, add H2O and H+ in acidic solution as needed, balance charge with electrons, then multiply to cancel electrons before adding.

A valid full redox equation conserves atoms and charge and contains no uncancelled electrons.

For MnO₄⁻ → Mn²⁺ in acid, balance O with 4H₂O, H with 8H⁺ and charge with 5e⁻: MnO₄⁻ + 8H⁺ + 5e⁻ → Mn²⁺ + 4H₂O. After combining halves, cancel electrons and any identical H⁺ or H₂O, then recheck both atoms and net charge.

To adapt an acidic half-equation to neutral or basic conditions, first balance it with H₂O, H⁺ and e⁻. Add the same number of OH⁻ to both sides to neutralize every H⁺, replace H⁺+OH⁻ by H₂O, then cancel water appearing on both sides. Recheck atoms and total charge; do not leave free H⁺ in a stated neutral medium unless the chemistry justifies it.

Balancing Redox Equations

[ 2 ]

The reaction continues until the violet colour disappears. The thiosulfate ion, S2O32−\mathrm{S}_{2} \mathrm{O}_{3}{ }^{2-}, is oxidized to SO2\mathrm{SO}_{2}, and Fe3+\mathrm{Fe}^{3+} is reduced to Fe2+\mathrm{Fe}^{2+}. Deduce the oxidation half-equation, and the overall redox equation for this second step of the reaction.

Oxidation half-equation:
Overall redox equation:

Redox Displacement

A more active metal more readily donates electrons to a less active metal ion. A halogen with greater reduction tendency oxidizes the halide of a weaker halogen.

Test a predicted displacement by placing one metal in the other metal's sulfate or comparing supplied electrode data.

Zinc displaces Cu²⁺ because Zn more readily oxidizes: Zn + Cu²⁺ → Zn²⁺ + Cu. Chlorine displaces Br⁻ because Cl₂ more readily reduces. Keep the metal and halogen trends in their correct electron directions instead of using one vague 'more reactive' rule.

Predicting Displacement Reactions

[ 2 ]

Discuss how the relative reactivity of copper and thallium could be established using the metals and aqueous solutions of their sulfates.

Metals with Dilute Acids

A metal above hydrogen in the activity series can donate electrons to acid and release hydrogen gas; a metal below hydrogen, such as copper, does not react with dilute hydrochloric acid.

metal+acid→salt+H2(g)metal + acid → salt + H2(g)

Balance the electron transfer behind the molecular equation: metal atoms are oxidized and 2H⁺ + 2e⁻ → H₂ is the reduction. Use the metal charge and acid anion to construct the salt rather than assuming every metal forms a 2+ ion.

Predicting Hydrogen Release

[ 2 ]

Outline, using an ionic equation, what is observed when magnesium powder is added to a solution of ammonium chloride.

Anodes, Cathodes and Polarity

Oxidation always occurs at the anode and reduction always occurs at the cathode. In a voltaic cell the anode is negative and cathode positive; in an electrolytic cell the anode is positive and cathode negative.
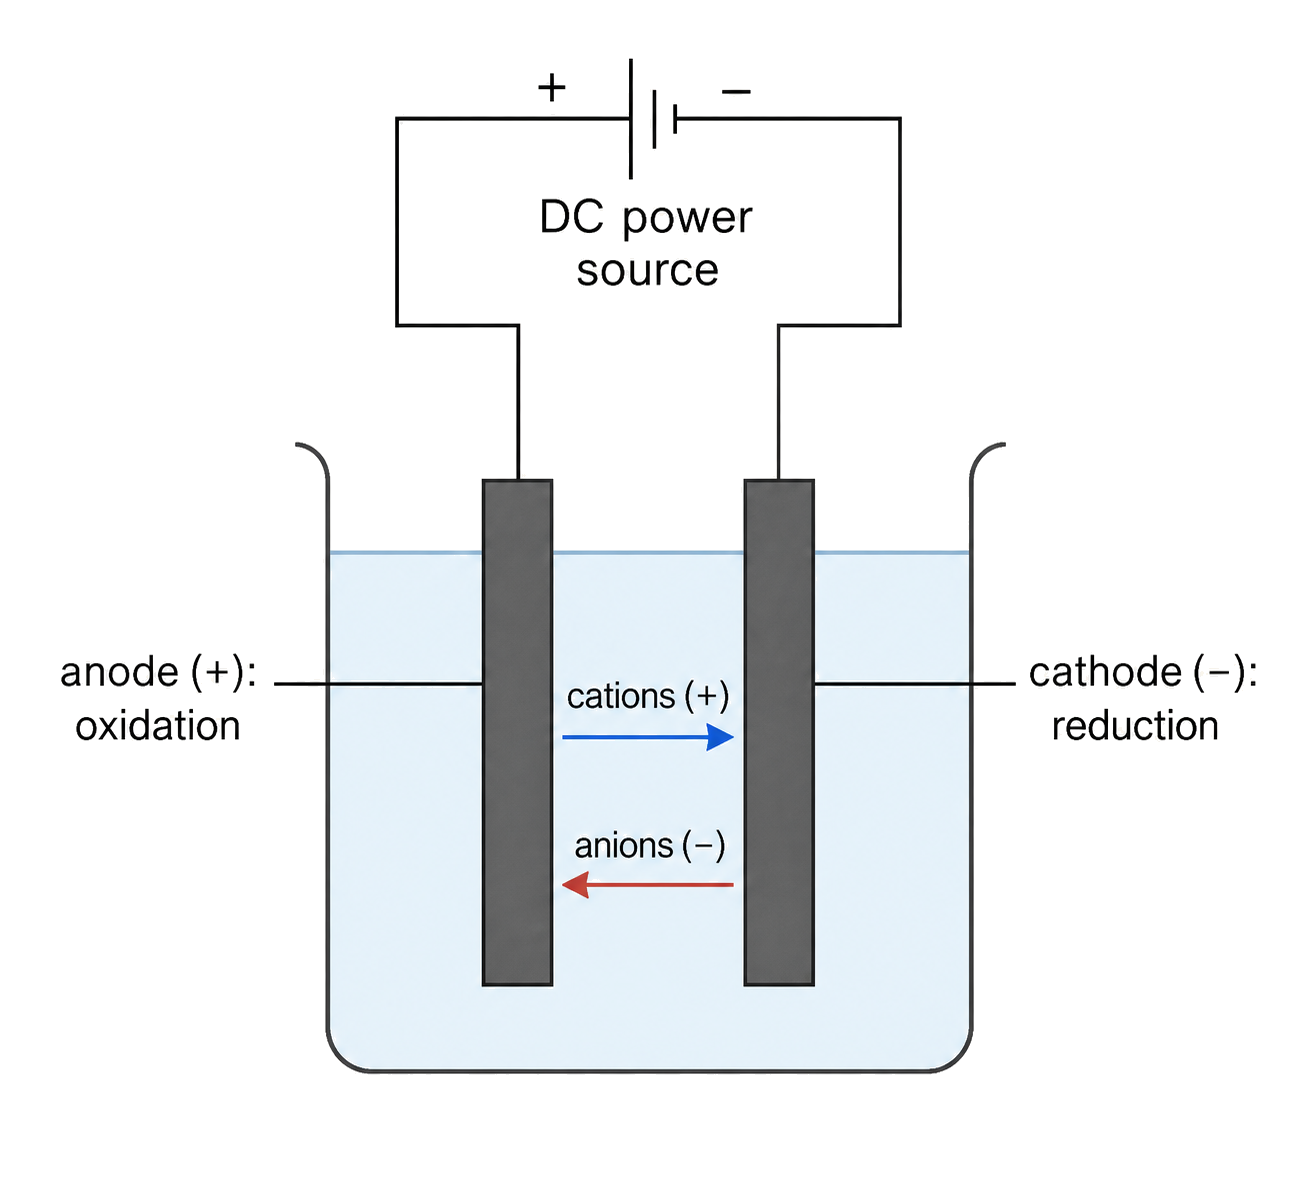
the electrolytic anode is positive and is the site of oxidation; the electrolytic cathode is negative and is the site of reduction; cations move to the cathode and anions move to the anode; the external DC source is explicit and no spontaneous operation is implied.

Name electrodes from the half-reactions before assigning signs. Electrons leave the anode and reach the cathode through the external circuit; a power supply reverses the polarities in an electrolytic cell but never changes where oxidation and reduction occur.

Labelling Electrochemical Cells

[ 2 ]

Annotate the electrolytic cell with the terms anode and cathode, and show the direction of ion movement.

Voltaic Cells

A voltaic cell uses a spontaneous redox reaction to convert chemical energy to electrical energy. Electrons flow through the wire from anode to cathode; the salt bridge carries ions to maintain charge neutrality.
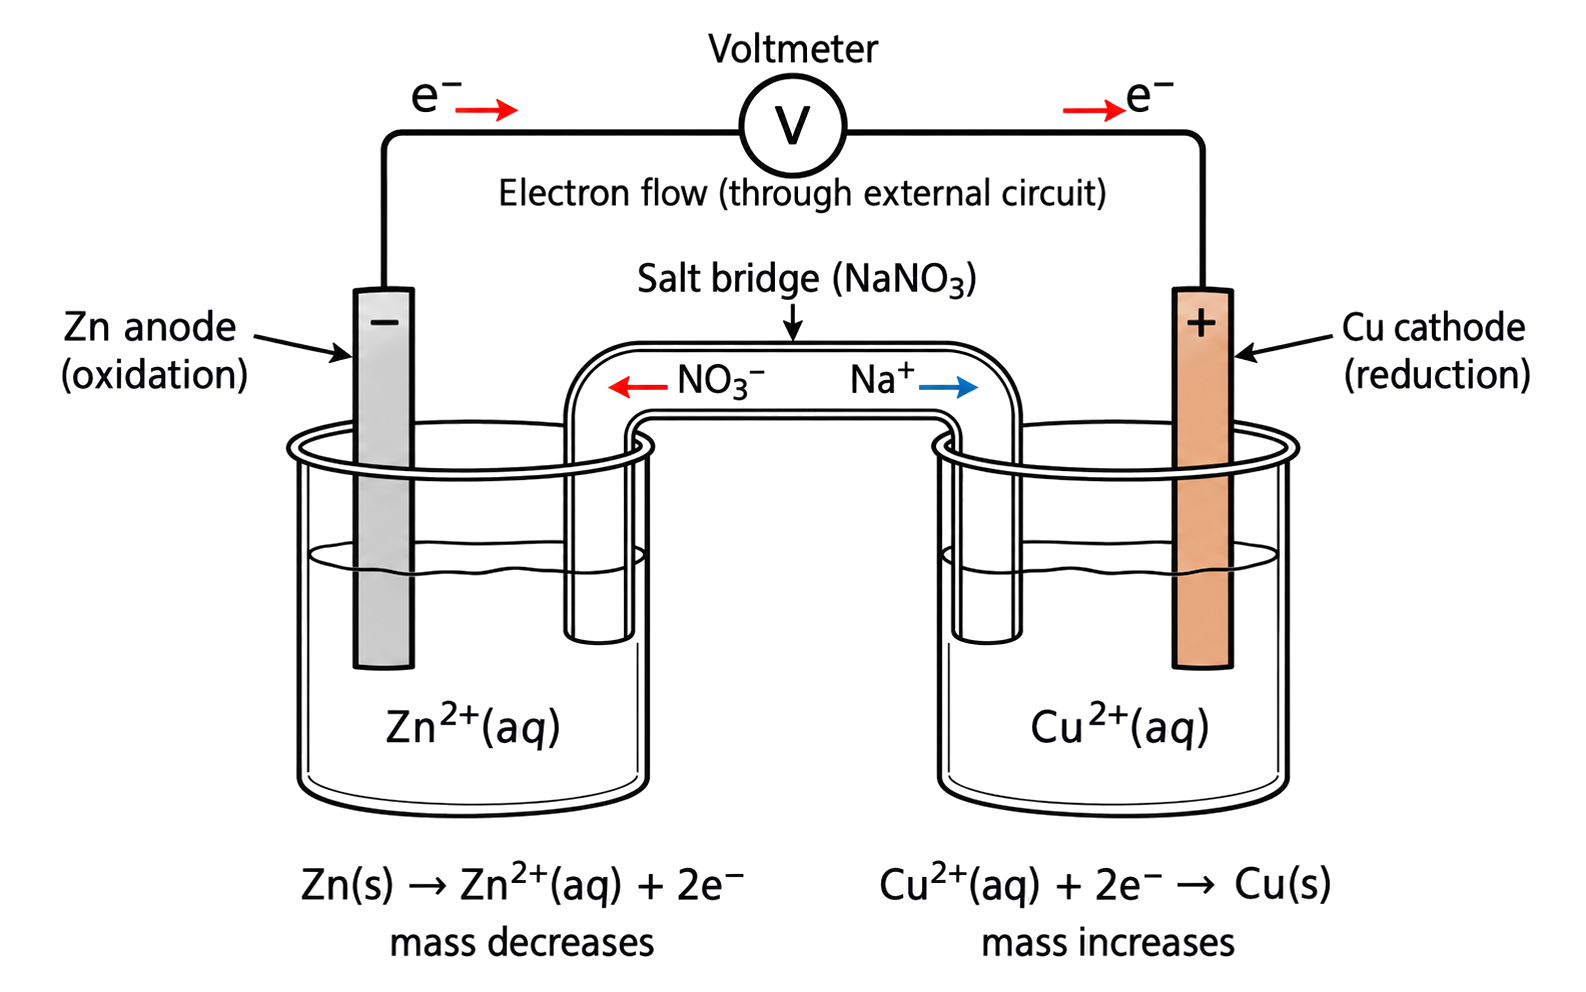
Zn is the negative anode and is oxidized by Zn(s) → Zn2+(aq) + 2e−; Cu is the positive cathode and Cu2+ is reduced by Cu2+(aq) + 2e− → Cu(s); external electrons flow from Zn through the voltmeter to Cu and never through the salt bridge; NO3− migrates toward the anode and Na+ migrates toward the cathode.

Both half-cells connect to the external circuit and the salt bridge must contact both solutions.

In a Zn|Zn²⁺ || Cu²⁺|Cu cell, Zn is oxidized at the negative anode and electrons travel through the wire to the positive Cu cathode, where Cu²⁺ is reduced. Salt-bridge anions migrate toward the anode compartment and cations toward the cathode compartment to prevent charge buildup; electrons do not flow through the bridge.

Completing a Voltaic-Cell Diagram

[ 3 ]

Simple cells rely on differences in standard electrode potential values between different elements and their ions. The following is an incomplete diagram for measuring a cell potential between Mn2+(aq)/Mn\mathrm{Mn}^{2+}(\mathrm{aq}) / \mathrm{Mn} and Ni2+(aq)/Ni\mathrm{Ni}^{2+}(\mathrm{aq}) / \mathrm{Ni} half-cells.

Draw the missing components and fully label the diagram to show how the cell potential can be measured.

Primary, Secondary and Fuel Cells

Cell Energy direction Reuse
primary chemical → electrical not readily reversible
secondary chemical ⇌ electrical recharge by external power
fuel chemical → electrical while reactants are supplied refill fuel

Write the discharge half-equations first. Charging a secondary cell requires an external potential to drive their reverse, whereas a primary cell is not designed for safe efficient reversal and a fuel cell continues only while reactants are supplied. Rechargeability is a reaction-design property, not simply the presence of a power socket.

Explaining Rechargeability

[ 1 ]

Outline how a rechargeable battery differs from a primary cell.

Molten-Salt Electrolysis

In molten salt there is no water: metal ions are reduced to metal at the cathode and anions are oxidized at the anode. For molten chloride, chloride forms chlorine gas.
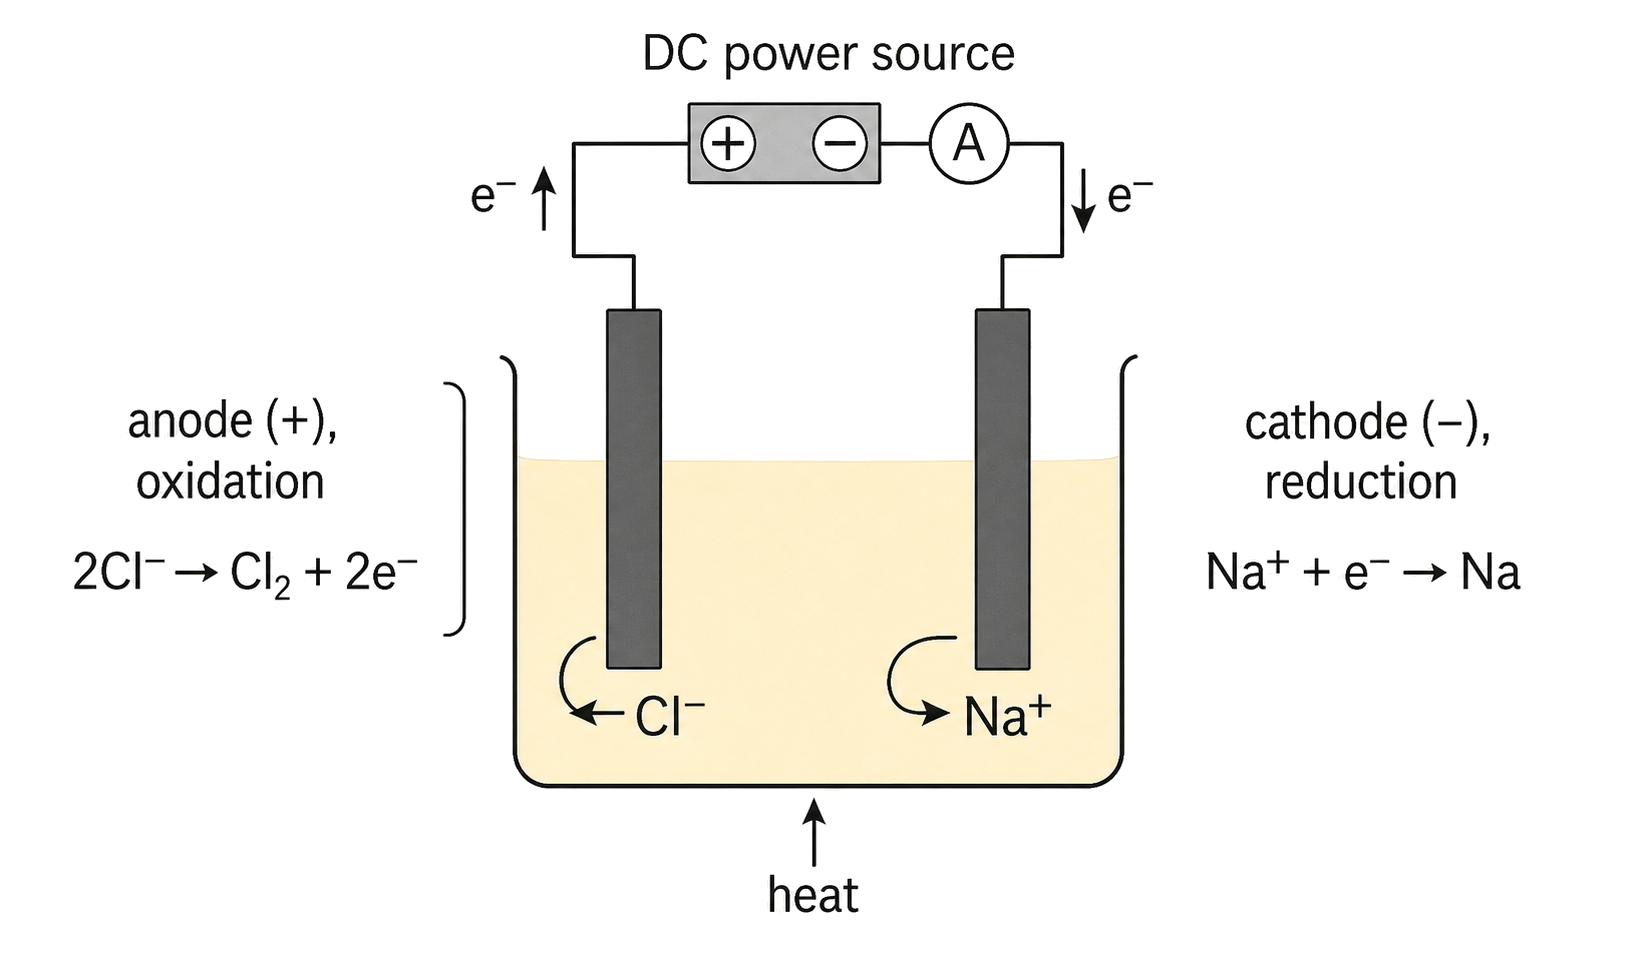
the electrolyte is heated molten NaCl and contains no water; Cl- moves to the positive anode and Na+ moves to the negative cathode; the anode half-equation is 2Cl- to Cl2 plus 2e-; the cathode half-equation is Na+ plus e- to Na.

M(n+)+ne−→Matcathode;2X−→X2+2e−atanodeM^(n+) + ne− → M at cathode; 2X− → X2 + 2e− at anode

Molten MgCl₂ contains only Mg²⁺ and Cl⁻: Mg²⁺ + 2e⁻ → Mg at the cathode and 2Cl⁻ → Cl₂ + 2e⁻ at the anode. The melt conducts by ion migration; do not introduce H₂, O₂ or water-based competition into a molten-salt question.

Deducing Molten-Electrolysis Products

[ 2 ]

Deduce the products of the electrolysis of molten cobalt(II) bromide, CoBr2(l)\mathrm{CoBr}_{2}(\mathrm{l}).

Product at anode:
Product at cathode:

Oxidation of Alcohols

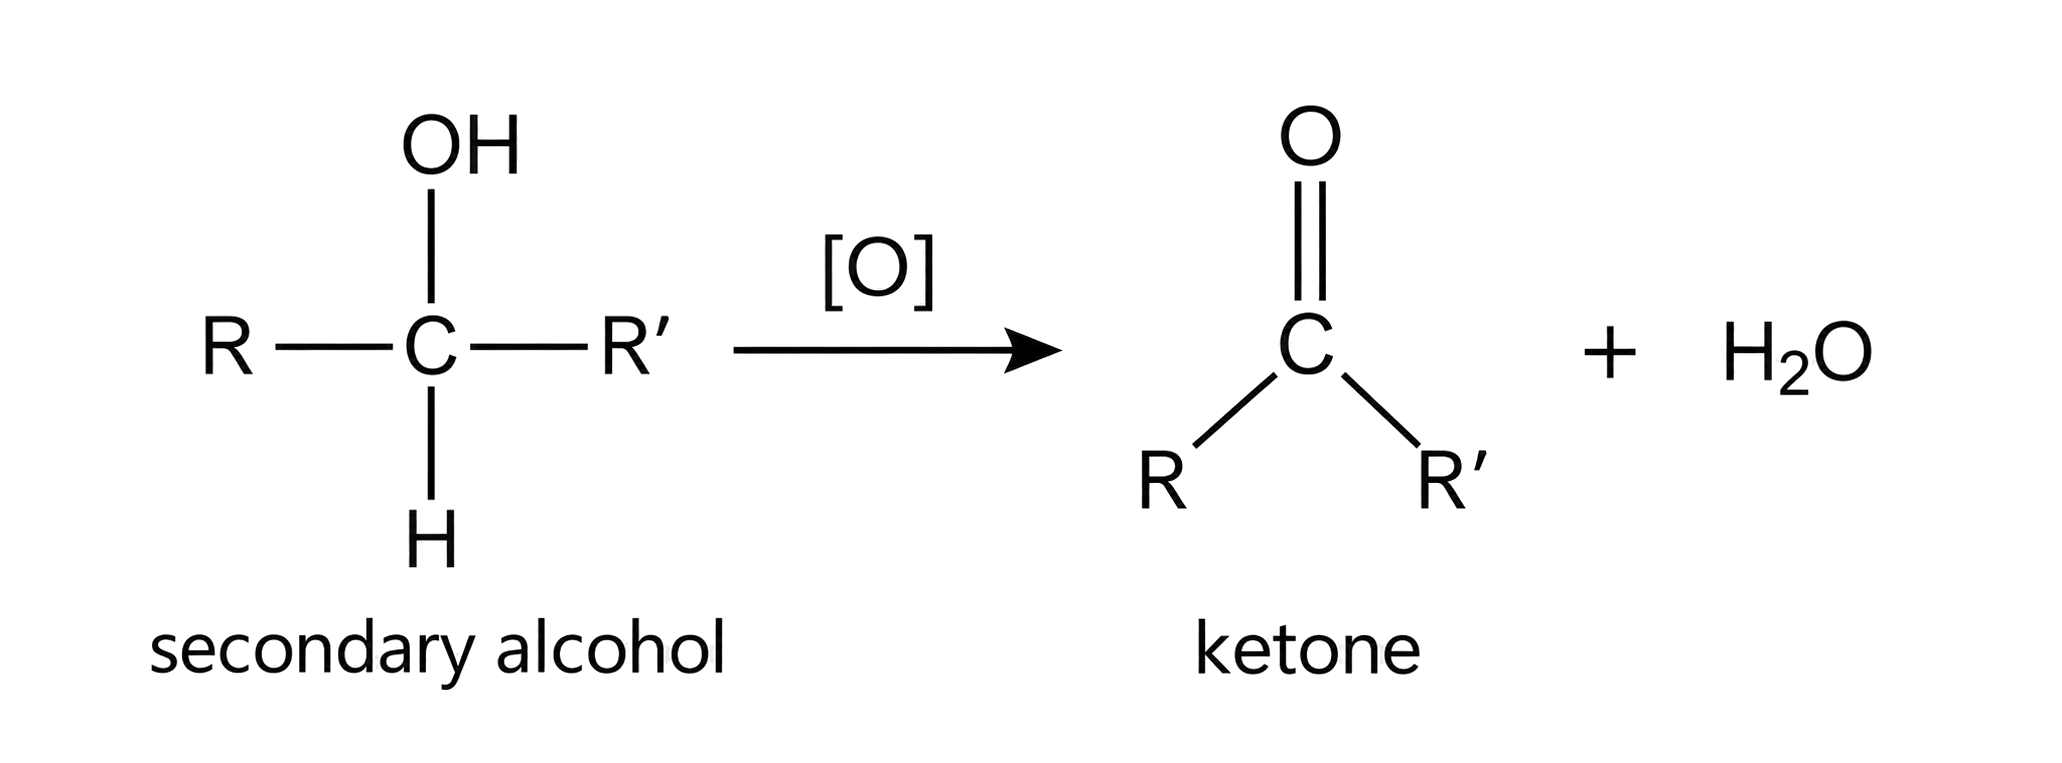
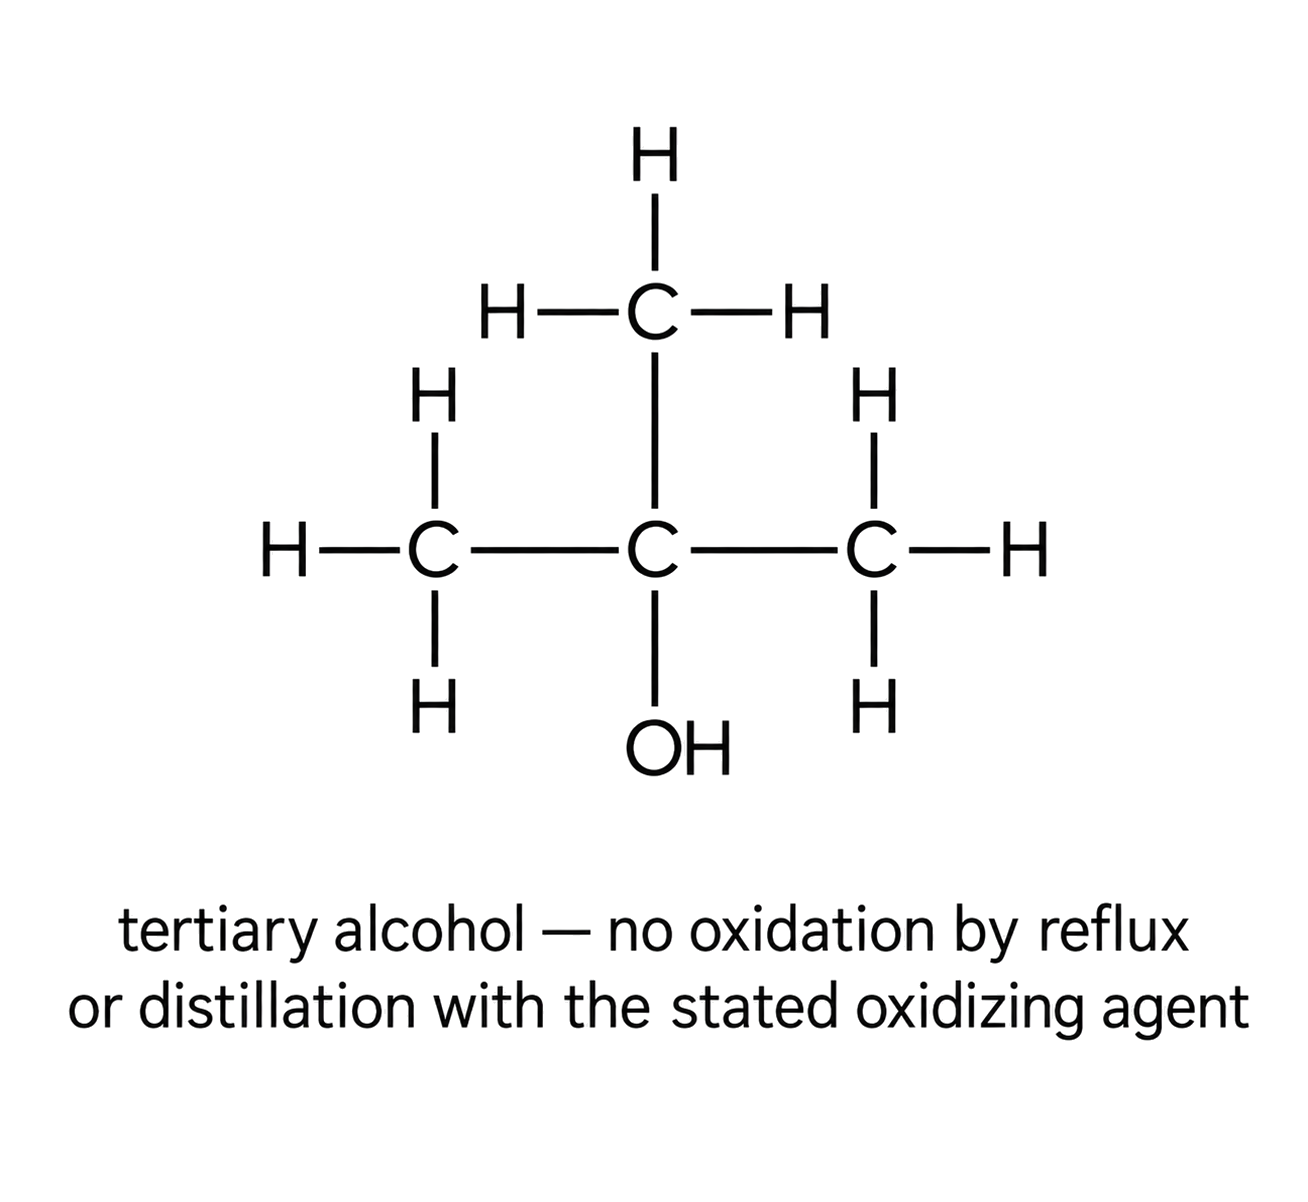
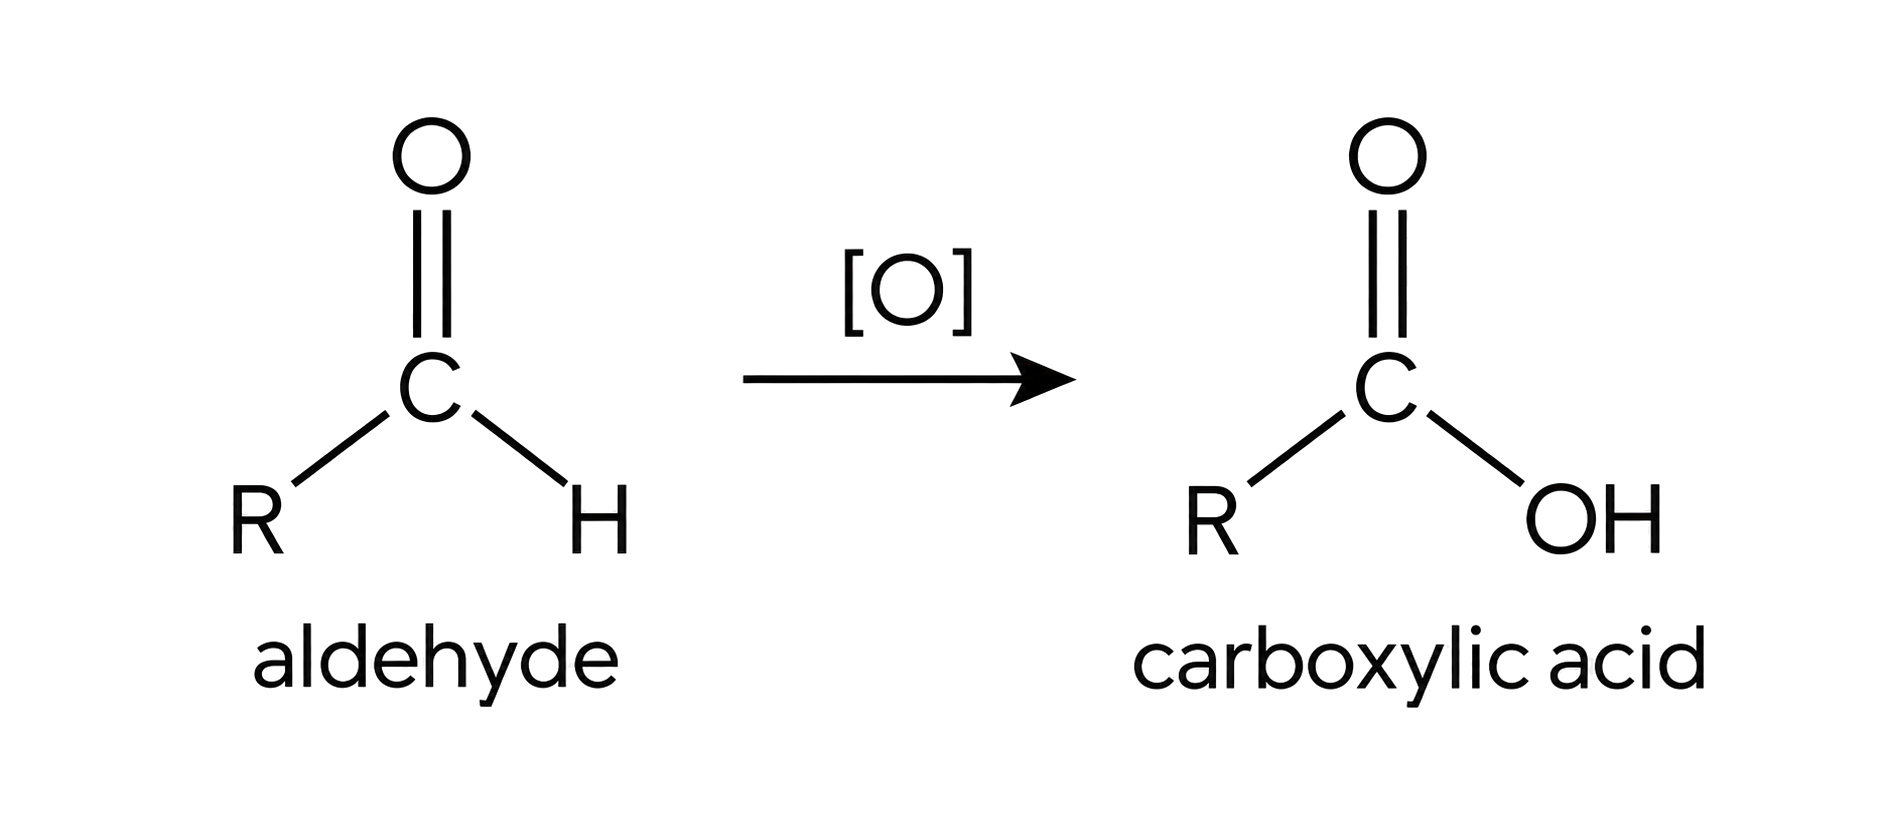
A primary alcohol oxidizes to an aldehyde and then a carboxylic acid; a secondary alcohol oxidizes to a ketone. Reflux supports further oxidation to the acid, while distillation can remove an aldehyde.
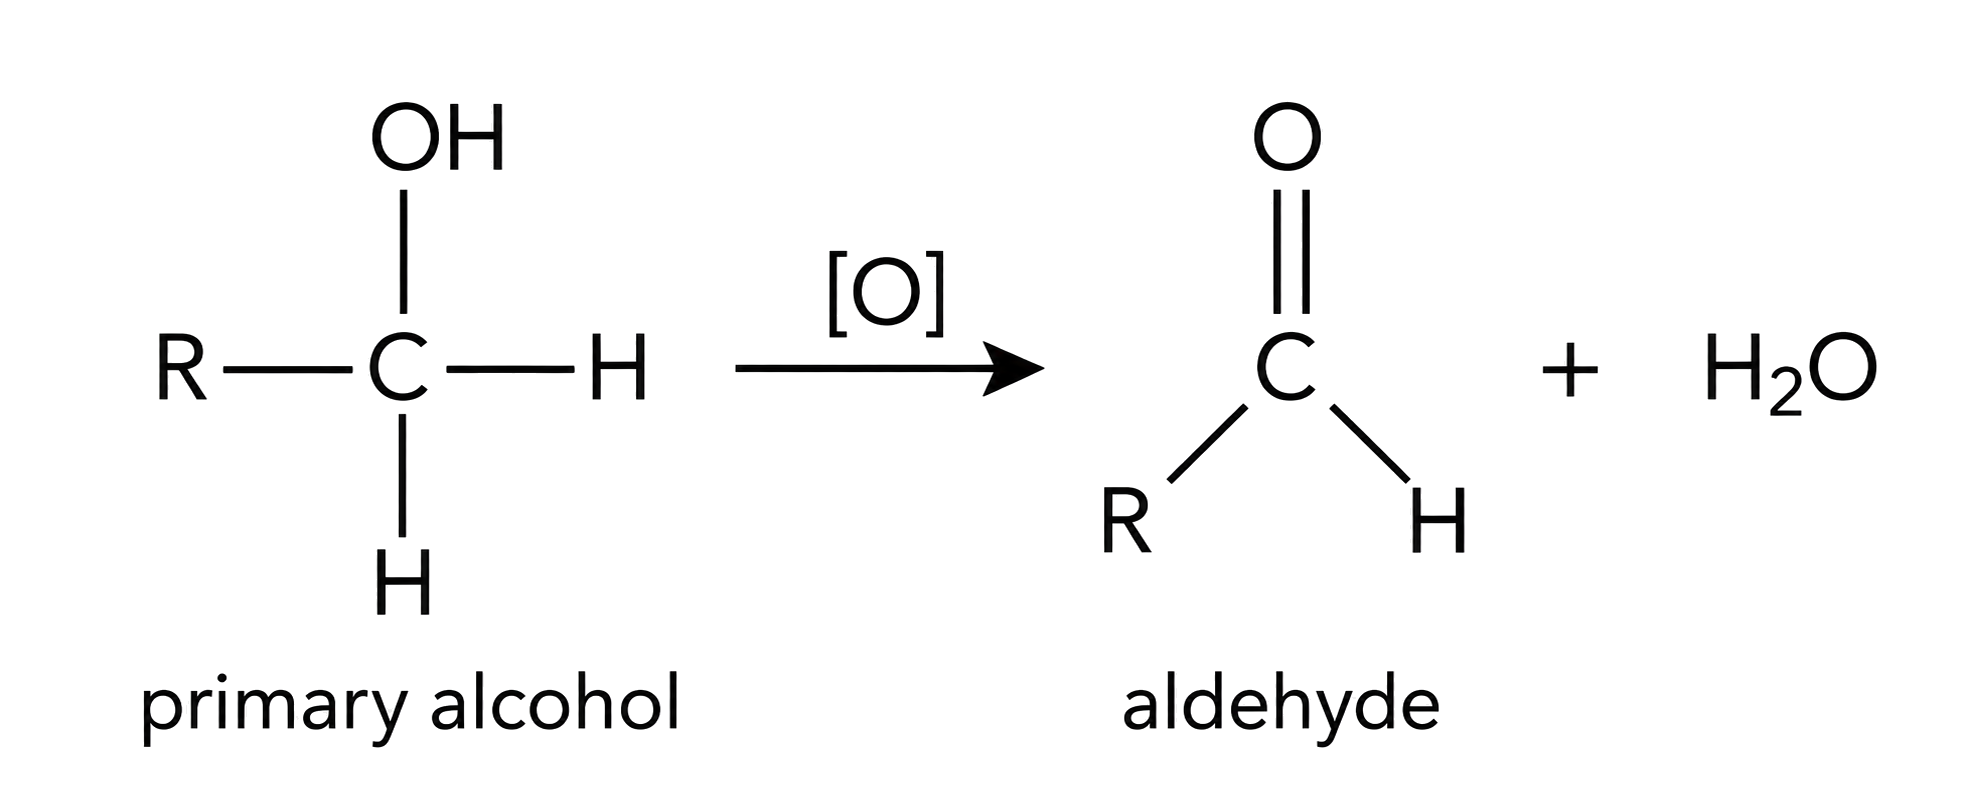
R–CH2OH becomes R–CHO; the aldehyde retains one H on the carbonyl carbon; H2O is shown as in the approved source.
R–CHO becomes R–COOH; the same carbonyl carbon and R group are retained; no unsupported water product is added.
R–CH(OH)–R′ becomes R–C(=O)–R′; both R groups and the functional carbon are retained; H2O is shown as in the approved source.
the structure is (CH3)3C–OH; the carbon bearing OH has no H; the non-reaction claim is explicitly limited to the stated reflux/distillation oxidizing conditions.

In a primary-alcohol experiment, distil the aldehyde as it forms to limit further oxidation; heat under reflux when the carboxylic acid is required. Tertiary alcohols lack the required hydrogen on the carbon bearing –OH and are not oxidized in the same way.

Choosing Alcohol-Oxidation Products

[ 2 ]

Deduce the organic products when butan-1-ol and butan-2-ol are separately heated under reflux with acidified potassium dichromate(VI).

Butan-1-ol:
Butan-2-ol:

Reduction of Carbonyl Compounds

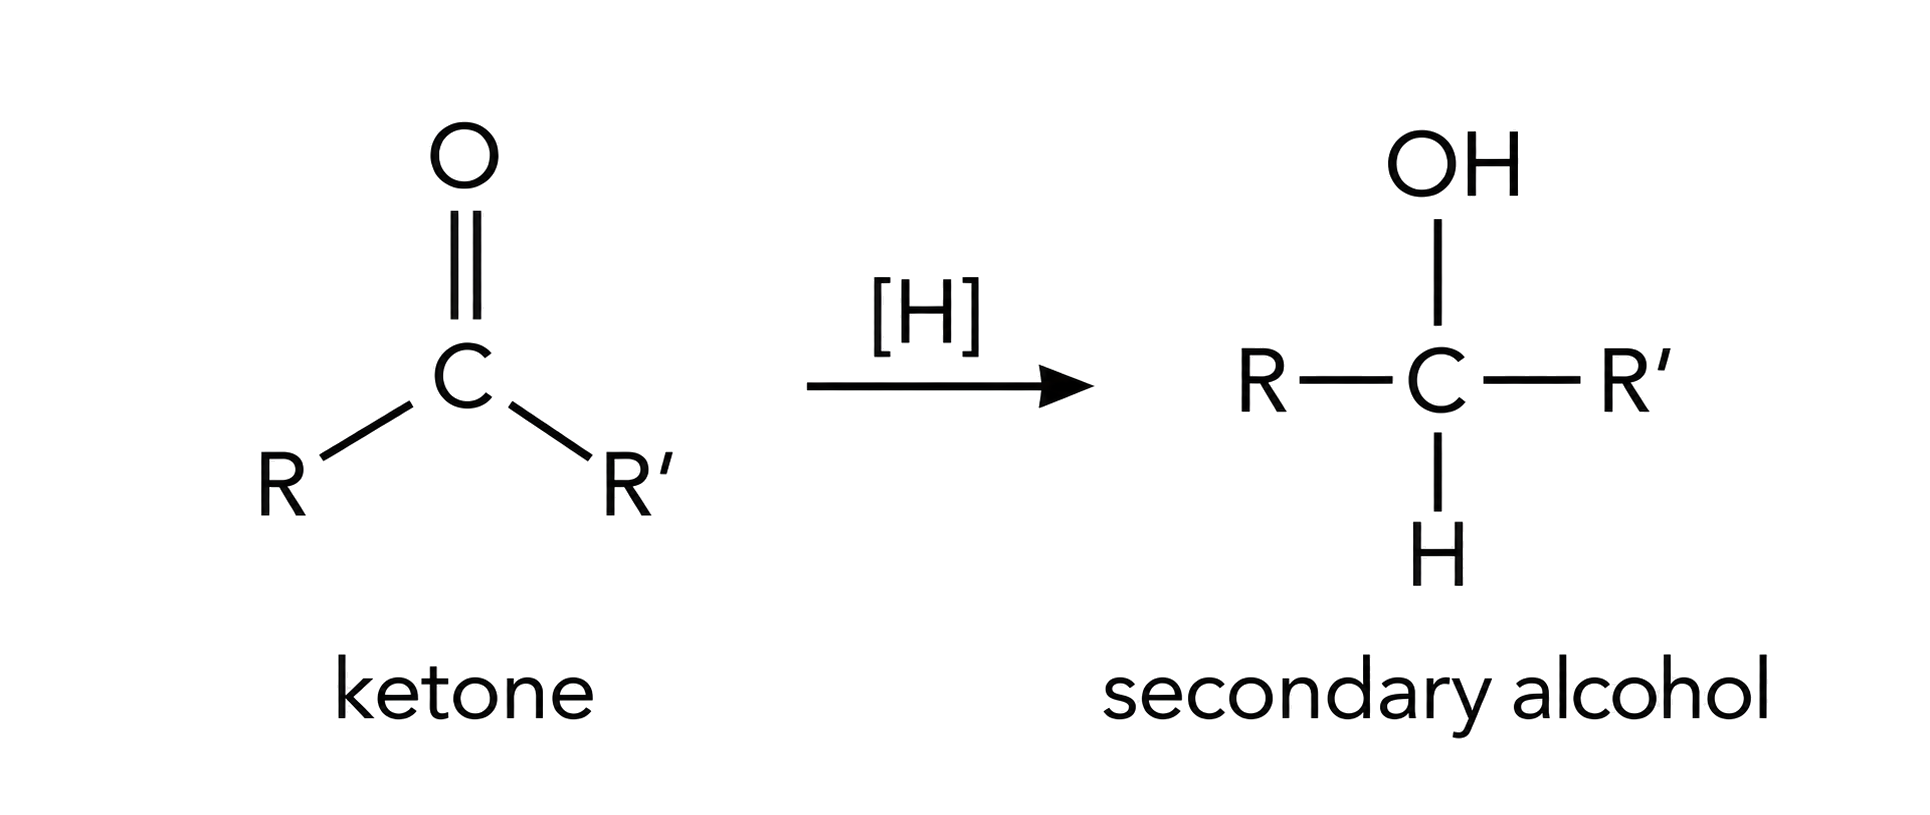
A carboxylic acid can be reduced through an aldehyde to a primary alcohol; a ketone is reduced to a secondary alcohol. Hydride ions supply the reduction equivalent in these transformations.
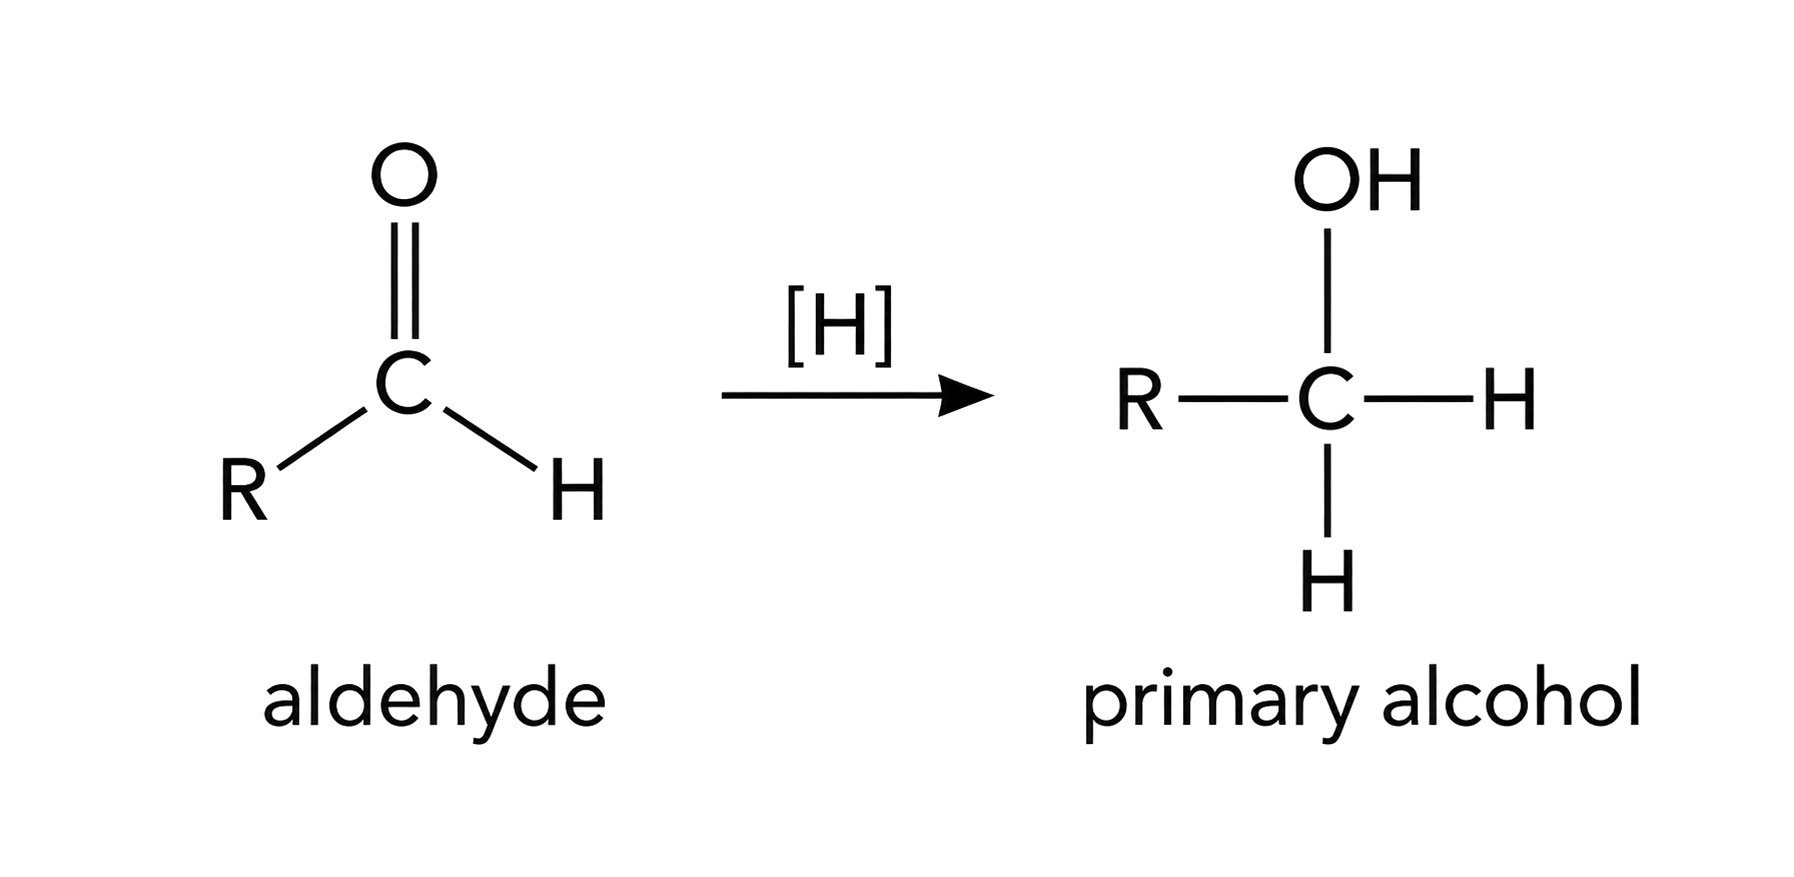
R–CHO becomes R–CH2OH; the product carbon has R, OH and two H substituents; the product is correctly labelled primary alcohol.
R–C(=O)–R′ becomes R–CH(OH)–R′; the product is correctly labelled secondary alcohol; the carbon skeleton is unchanged.

Track the carbon functional group rather than only the reagent: an aldehyde gives a primary alcohol and a ketone gives a secondary alcohol. Hydride supplies an electron-rich H unit to the carbonyl carbon; named reducing agents and detailed mechanisms are outside this objective.

Deducing Reduction Products

[ 1 ]

Which product may be obtained by the reduction of CH3CH2COOH\mathrm{CH}_{3} \mathrm{CH}_{2} \mathrm{COOH} ?

Hydrogenation of Alkenes and Alkynes

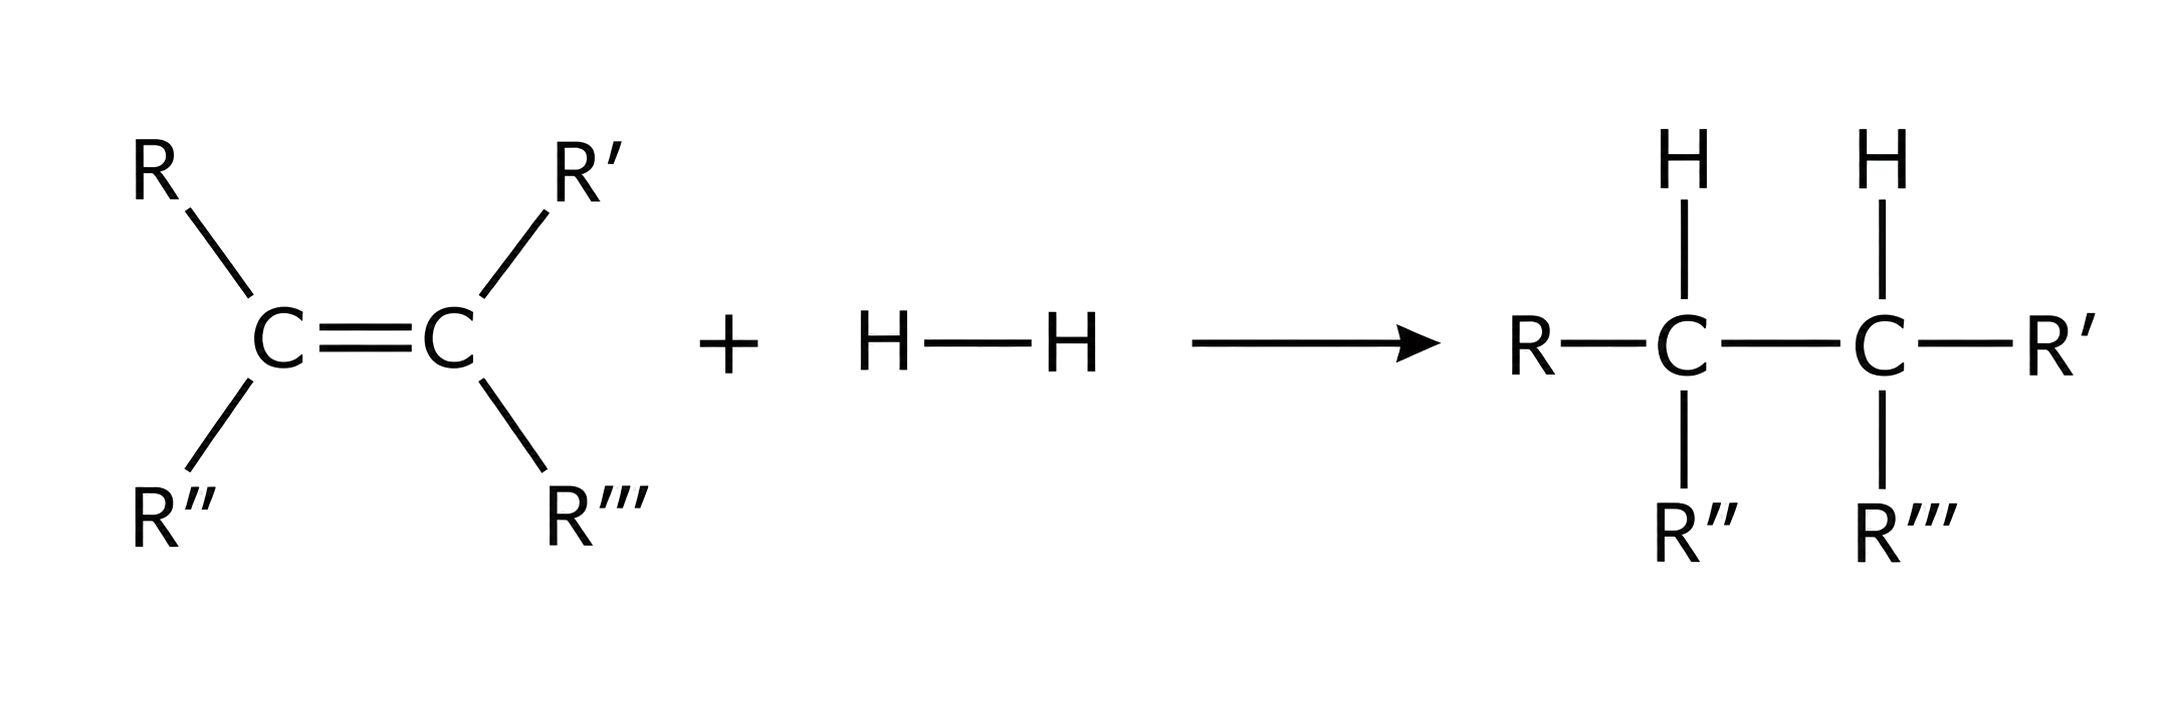
Hydrogenation adds H2 across π bonds. Continue addition until the required saturated product is formed; nickel, palladium or platinum catalysts with heat or pressure are typical conditions.
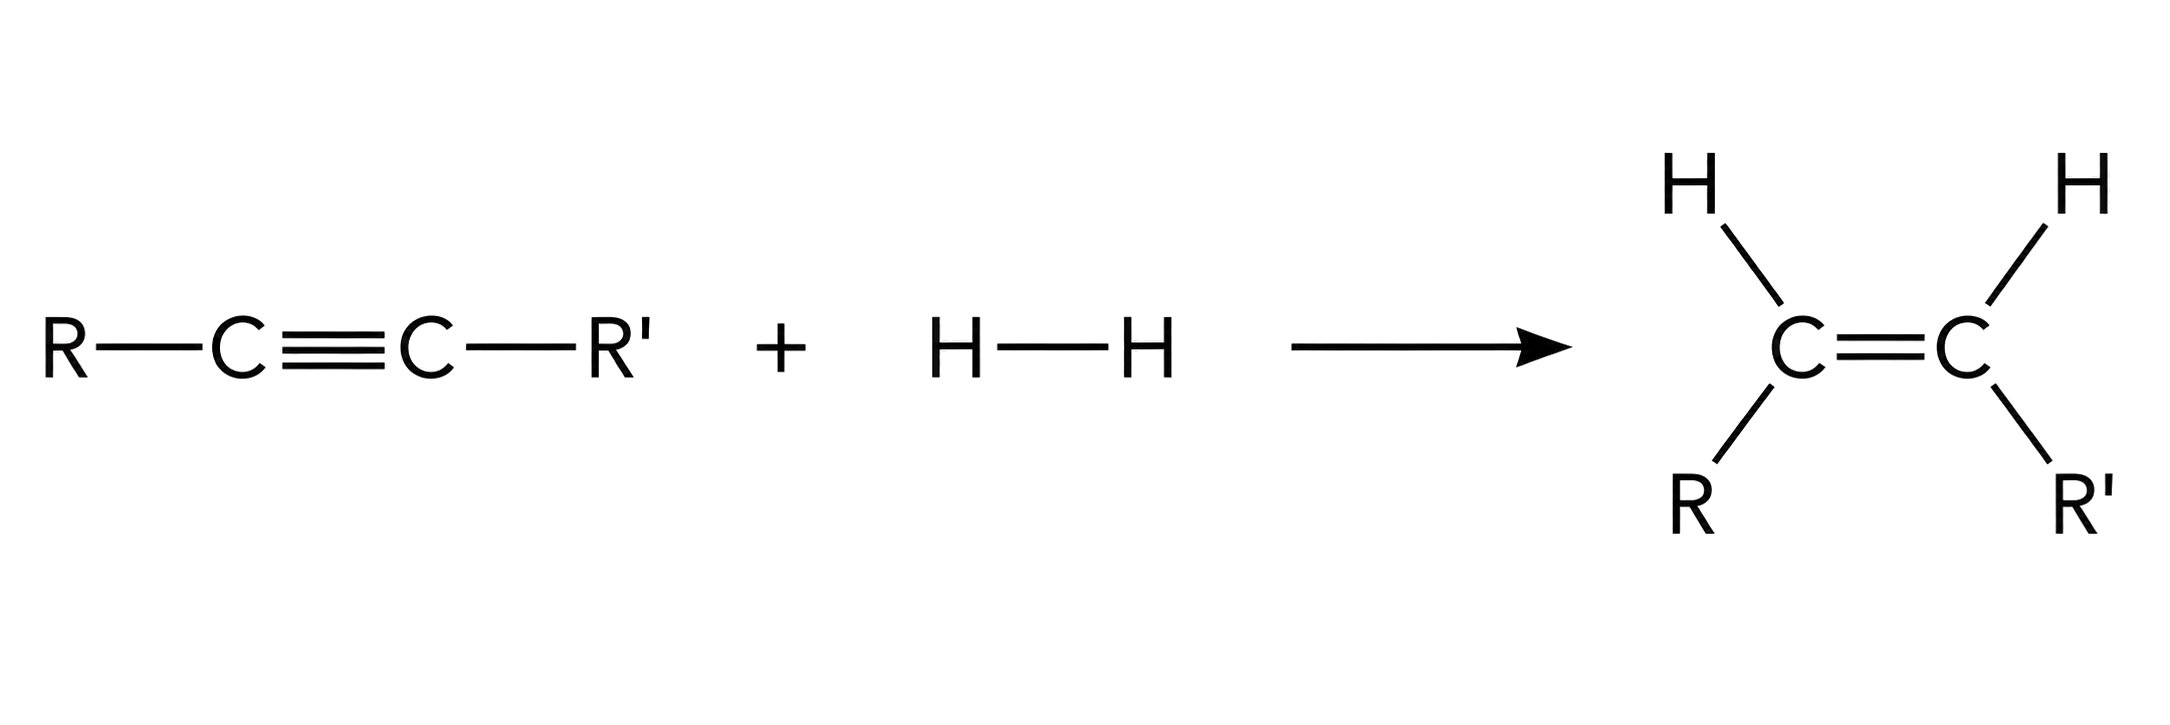
R–C≡C–R′ receives exactly one H2; one H is added to each alkyne carbon and C≡C becomes C=C; R and R′ remain on their original carbons; no unsupported stereochemical outcome is claimed.
one H2 is added across C=C; C=C becomes C–C while the sigma carbon skeleton remains intact; R, R′, R″ and R‴ remain on the correct original carbon; each alkene carbon gains exactly one H.
R–C≡C–R′ receives exactly 2 H2; the product is R–CH2–CH2–R′; the triple bond becomes a single bond without breaking the carbon skeleton.

Count π bonds to determine hydrogen demand: one mole of H₂ saturates one C=C, while full conversion of one C≡C to C–C needs two moles of H₂. Keep the carbon skeleton unchanged when drawing the product.
Deducing Hydrogenation Products

[ 2 ]

State the reagent and conditions needed and draw the structural formula of the product.

The Standard Hydrogen Electrode

HL only

E°(SHE)=0VbyconventionE°(SHE) = 0 V by convention

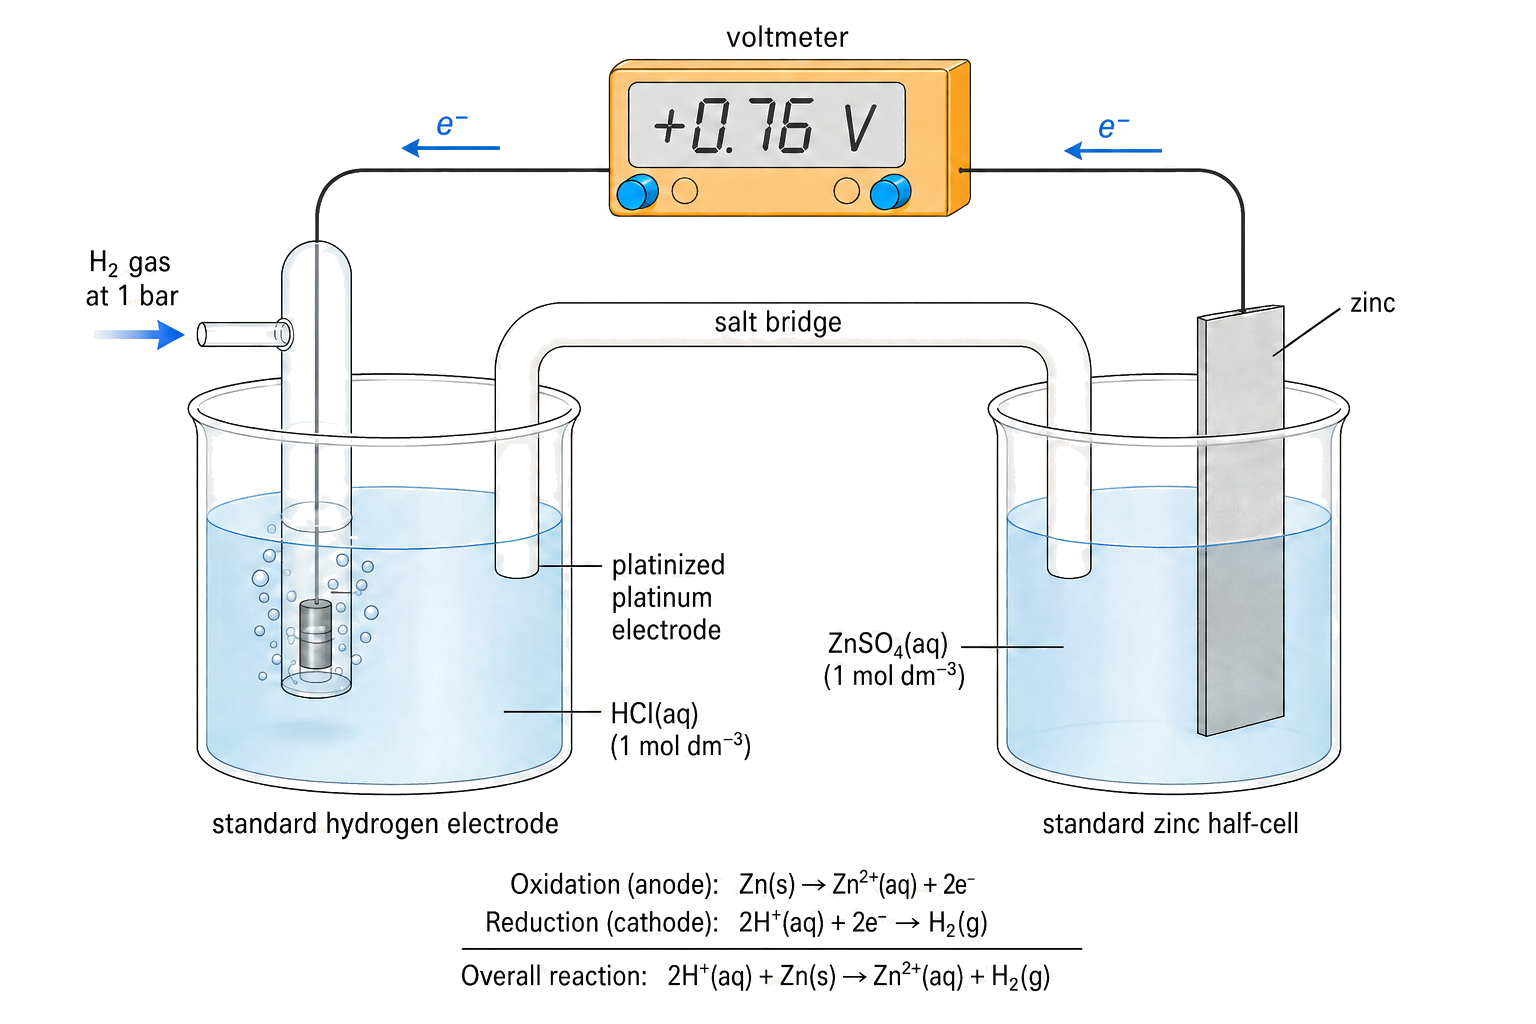
the SHE uses platinized platinum, H2 at 1 bar, and HCl at 1 mol dm-3; the zinc half-cell uses zinc in ZnSO4 at 1 mol dm-3; electrons flow externally from zinc to the SHE and the voltmeter reads +0.76 V; zinc oxidation, H+ reduction, and the overall reaction are balanced and correctly assigned.

Use reduction-form data: a more positive E° means greater tendency to be reduced and stronger oxidizing behaviour. A very negative metal reduction potential indicates ease of reverse oxidation and strong reducing behaviour.

The standard hydrogen electrode uses H₂(g) at 100 kPa in contact with aqueous H⁺ of unit activity (commonly represented as 1 mol dm⁻³) at 298 K on an inert platinum surface, and is assigned E° = 0.00 V. Pair an unknown half-cell with this reference, use polarity to identify reduction, then interpret more positive reduction potential as stronger oxidizing tendency.

Interpreting Standard Potentials

HL only

[ 2 ]

Comment on the sign and value of the standard reduction potential of lithium that make it suitable to use in the battery. Use section 19 of the data booklet.

Standard Cell Potential

HL only

E°cell=E°cathode−E°anode(usingtabulatedreductionpotentials)E°cell = E°cathode − E°anode (using tabulated reduction potentials)

the SHE uses platinized platinum, H2 at 1 bar, and HCl at 1 mol dm-3; the zinc half-cell uses zinc in ZnSO4 at 1 mol dm-3; electrons flow externally from zinc to the SHE and the voltmeter reads +0.76 V; zinc oxidation, H+ reduction, and the overall reaction are balanced and correctly assigned.

A positive E°cell indicates a spontaneous voltaic direction. Reverse the direction if the calculated sign is negative.

Select the more positive reduction potential as the cathode reaction, keep both tabulated values as reduction potentials, and calculate E°cell = E°cathode − E°anode. Do not multiply an electrode potential when a half-equation is scaled.

Worked Ecell∘E^\circ_{cell} example: E∘(AgX+/Ag)=+0.80 VE^\circ(\ce{Ag+/Ag})=+0.80\,\mathrm{V} and E∘(CuX2+/Cu)=+0.34 VE^\circ(\ce{Cu^{2+}/Cu})=+0.34\,\mathrm{V}. Silver is the cathode, so Ecell∘=Ecathode∘−Eanode∘=0.80−0.34=+0.46 VE^\circ_{cell}=E^\circ_{cathode}-E^\circ_{anode}=0.80-0.34=+0.46\,\mathrm{V}. The positive result predicts the spontaneous reaction 2 AgX++Cu→2 Ag+CuX2+\ce{2Ag+ + Cu -> 2Ag + Cu^{2+}} under standard conditions. Do not multiply E∘E^\circ when doubling the silver half-equation.

Calculating E°cell

HL only

[ 1 ]

Calculate the standard cell potential, Ecell 0E_{\text {cell }}^{0}, for this cell. Use section 19 of the data booklet.

Gibbs Energy and Cell Potential

HL only

ΔG°=−nFE°cellΔG° = −nFE°cell

n is the moles of electrons transferred and F is Faraday's constant. Positive E°cell gives negative ΔG° and a spontaneous reaction.

Find n from the balanced overall redox equation, not from a single unscaled half-equation. With E° in volts and F in C mol⁻¹, ΔG° is obtained in J mol⁻¹; convert to kJ mol⁻¹ only at the end.

Worked ΔG∘\Delta G^\circ example: for 2 HX++Zn→ZnX2++HX2\ce{2H+ + Zn -> Zn^{2+} + H2}, n=2n=2 and Ecell∘=+0.76 VE^\circ_{cell}=+0.76\,\mathrm{V}. Using F=9.65×10⁴ C mol−1F=9.65\times10^4\,\mathrm{C\,mol^{-1}}, ΔG∘=−(2)(9.65×10⁴)(0.76)=−1.47×10⁵ J mol−1=−147 kJ mol−1\Delta G^\circ=-(2)(9.65\times10^4)(0.76)=-1.47\times10^5\,\mathrm{J\,mol^{-1}}=-147\,\mathrm{kJ\,mol^{-1}}. Its negative sign agrees with a spontaneous standard-cell reaction.

Calculating ΔG° for a Cell

HL only

[ 2 ]

Calculate the standard Gibbs free energy of the cell, in kJmol−1\mathrm{kJ} \mathrm{mol}^{-1}. Use sections 1, 2 and 24 of the data booklet.

Aqueous Electrolysis

HL only

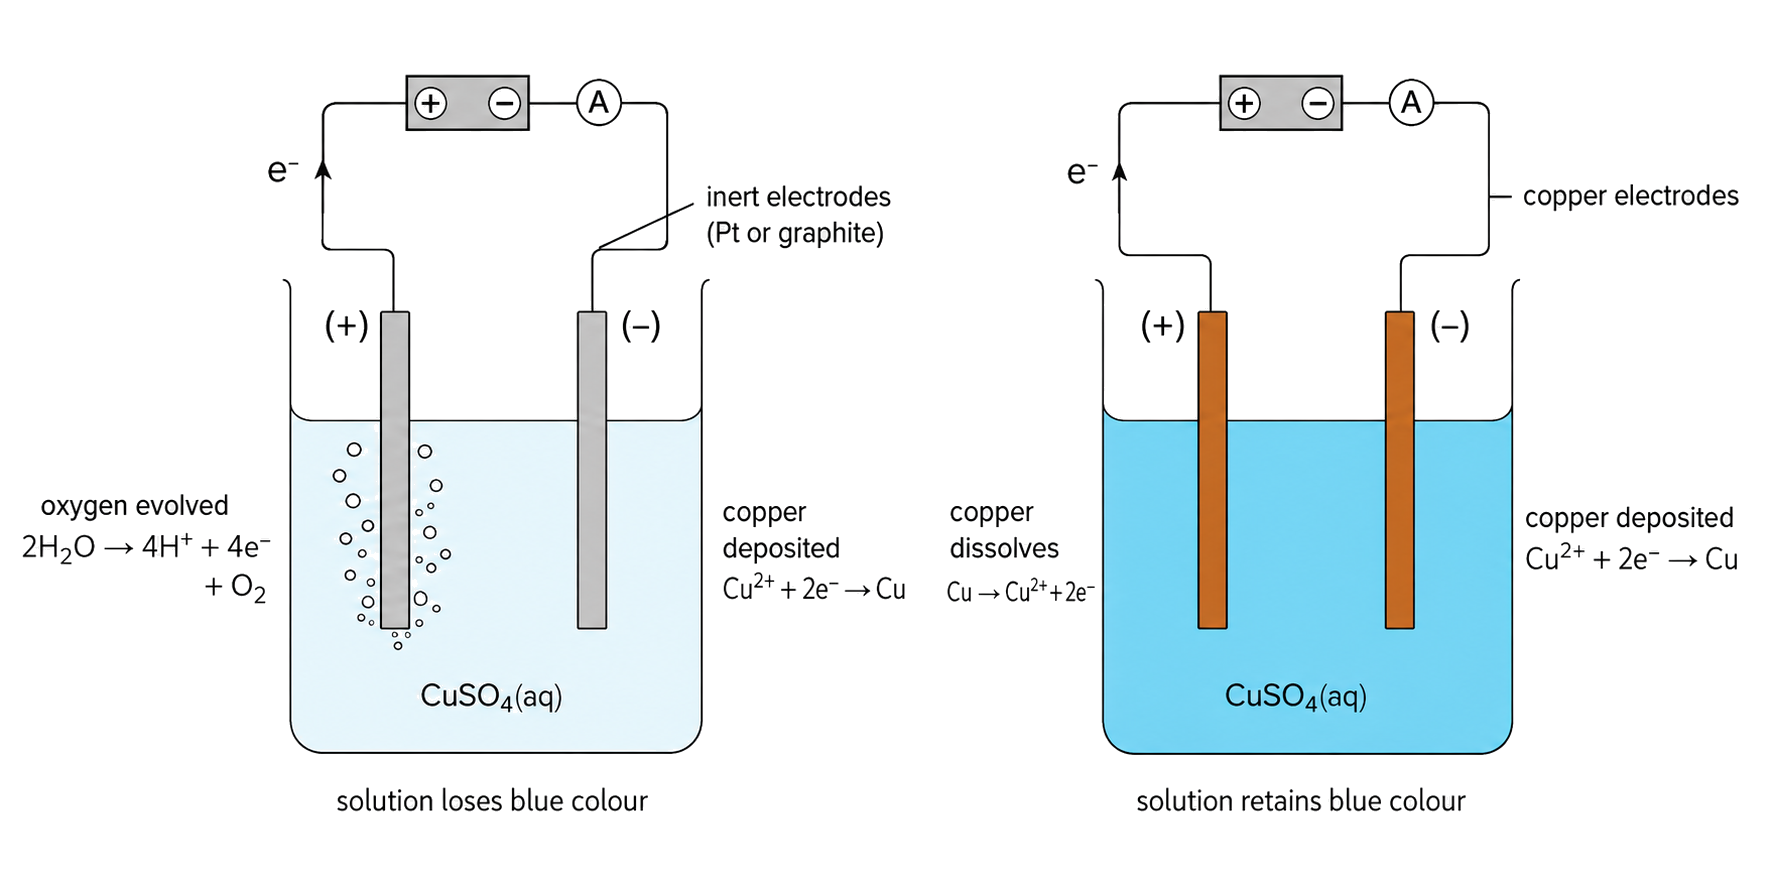
At each electrode compare the possible aqueous species using reduction or oxidation tendencies. Water may react instead of sulfate or in dilute halide solution; concentrated halide can be oxidized, while molten salt contains no water.
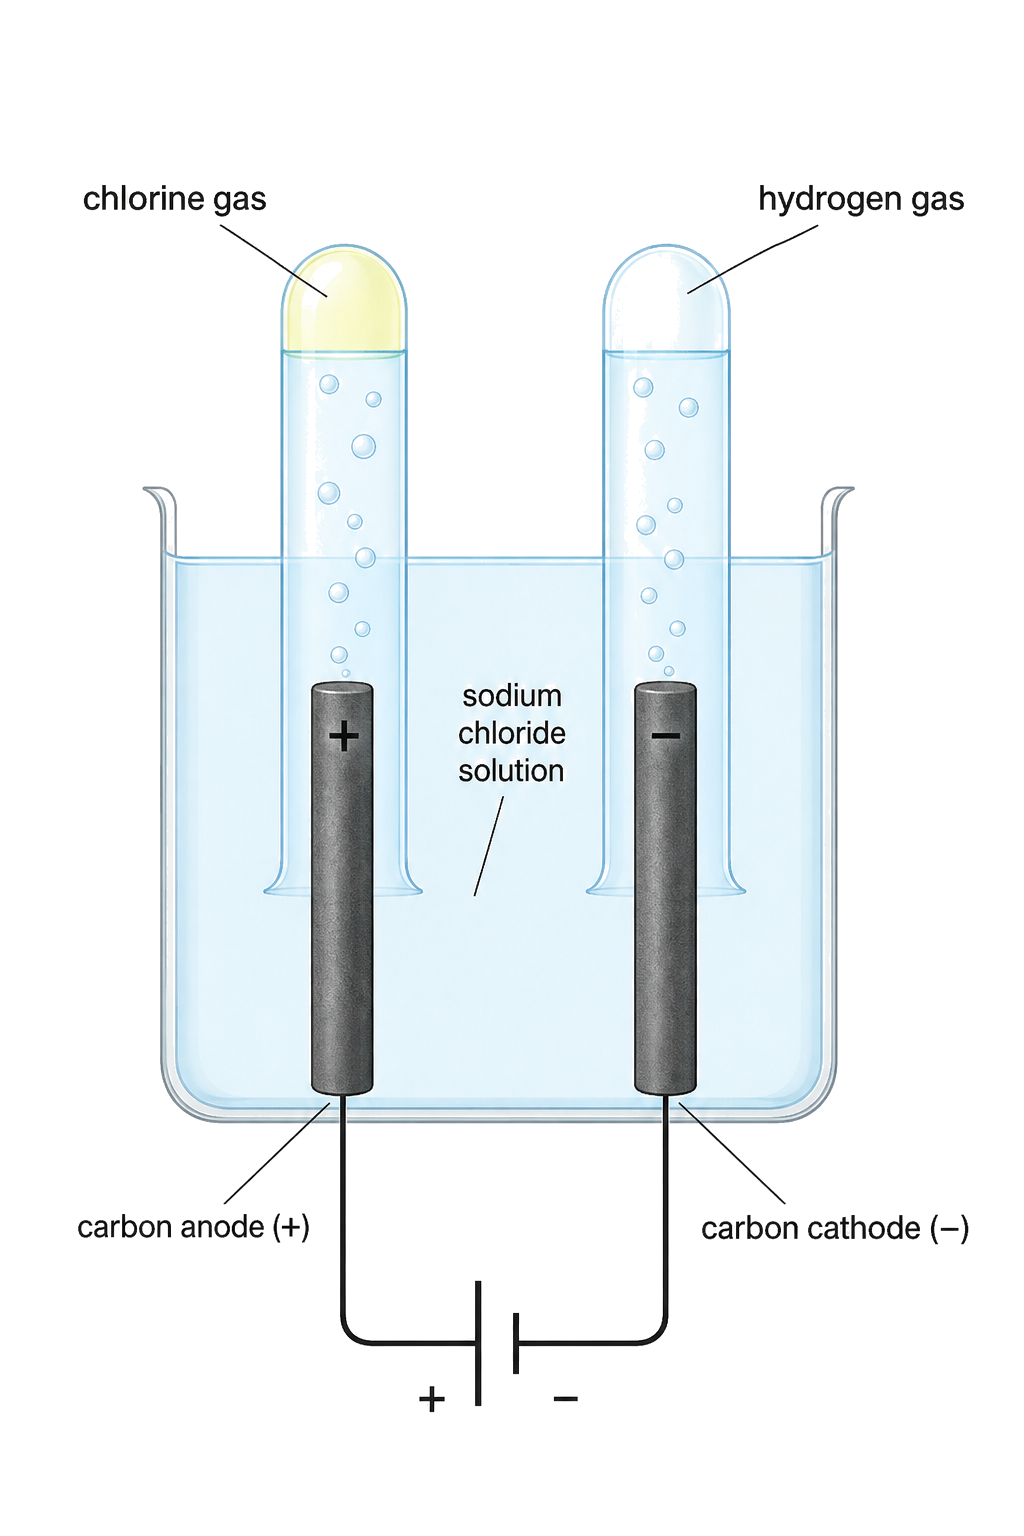
aqueous sodium chloride and inert carbon electrodes are explicit; chlorine is collected at the positive anode; hydrogen is collected at the negative cathode; sodium metal is not shown as an aqueous cathode product.
with inert electrodes oxygen evolves at the positive anode and copper deposits at the negative cathode; the inert-electrode solution loses blue colour as Cu2+ is removed; with copper electrodes the positive copper anode dissolves and copper deposits at the negative cathode; the copper-electrode solution retains blue colour because Cu2+ is replenished.

List the solute ion and water as competing possibilities at each electrode, then use electrode-potential data together with stated concentration conditions to select products. In aqueous sulfate, water commonly supplies the anode gas; in concentrated halide, halogen formation may compete. Do not transfer molten-salt products automatically to solution.

Selecting Aqueous-Electrolysis Products

HL only

[ 2 ]

Determine the products formed at each electrode during the electrolysis of an aqueous solution of sodium bromide. Use section 24 in the data booklet.

Positive electrode (anode):
Negative electrode (cathode):

Electroplating with Electrolysis

HL only

The object being coated is the cathode, where coating-metal ions are reduced to metal. Use an electrolyte containing those ions; a coating-metal anode can replenish them.
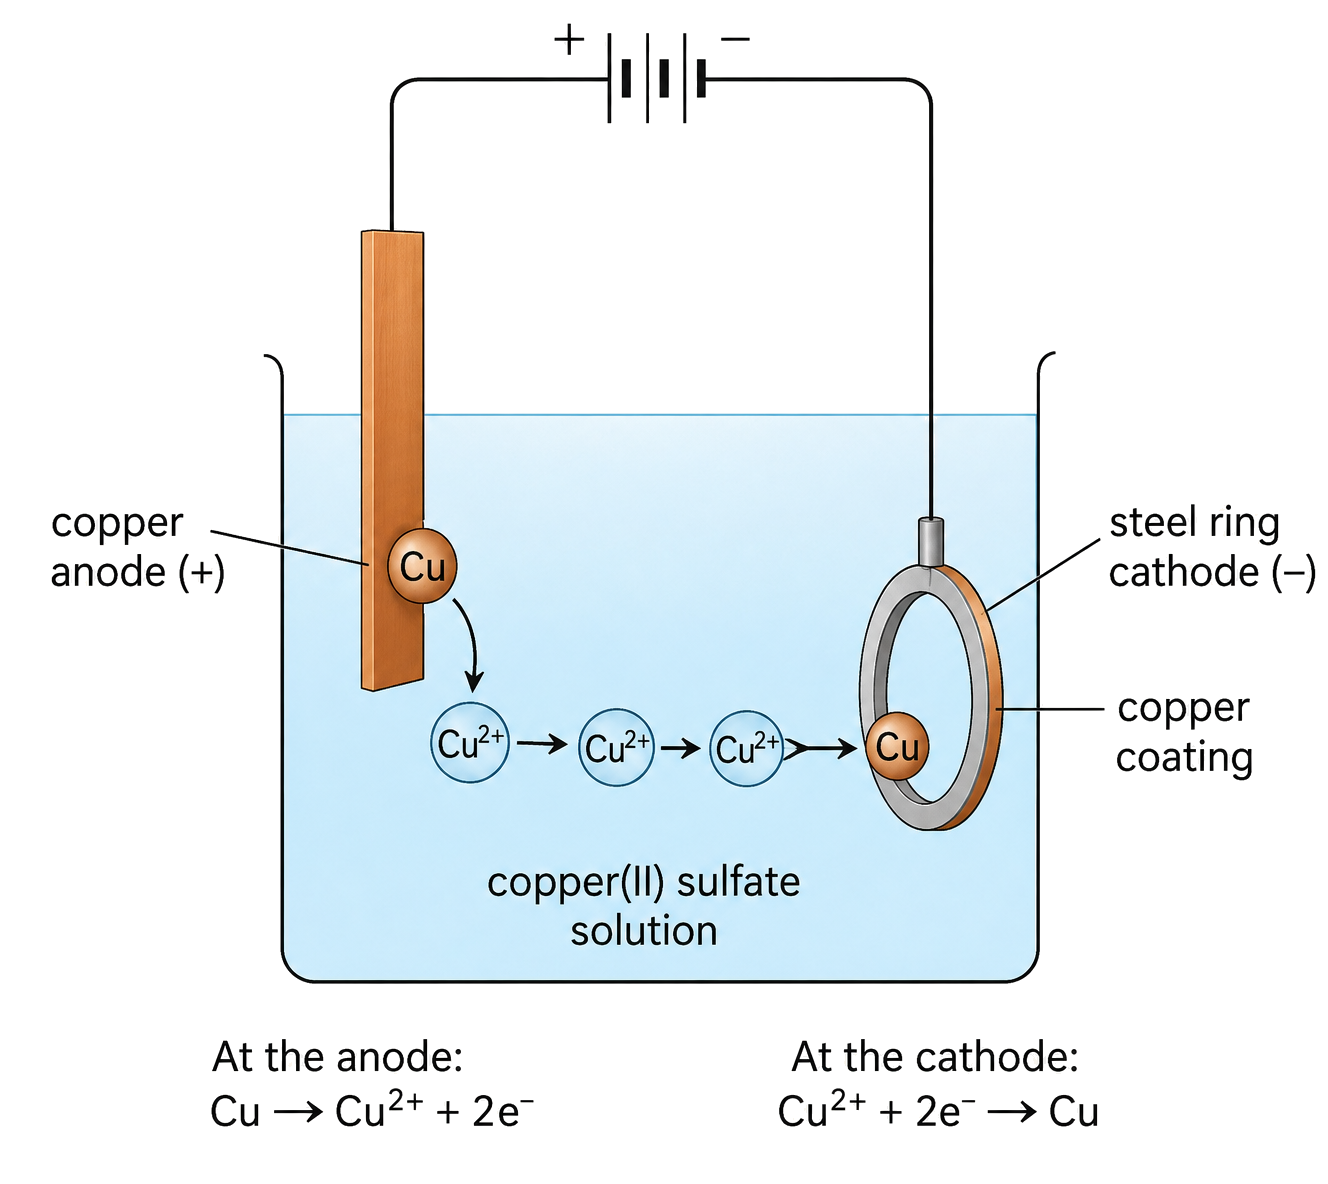
the soluble copper anode is positive and the steel ring cathode is negative; Cu2+ moves through copper(II) sulfate solution toward the ring; copper dissolves at the anode and is reduced as a coating at the cathode; both copper half-equations are correct and electron-balanced.

M(n+)+ne−→M(s)attheobjectcathodeM^(n+) + ne− → M(s) at the object cathode

Trace metal atoms through the circuit: M atoms may oxidize at a soluble anode to maintain Mⁿ⁺, while Mⁿ⁺ gains electrons and deposits on the object. Reversing the object to the anode would remove metal rather than coat it.

Designing an Electroplating Cell

HL only

[ 3 ]

Describe how electrolysis can be used to electroplate a bracelet with a layer of silver metal. Include the choice of electrodes and electrolyte needed in your description.

Electron Transfer Reactions Summary

Retrieve the route: assign oxidation states, balance half-equations, predict displacement, label cells, trace electrons and ions, follow organic redox pathways, calculate potentials and choose electrolysis products.

Check electron loss/gain, anode/cathode versus polarity, spontaneous sign, salt-bridge direction, ions present, organic functional-group direction and object-cathode placement.

3.3 Electron sharing reactions (Radicals)

Syllabus
First assessment 2025
Topic
3.3
Level
HL

Free Radicals

A radical is a highly reactive species containing an unpaired electron. Show the unpaired electron with a dot next to the atom that carries it.
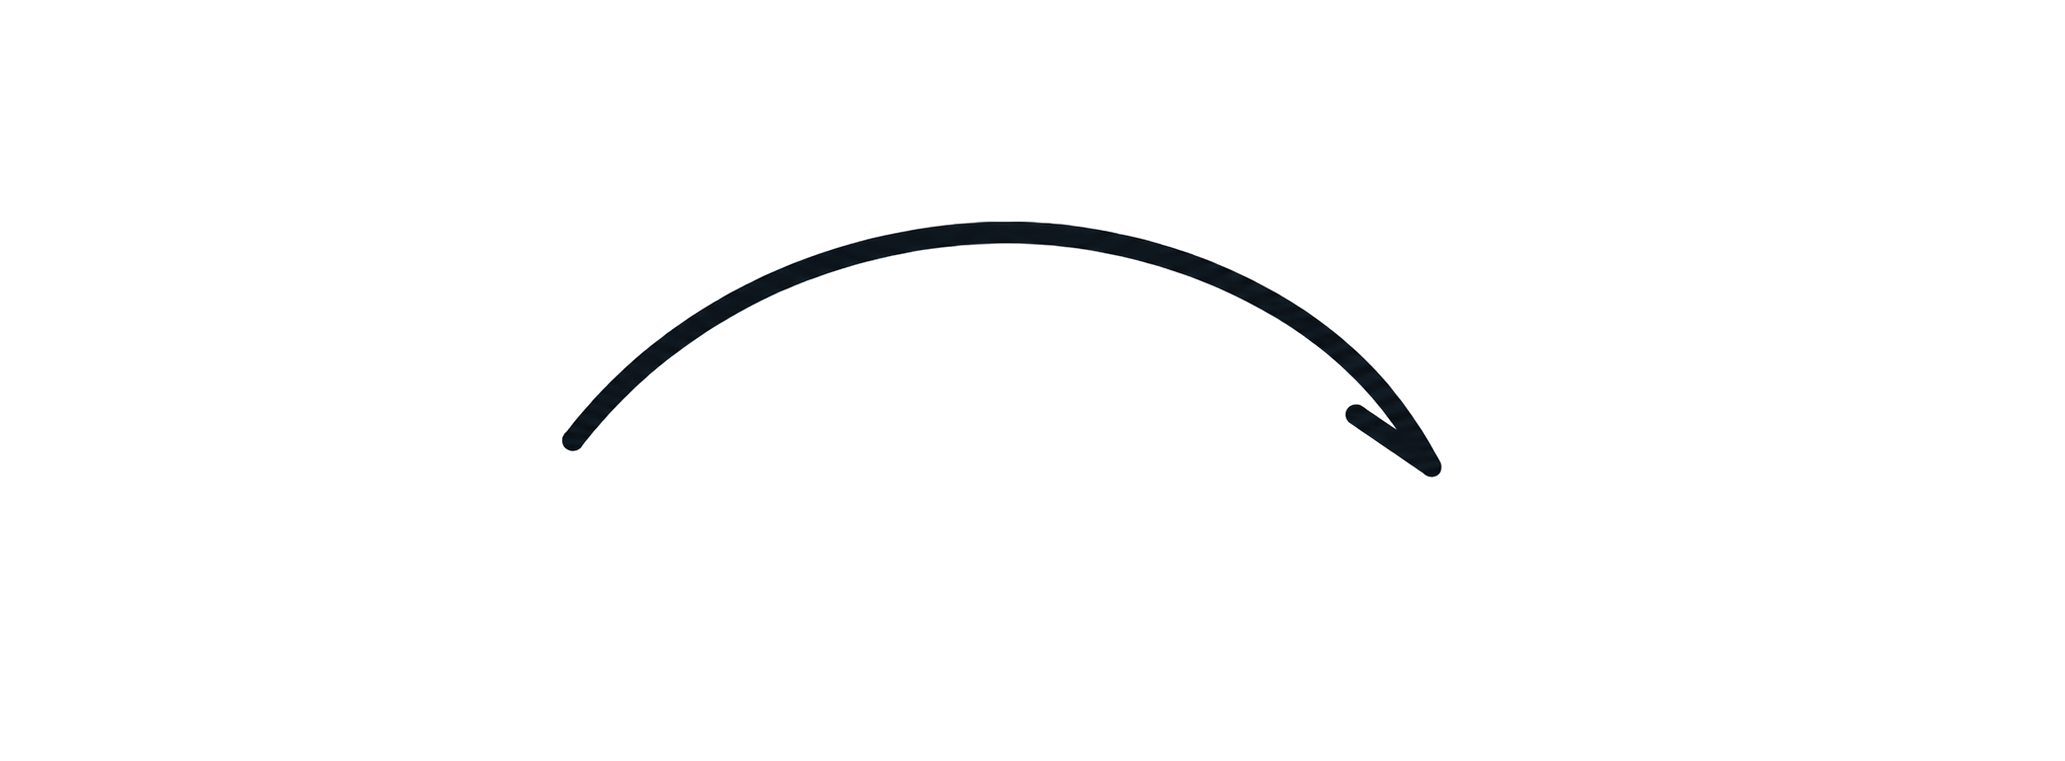
the arrow has one curved shaft and exactly one terminal barb; the symbol therefore represents movement of one electron; no double-barbed arrow or unrelated molecular content is present.
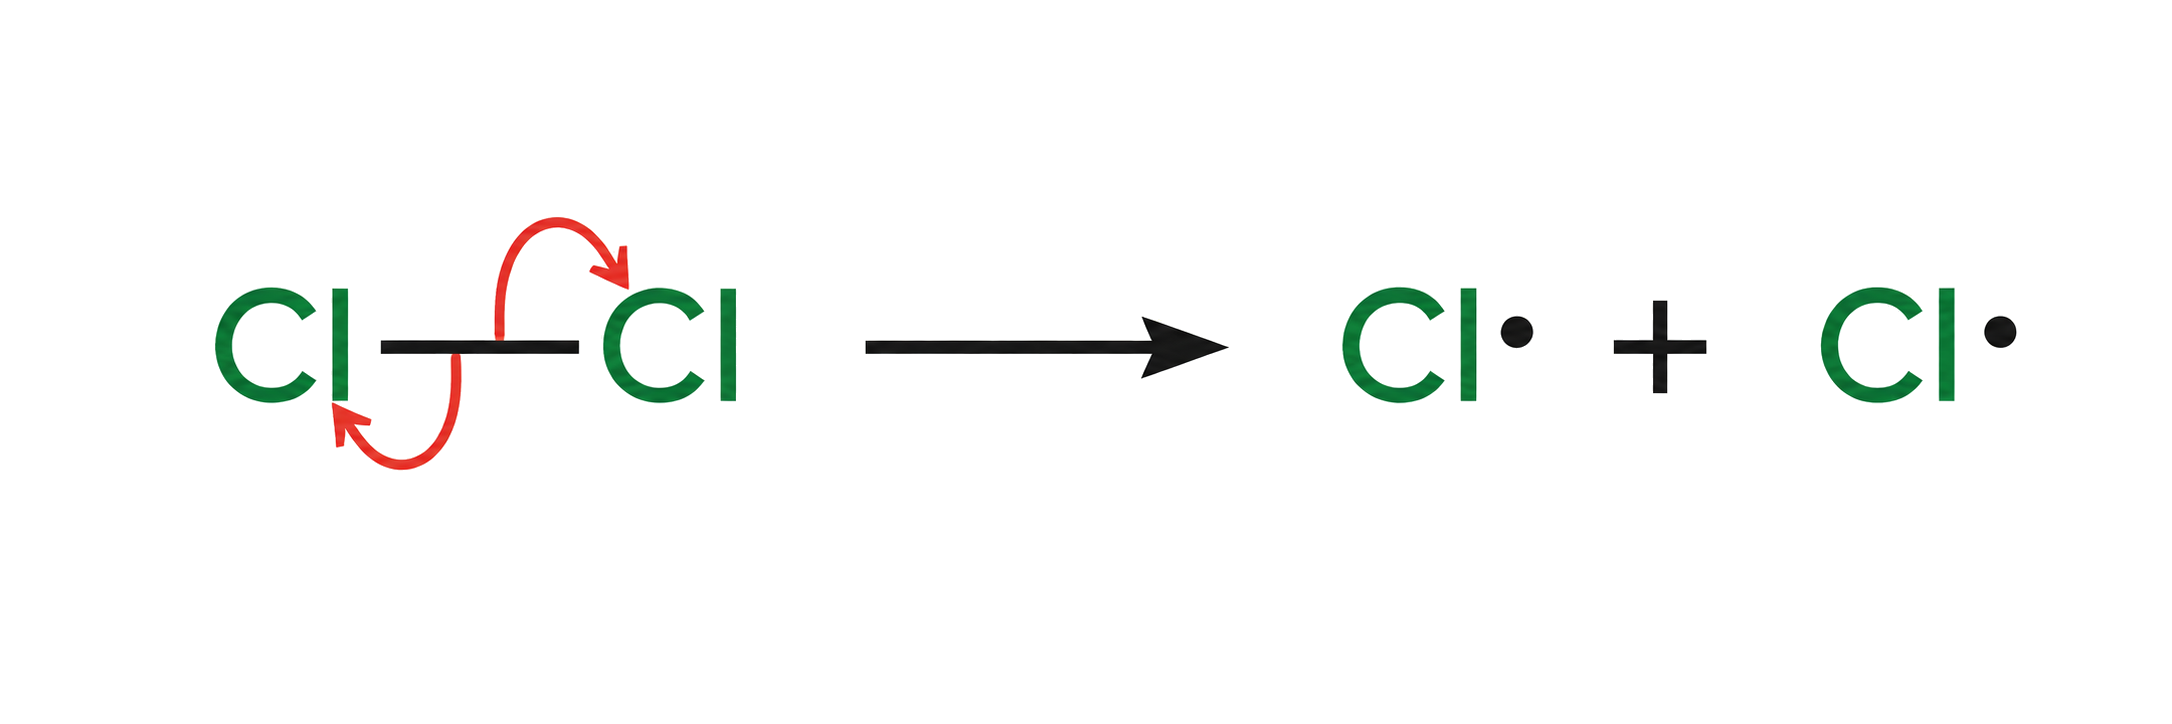
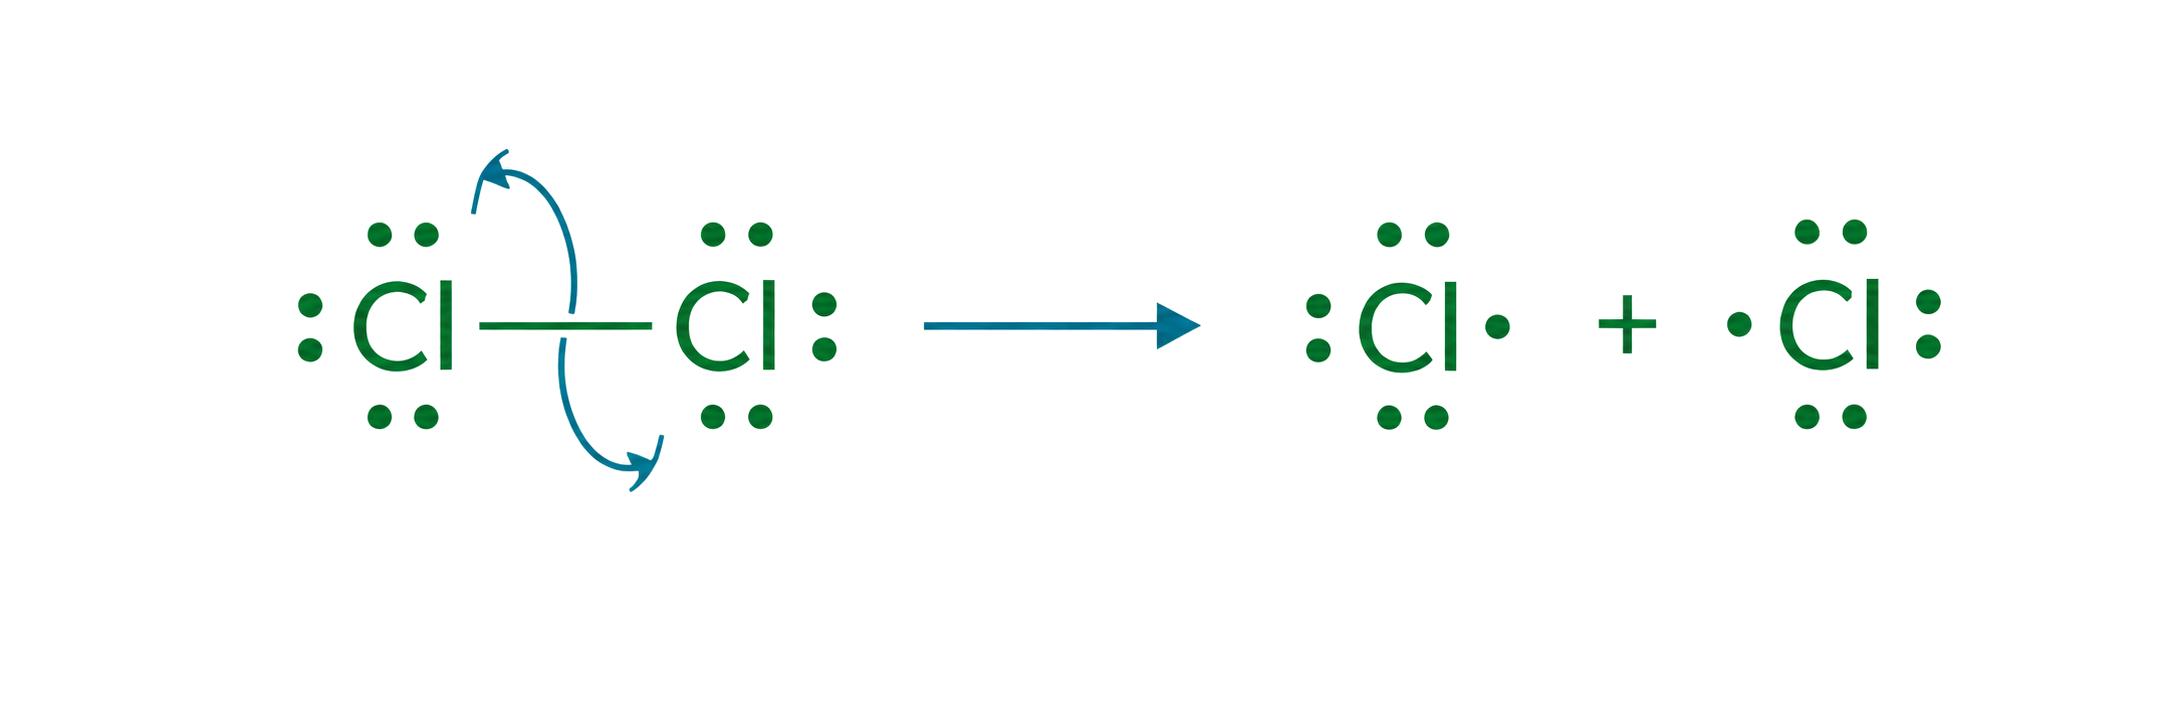
two single-barbed arrows split the Cl-Cl bonding pair equally; one arrow terminates at each chlorine atom; the products are exactly two neutral Cl radicals with one radical dot each.

Homolytic fission creates radicals because each covalent-bond fragment retains one bonding electron.

The dot in Cl· or CH₃· represents one unpaired electron, not a positive or negative charge. Radicals react readily because pairing that electron can form a bond; track the dot through every equation so electron and atom accounting remain explicit.
Recognizing Radical Notation

[ 1 ]

Which radical is most likely to form during the breakdown of one covalent bond of dichlorofluoromethane, CHCl2 F\mathrm{CHCl}_{2} \mathrm{~F}, in the upper atmosphere?

Homolytic Fission and Initiation

Cl2(g)→2Cl⋅(g)underUVlightorheatCl2(g) → 2Cl·(g) under UV light or heat

two single-barbed arrows split the Cl-Cl bonding pair equally; one arrow terminates at each chlorine atom; the products are exactly two neutral Cl radicals with one radical dot each.
Cl2 homolysis is shown with two single-electron fish-hook arrows; each reactant chlorine has three lone pairs; each product chlorine has three lone pairs and one unpaired electron; no formal charge is introduced.

Homolytic cleavage gives one electron to each fragment. Use single-electron arrows to show radical movement in the chain mechanism.

Draw two single-barbed arrows from the breaking X–X bond, one toward each atom, to account for both electrons. The UV or heat step creates radicals and is initiation; a step that consumes one radical and forms another belongs to propagation.

Writing the Initiation Step

[ 1 ]

Write an equation for the initiation reaction.

Free-Radical Substitution

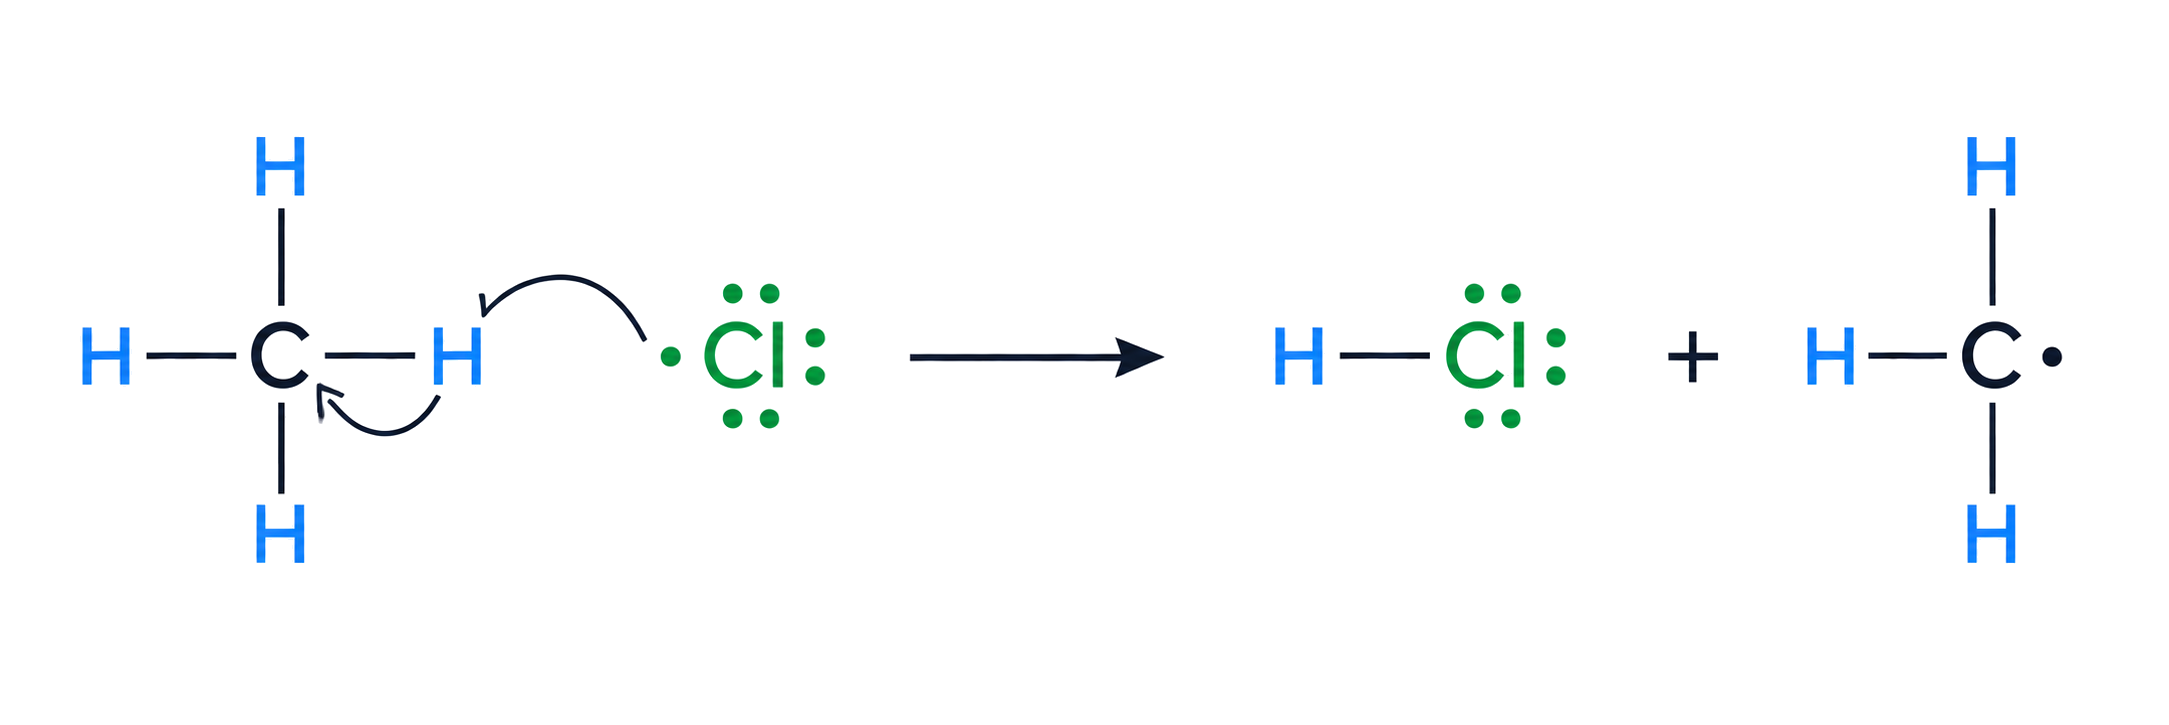
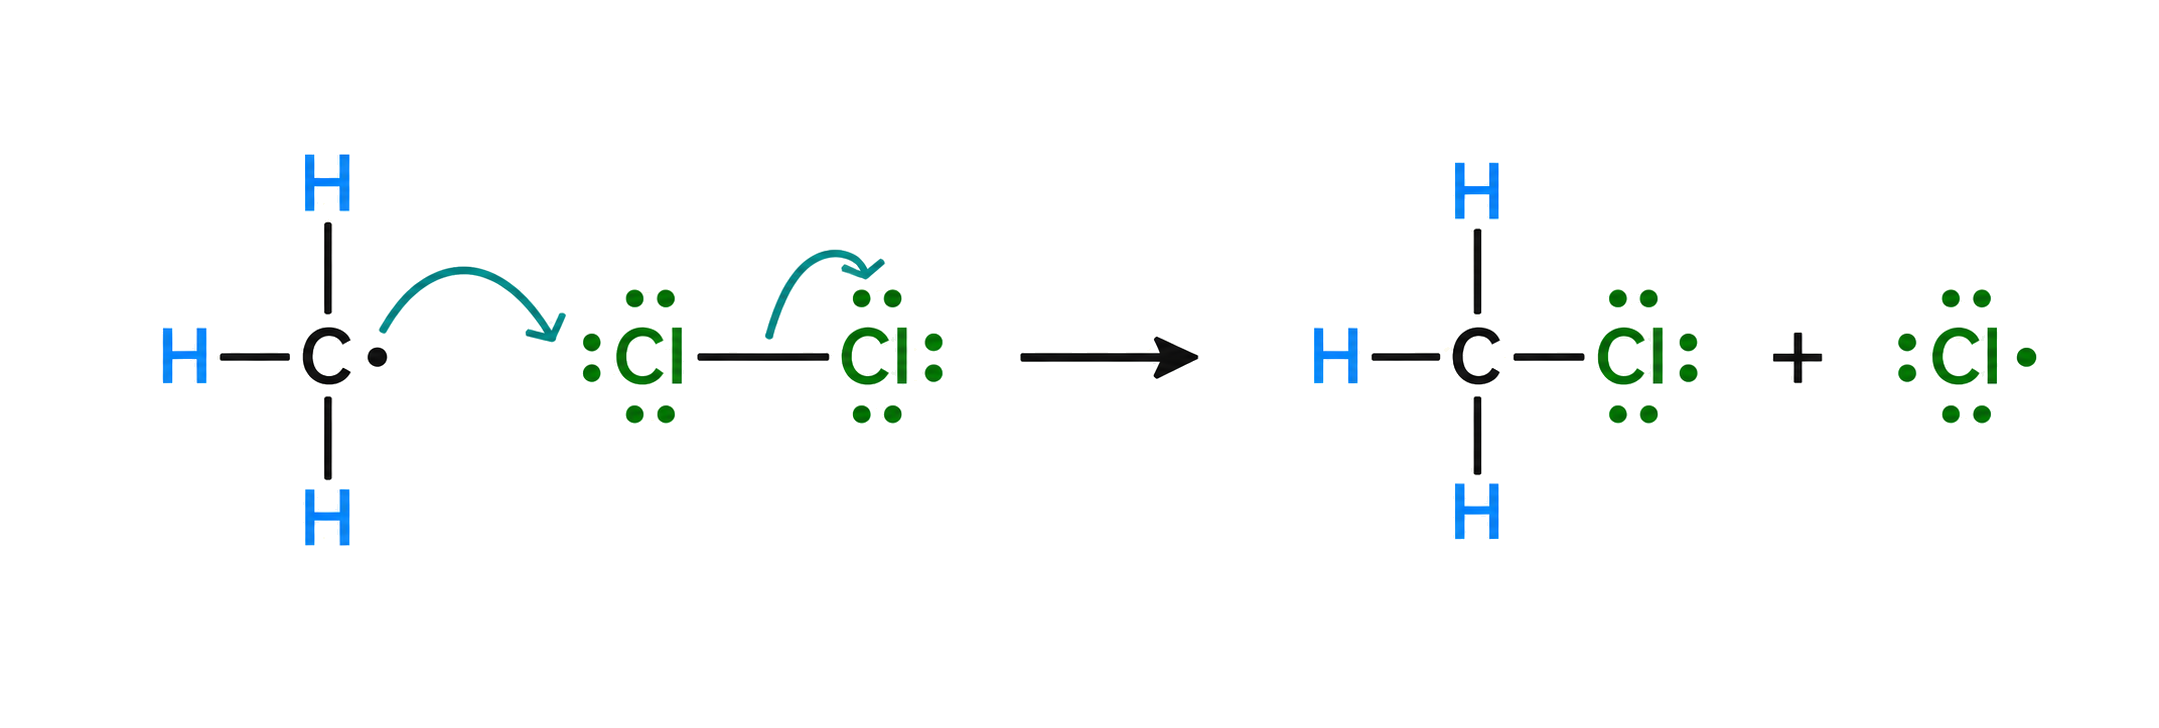
Initiation creates radicals; propagation abstracts H from an alkane and then regenerates the halogen radical; termination combines radicals. A mixture can form because substitution may occur at different positions.
Cl2 homolysis is shown with two single-electron fish-hook arrows; each reactant chlorine has three lone pairs; each product chlorine has three lone pairs and one unpaired electron; no formal charge is introduced.
the equation is Cl radical plus CH4 to HCl plus CH3 radical; one fish-hook arrow begins at the chlorine radical electron and ends at H; the second fish-hook arrow begins at the breaking C-H bond and ends at carbon; four reactant hydrogens become one HCl hydrogen plus three methyl-radical hydrogens.
the equation is CH3 radical plus Cl2 to CH3Cl plus Cl radical; the carbon radical electron moves to the nearer chlorine; one electron from the Cl-Cl bond moves to the departing chlorine; the chlorine radical is regenerated and all atoms are conserved.

C2H6+Cl⋅→C2H5⋅+HCl;C2H5⋅+Cl2→C2H5Cl+Cl⋅C2H6 + Cl· → C2H5· + HCl; C2H5· + Cl2 → C2H5Cl + Cl·

For methane chlorination, initiation forms 2Cl· from Cl₂ under UV. Propagation uses Cl· + CH₄ → HCl + CH₃· and CH₃· + Cl₂ → CH₃Cl + Cl·; termination combines two radicals. Further substitution creates a mixture, so the mechanism does not guarantee only CH₃Cl.

Writing Chain-Substitution Equations

[ 4 ]

Explain the reaction mechanism by writing equations for each step.

One initiation step:
Two propagation steps:

One termination step:

Radical Chain Summary

Retrieve the route: locate the unpaired electron, split the bond homolytically, initiate with UV or heat, propagate by single-electron steps and terminate by radical combination.

Every propagation step must regenerate a radical, and every radical symbol and single-electron movement must be shown where required.

3.4 Electron-pair sharing reactions

Syllabus
First assessment 2025
Topic
3.4
Level
HL

Recognizing Nucleophiles

A nucleophile is an electron-rich species that donates an electron pair to an electron-deficient centre.
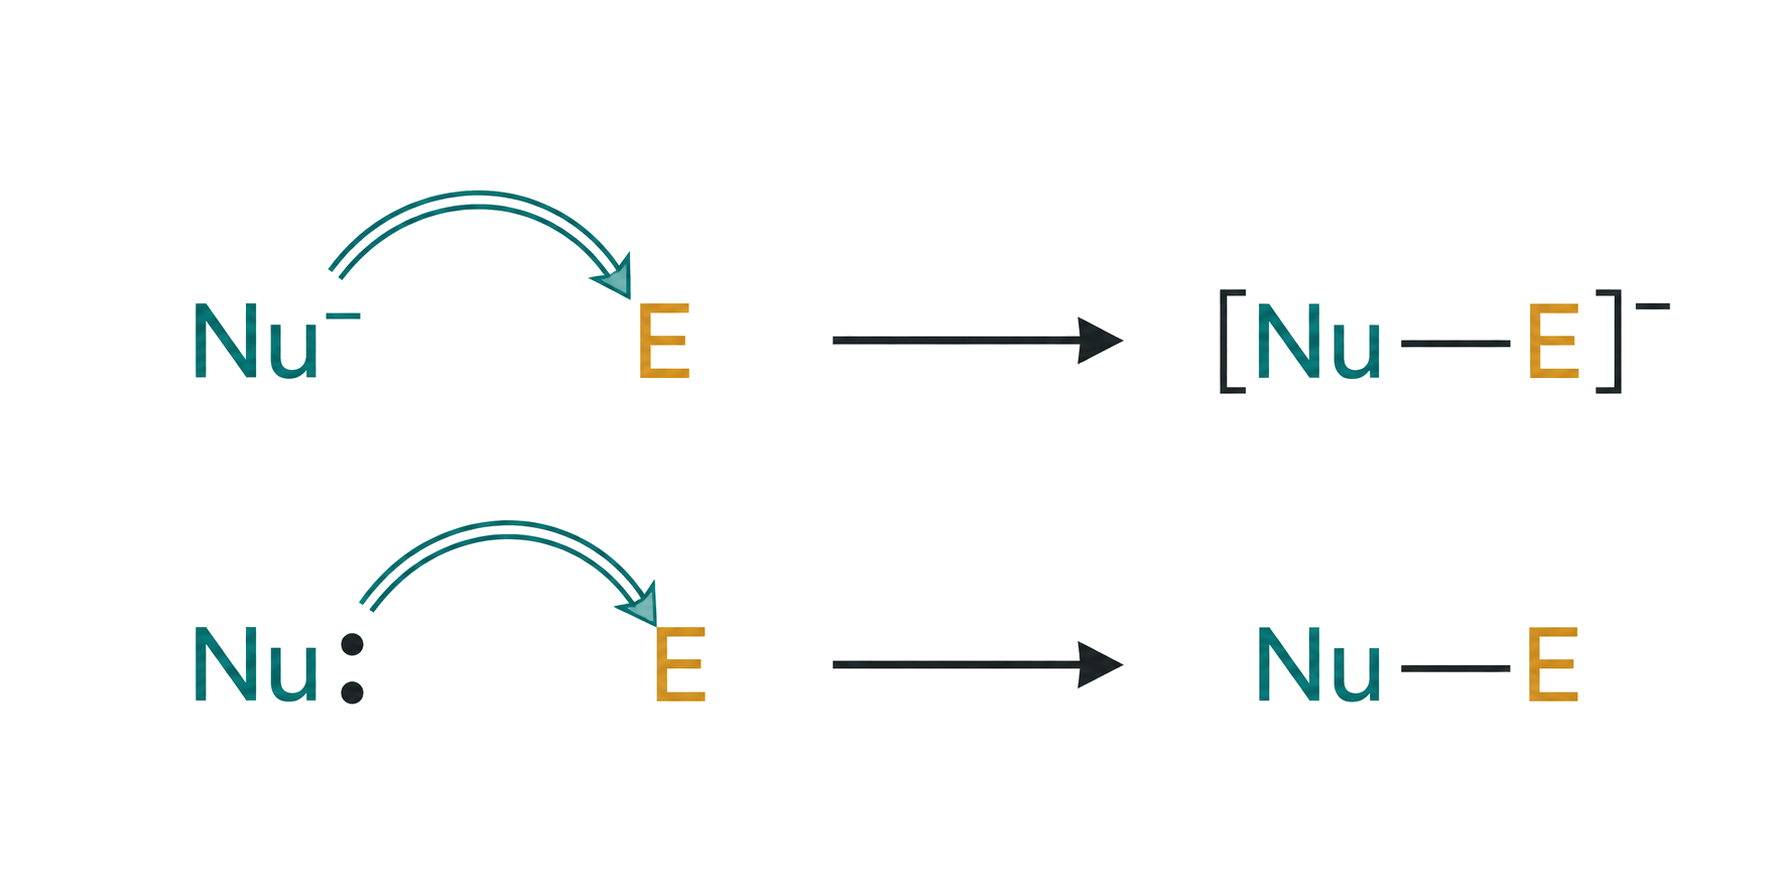
both curved arrows begin at the nucleophile electron pair and end at E; Nu- gives a negatively charged adduct while neutral Nu gives a neutral adduct; the coordinate bond is shown as electron-pair donation from Nu to E.

Look for an available electron pair: OH⁻ and CN⁻ use a negative charge and lone pair, while NH₃ uses a lone pair without being an anion. A curly arrow must start at that pair and point toward the atom where the new bond forms.

Identifying Nucleophiles

[ 1 ]

Identify a nucleophile which could be used for this reaction.

Nucleophilic Substitution

The nucleophile donates a pair to carbon while the leaving group departs with its bonding pair. Deduce the product by replacing the leaving group with the nucleophile.
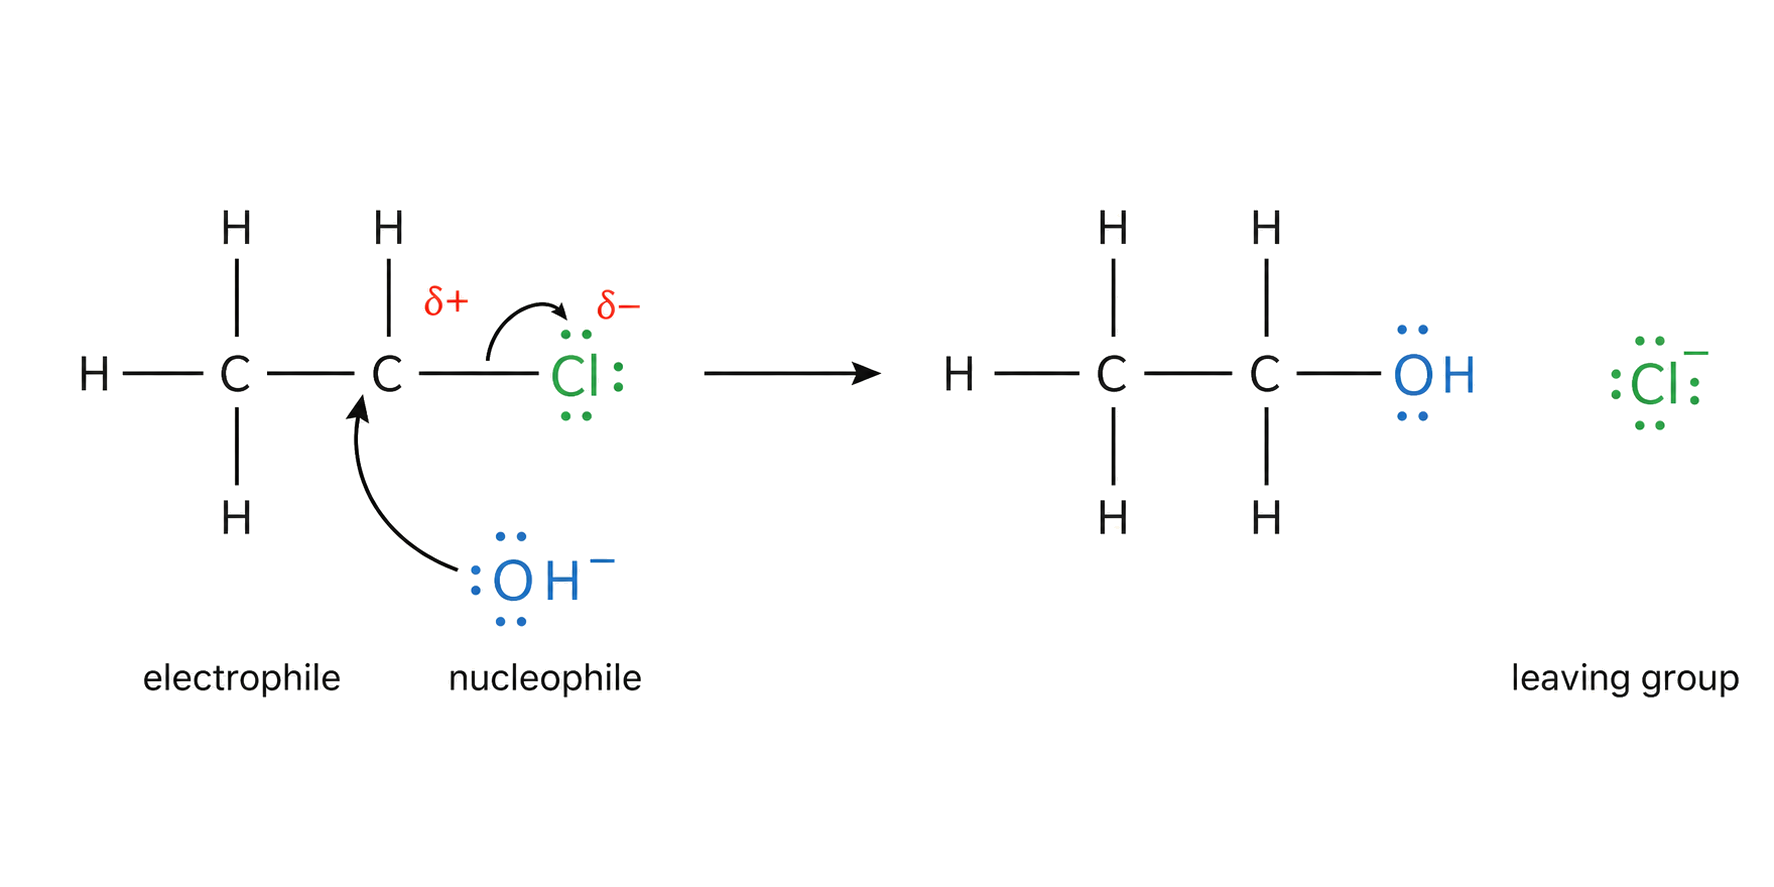
the hydroxide arrow starts at oxygen and ends at the carbon bearing chlorine; the leaving-group arrow starts at the C-Cl bond and ends at chlorine; carbon is delta positive and chlorine delta negative; CH3CH2OH and Cl- conserve every atom and charge.

For CH₃CH₂Br + OH⁻, the C–O bond forms as the C–Br bond breaks, giving CH₃CH₂OH + Br⁻. Account for charge and every atom in the product; the leaving group takes the bonding pair rather than departing as a neutral bromine atom.

Deducing Substitution Products

[ 4 ]

Explain the mechanism of the reaction, using curly arrows to represent the movement of electron pairs.

Heterolytic Fission

In heterolytic fission both bonding electrons remain with one fragment, producing ions. Curly arrows show movement of an electron pair.
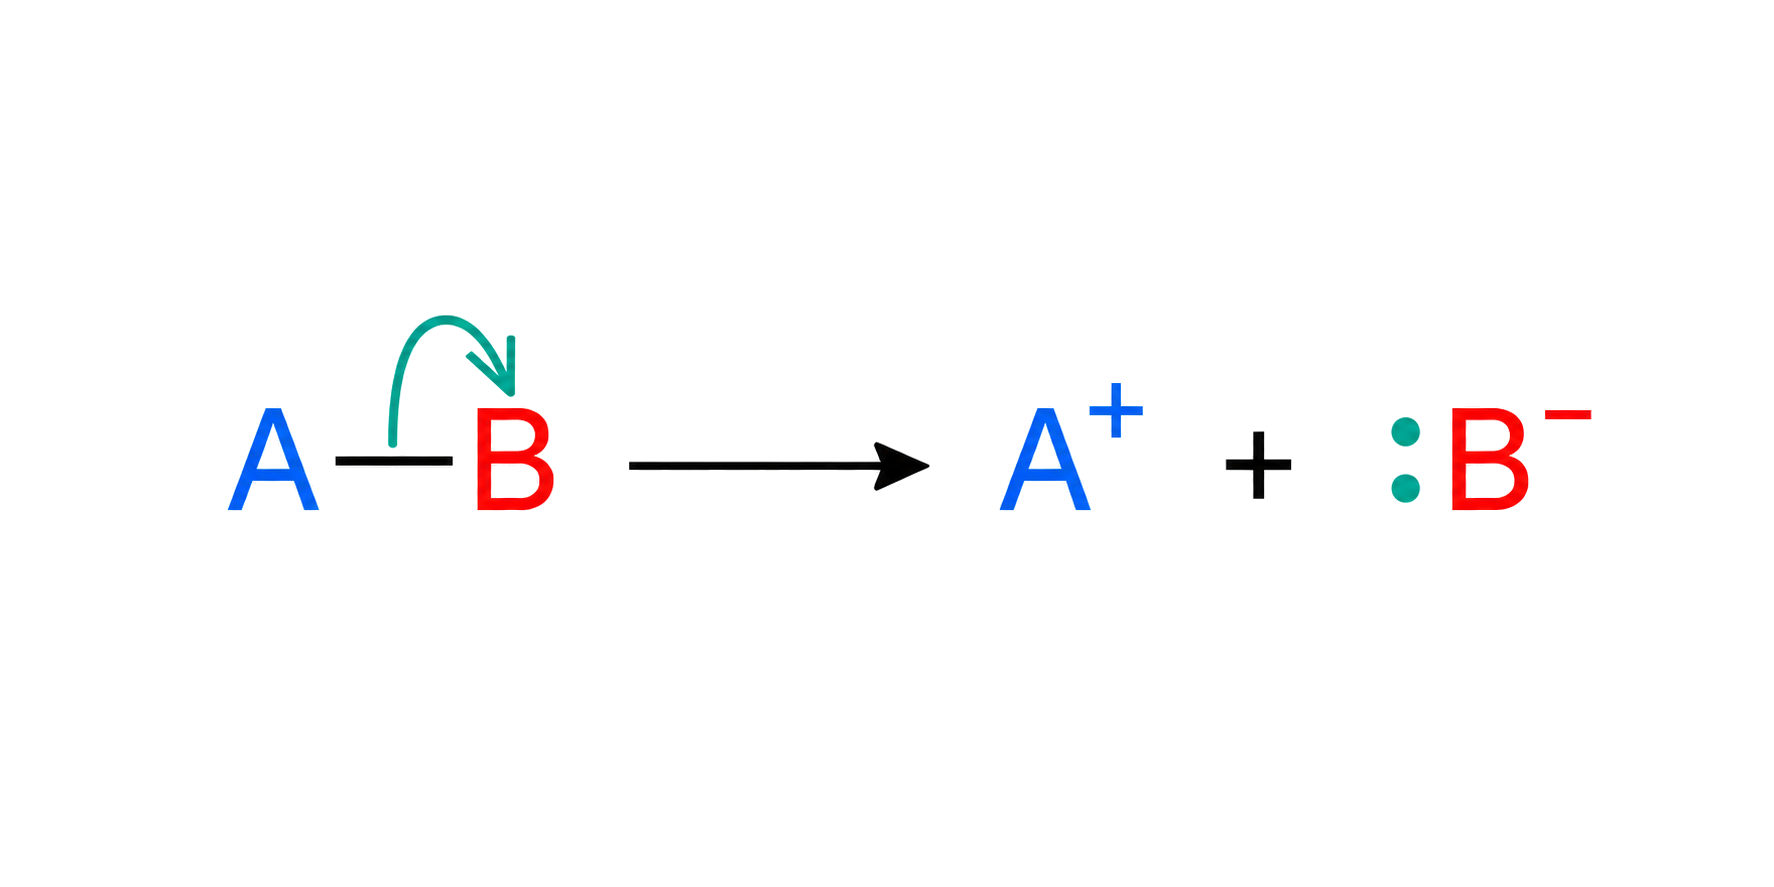
a full two-electron curly arrow begins at the A-B bond and ends at B; both bonding electrons are transferred to B; the products are exactly A plus and B minus.

Place the curly-arrow tail on the bond being broken and its head on the fragment receiving both electrons. Then assign charges from electron ownership: heterolysis creates ions, unlike homolysis, which gives one electron to each radical.

Showing Electron-Pair Movement

[ 2 ]

Contrast homolytic and heterolytic fission.

Homolytic fission:

Heterolytic fission:

Recognizing Electrophiles

An electrophile is an electron-deficient species that accepts an electron pair from a nucleophile.
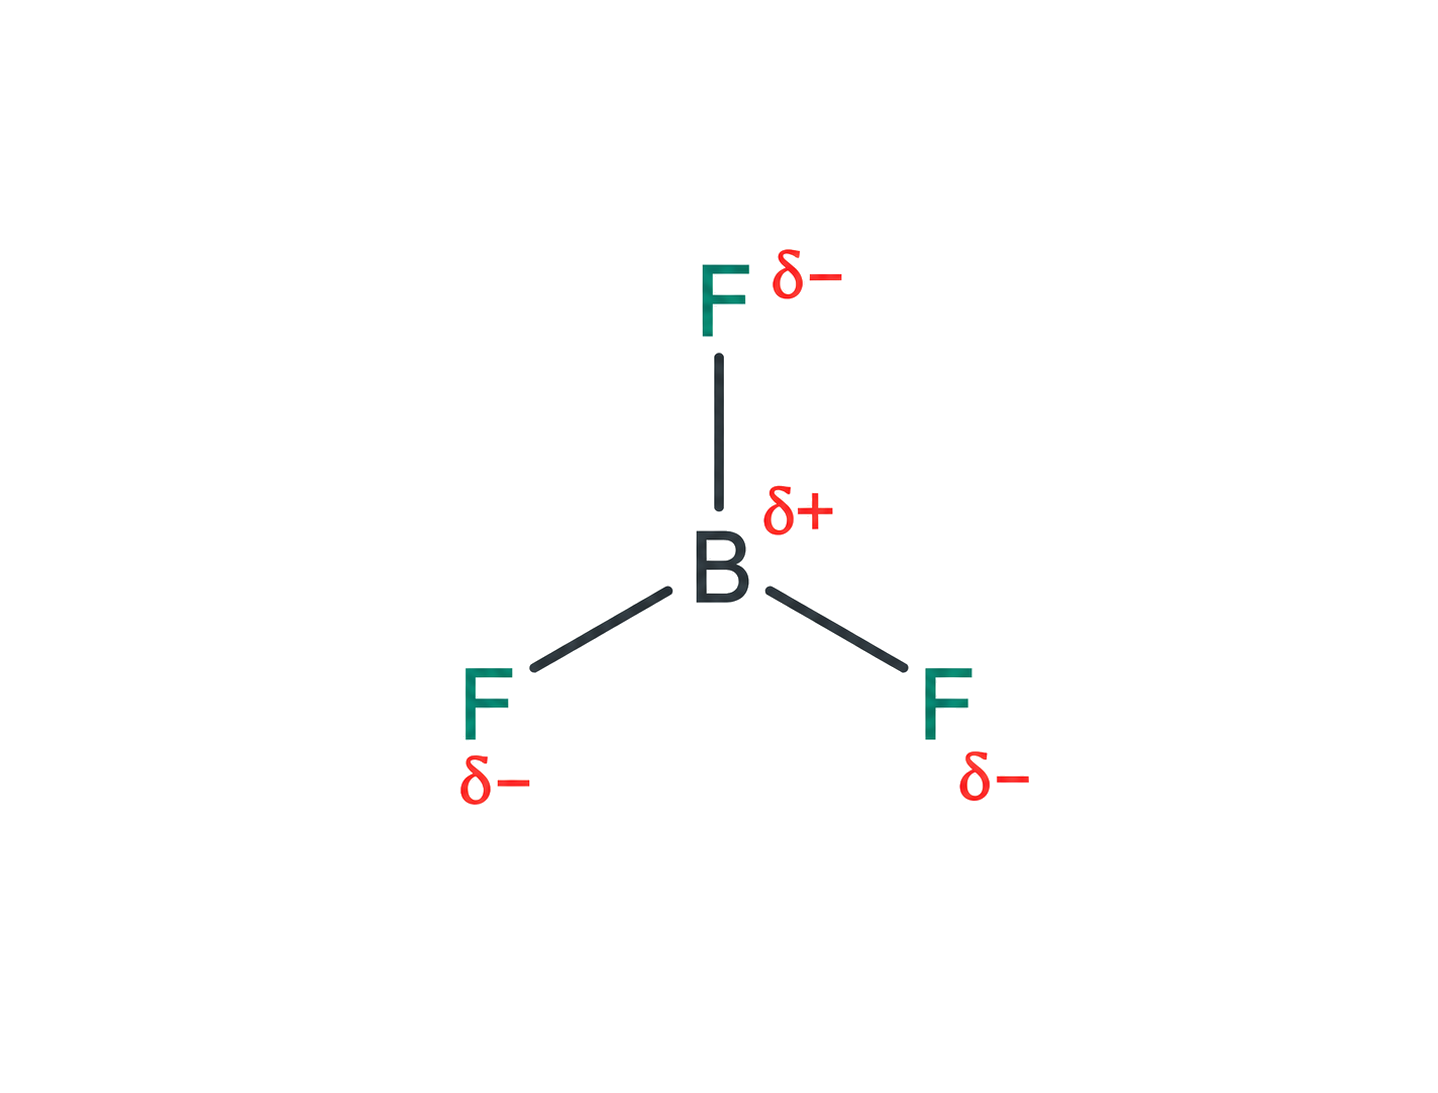
BF3 is trigonal planar with exactly three B-F single bonds; boron is labelled delta positive and each fluorine delta negative; no lone pair, formal charge, or fourth ligand is added to boron.
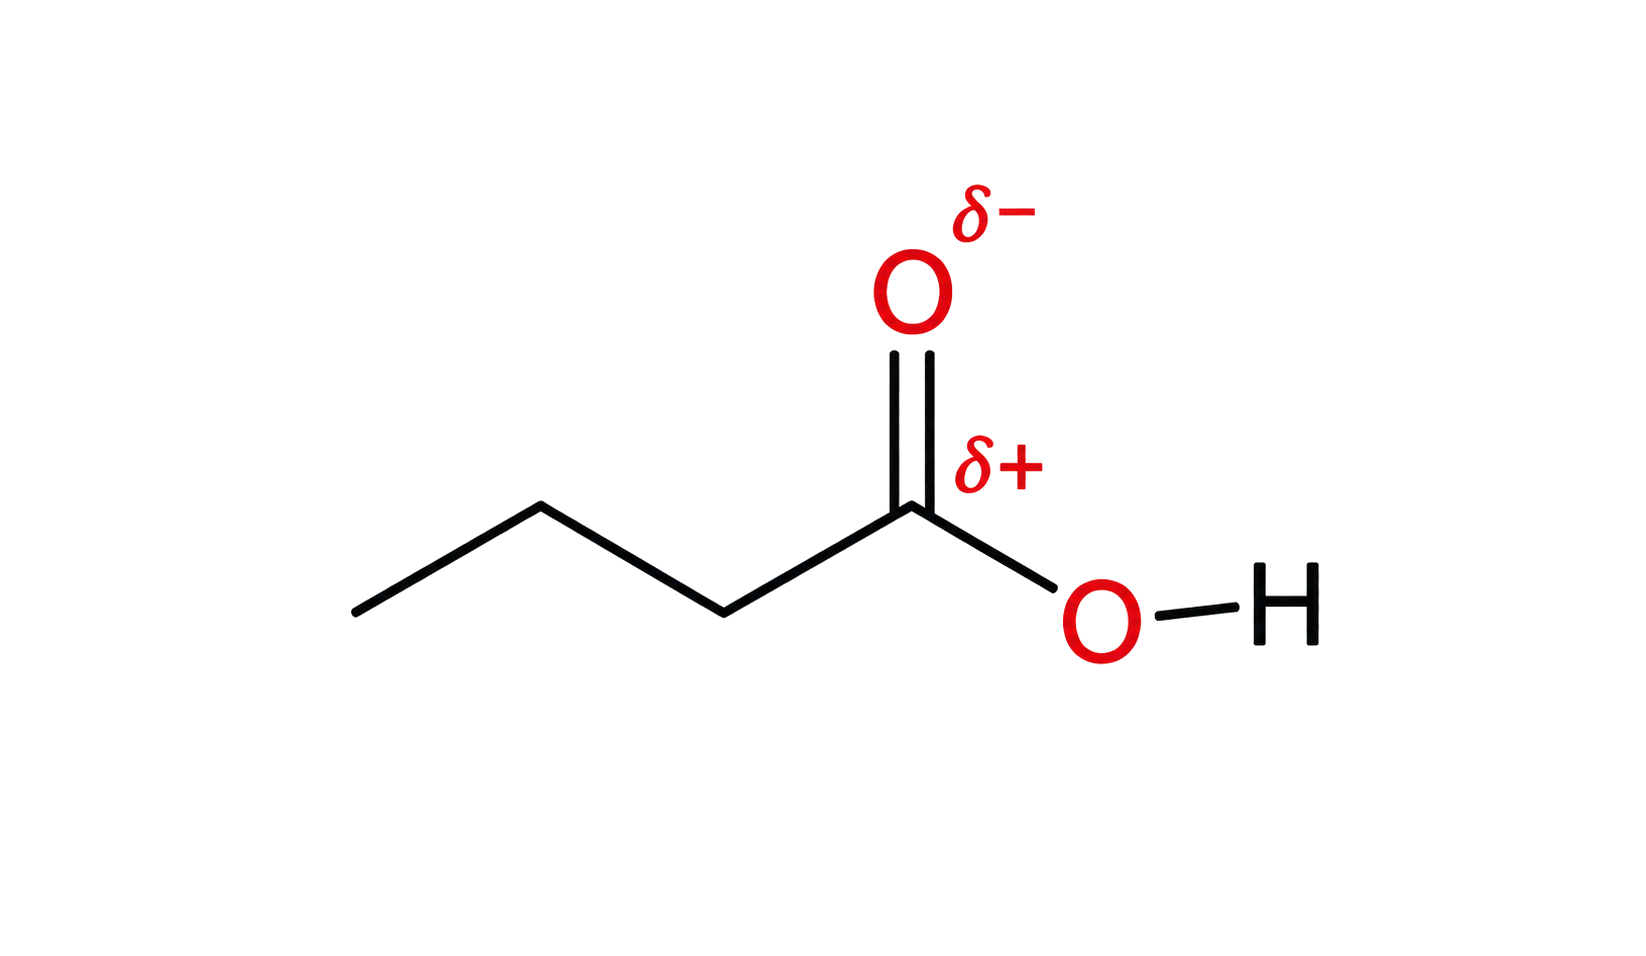
the skeletal structure is butanoic acid with four carbons; the C=O double bond and O-H bond are distinct; the carbonyl carbon is delta positive and carbonyl oxygen delta negative.

Identify the electron-poor atom, not merely a positive-looking formula. H⁺ and carbocations are electrophiles, and the δ⁺ carbon in a polar C–X bond can also accept a pair. The incoming curly arrow ends at this acceptor.

Recognizing Electrophiles

[ 1 ]

Which species is the electrophile?

CH3Br+OH−→CH3OH+Br−\mathrm{CH}_{3} \mathrm{Br}+\mathrm{OH}^{-} \rightarrow \mathrm{CH}_{3} \mathrm{OH}+\mathrm{Br}^{-}

Electrophilic Addition to Alkenes

The electron-rich C=C attacks an electrophile. Deduce addition products with water, halogens or hydrogen halides within the SL mechanism boundary.
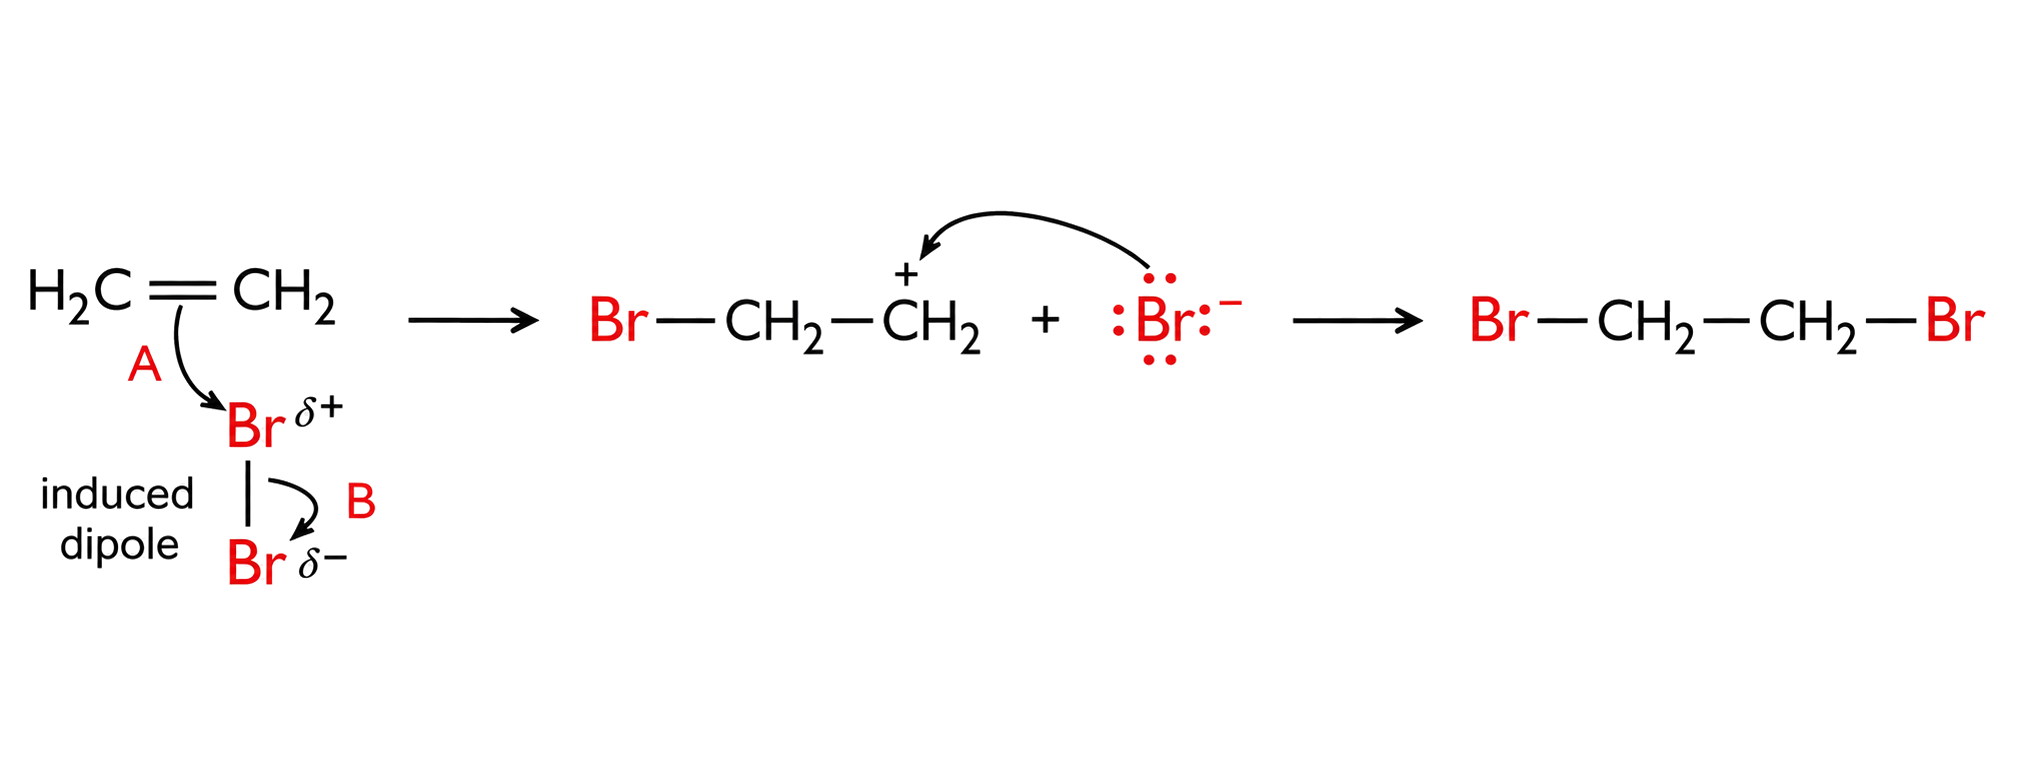
the C=C pi pair attacks Br delta positive and the Br-Br pair moves to Br delta negative; the textbook bromoethyl carbocation plus Br- intermediate is preserved; Br- attacks C+ and forms neutral 1,2-dibromoethane; two carbons, four hydrogens, and two bromines are conserved.

Treat the C=C as the reactive site and place the two added groups on its two carbon atoms. Bromine addition removes the double bond and forms a dibromoalkane; hydration forms an alcohol. At SL, deducing these products does not require a mechanism.

Reagent Groups added across C=C Product check
X₂ (for example Br₂) X and X vicinal dihalogenoalkane; C=C becomes C–C
HX H and X halogenoalkane; conserve the H and halogen from HX
H₂O/steam under acid-catalysed hydration conditions H and OH alcohol; conserve the carbon skeleton

At SL, use reagent and atom conservation to deduce products; curly-arrow mechanisms are not assessed in this card.

Deducing Alkene-Addition Products

[ 1 ]

Predict the product of the reaction between ethene and bromine.

Lewis Acids and Bases

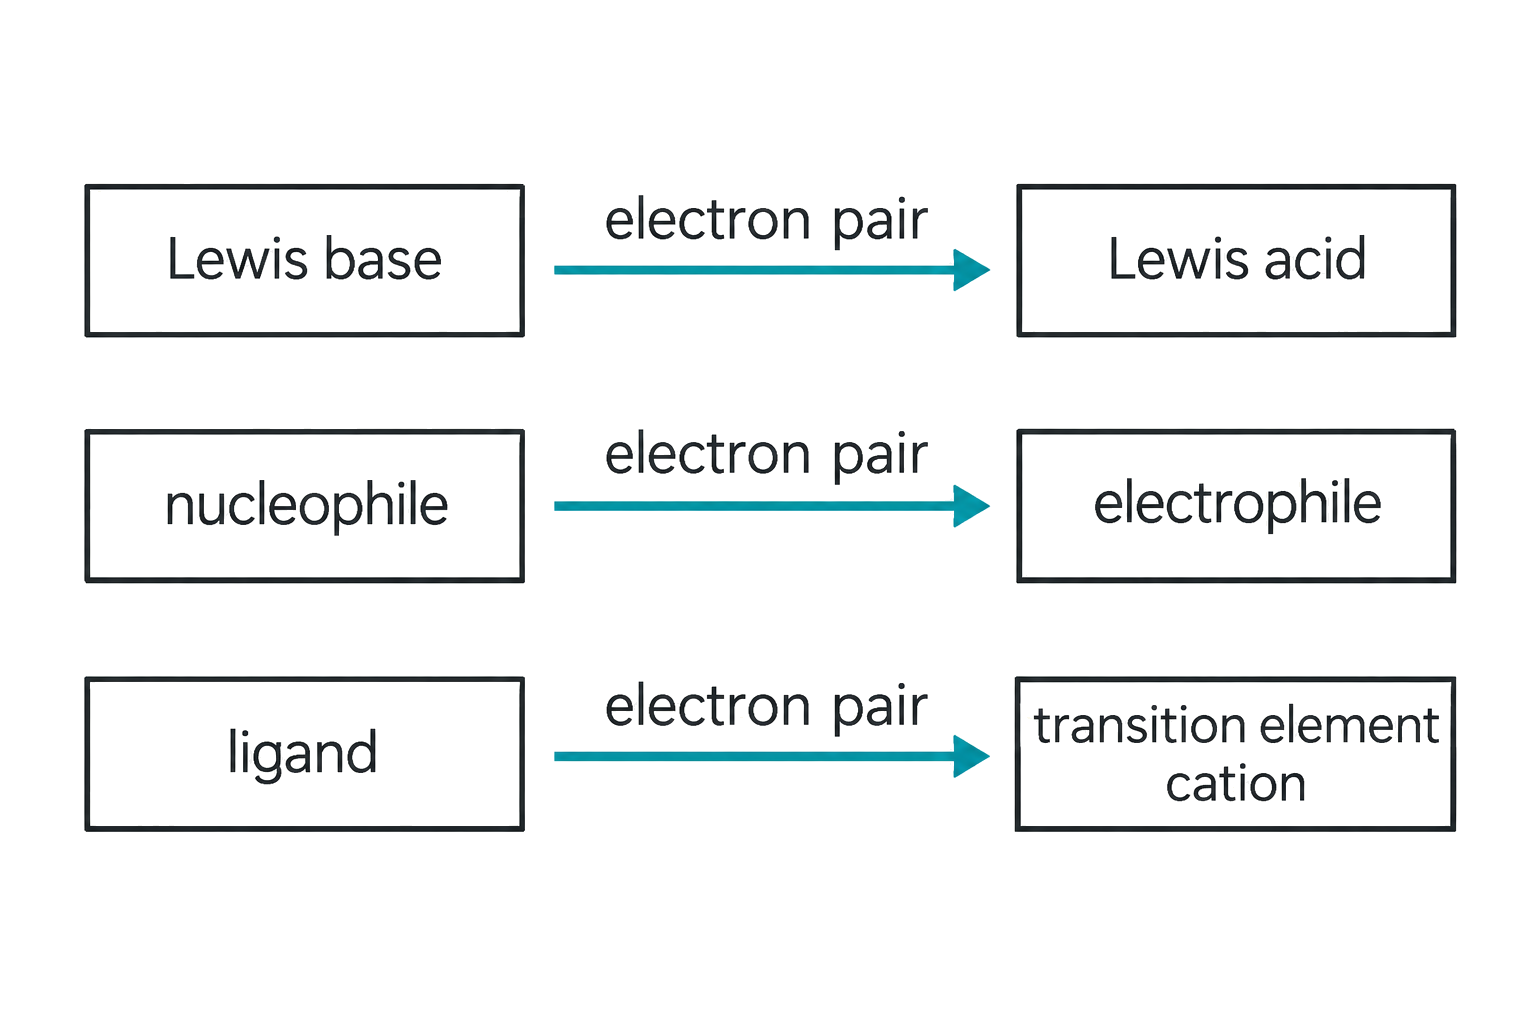
A Lewis acid accepts an electron pair; a Lewis base donates an electron pair. Nucleophiles correspond to Lewis bases and electrophiles to Lewis acids.

both curved arrows begin at the nucleophile electron pair and end at E; Nu- gives a negatively charged adduct while neutral Nu gives a neutral adduct; the coordinate bond is shown as electron-pair donation from Nu to E.

In BF₃ + NH₃ → F₃B←NH₃, NH₃ donates the pair and is the Lewis base; BF₃ accepts it and is the Lewis acid. Classify the roles from electron-pair movement rather than from whether H⁺ appears.

Classifying Lewis Acids and Bases

[ 1 ]

What is the role of the CN−\mathrm{CN}^{-}ion in the reaction of 1-chloropropane with excess KCN in ethanol?

C3H7Cl+KCN→C3H7CN+KCl\mathrm{C}_{3} \mathrm{H}_{7} \mathrm{Cl}+\mathrm{KCN} \rightarrow \mathrm{C}_{3} \mathrm{H}_{7} \mathrm{CN}+\mathrm{KCl}

Coordination Bonds

A ligand acts as a Lewis base and donates an electron pair to a Lewis-acid transition-metal cation, forming a coordinate bond.
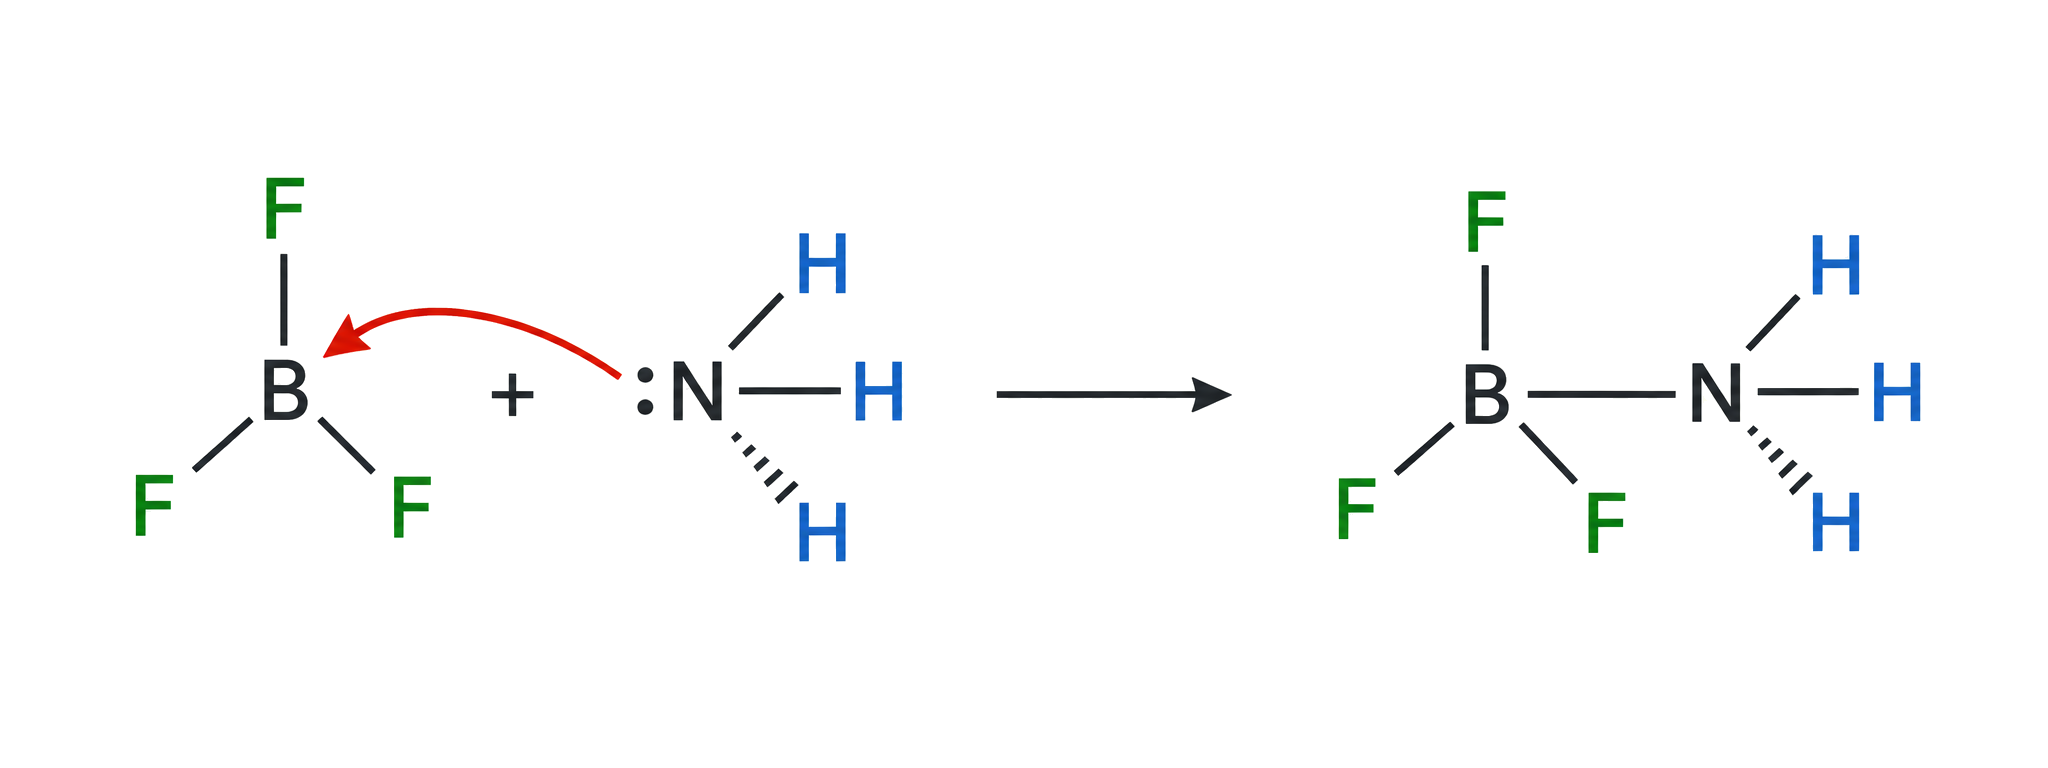
the full curly arrow starts at the nitrogen lone pair and ends at boron; reactants contain exactly BF3 and NH3; the product contains one B-N coordination bond with three F and three H; no proton transfer or extra ligand is implied.
Lewis base donates an electron pair to Lewis acid; nucleophile donates an electron pair to electrophile; ligand donates an electron pair to transition element cation; all three arrows run from donor on the left to acceptor on the right.

Show a coordination bond with an arrow from a ligand lone pair to the metal ion. The arrow records the origin of the shared pair; after formation the bond is not a different electrostatic species from other covalent bonds.

Explaining Coordinate-Bond Formation

[ 1 ]

Outline how ammonia acts as a Lewis base when it forms the complex ion

[Cu(NH3)4(H2O)2]2+(aq).\left[\mathrm{Cu}\left(\mathrm{NH}_{3}\right)_{4}\left(\mathrm{H}_{2} \mathrm{O}\right)_{2}\right]^{2+}(\mathrm{aq}) .

Ligands and Complex Ions

Identify the central transition-metal cation and the surrounding ligands. Each ligand donates an electron pair to the metal centre.
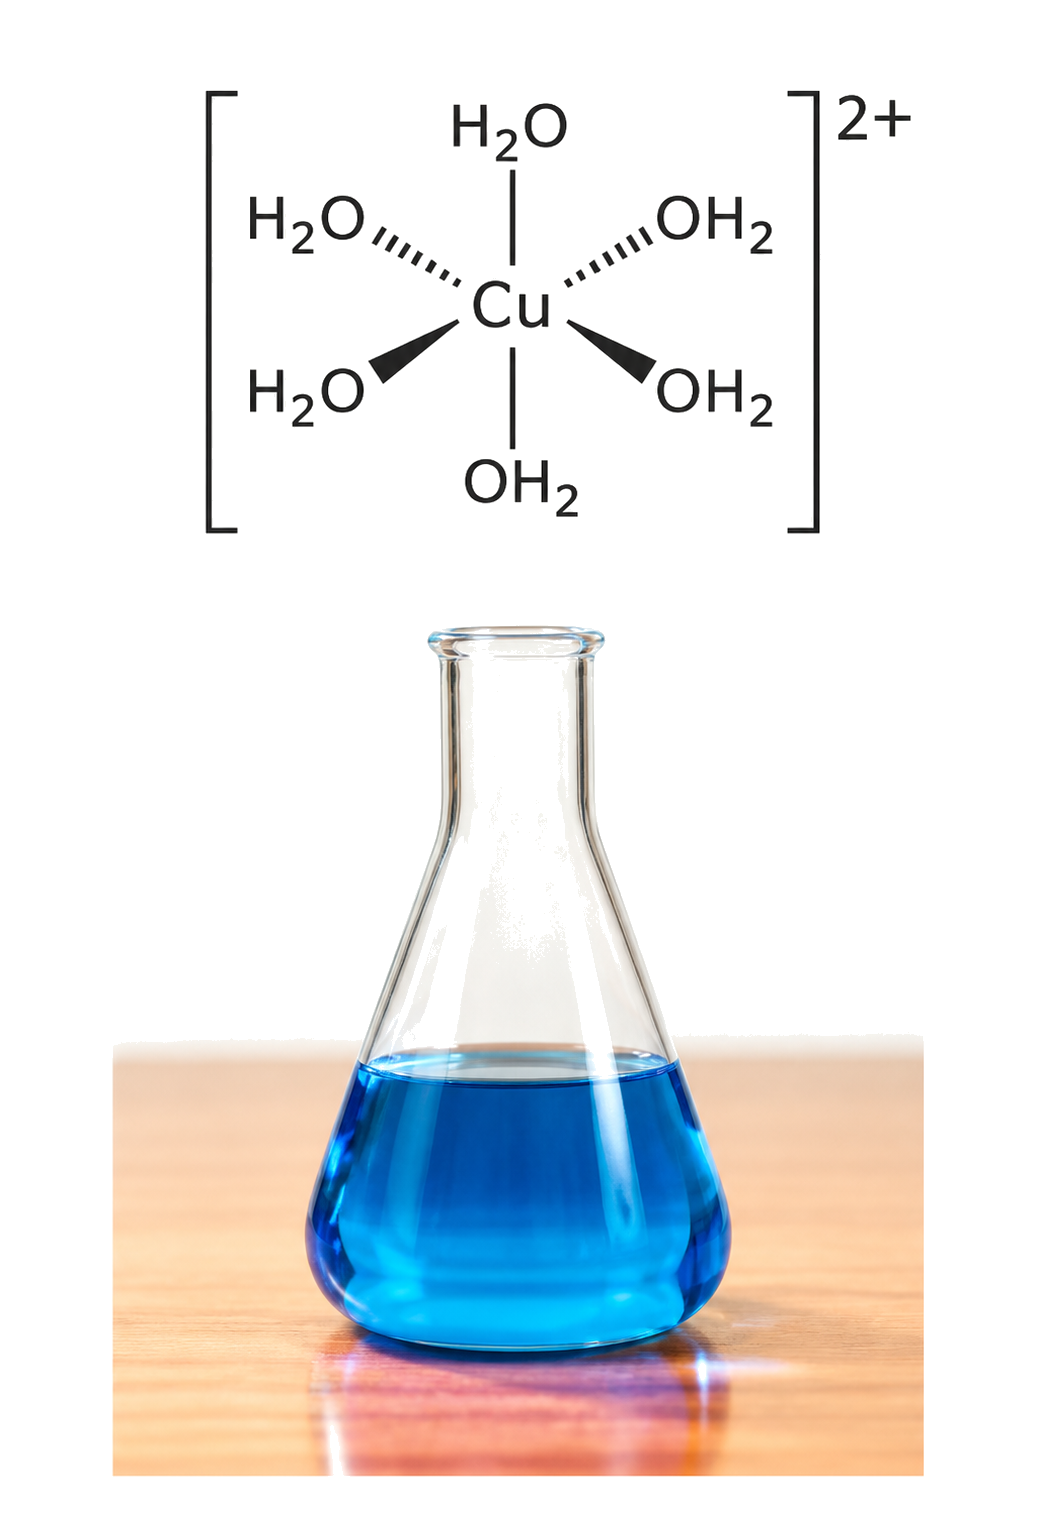
the bracketed ion contains one Cu and exactly six neutral H2O ligands; oxygen is the donor atom directed toward Cu; the six-coordinate geometry is octahedral; the overall charge is exactly 2 plus.

Read [Cu(NH₃)₄]²⁺ as one Cu centre with four NH₃ ligands and coordination number 4. Use ligand charges and the overall bracket charge to deduce the metal oxidation state; do not confuse coordination number with oxidation state.

Identifying Complex-Ion Components

[ 1 ]

Which statements are correct for the complex ion [FeCl4]2−\left[\mathrm{FeCl}_{4}\right]^{2-} ?

I. Chloride ions are behaving as ligands.
II. The oxidation state of iron is +3 .
III. Iron ion forms coordination bonds with chloride ions.

SN1 and SN2 Substitution

HL only
Mechanism Typical substrate/context Steps and stereochemical result
SN2 primary; some secondary one concerted backside attack; inversion at a stereogenic centre
SN1 tertiary; some secondary two steps through a planar carbocation; both configurations can form
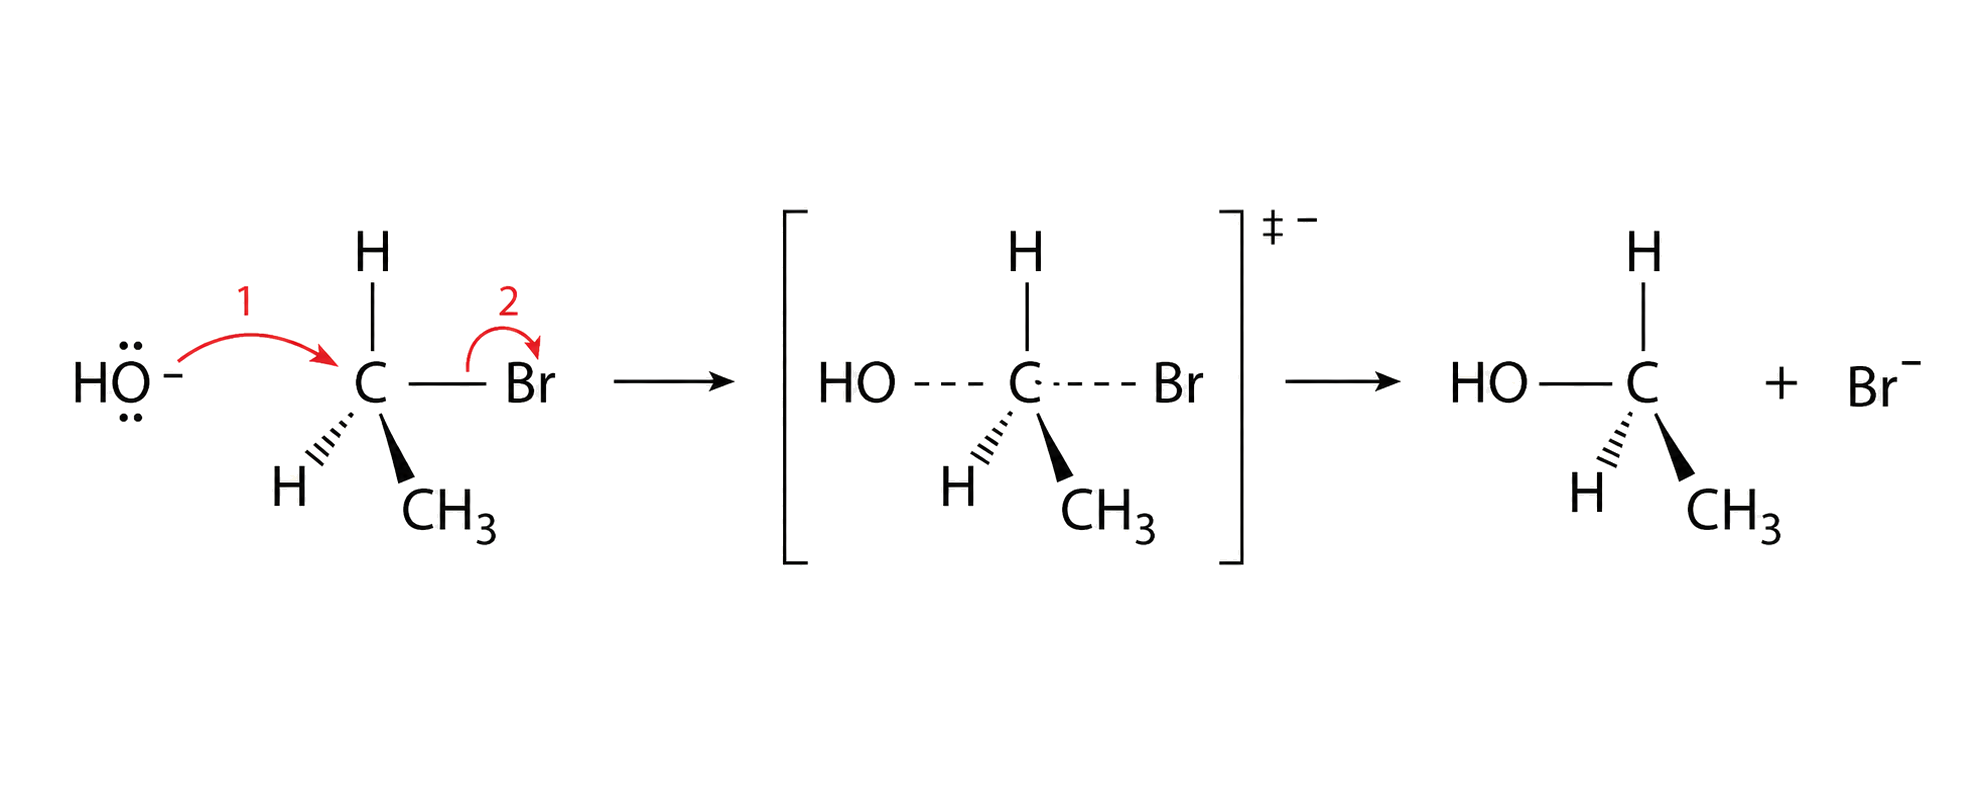
hydroxide attacks opposite bromine in one concerted step; the transition state is bracketed, overall negative, and has partial HO-C and C-Br bonds; no carbocation intermediate is introduced; ethanol and Br- are both shown with conserved atoms and charge.
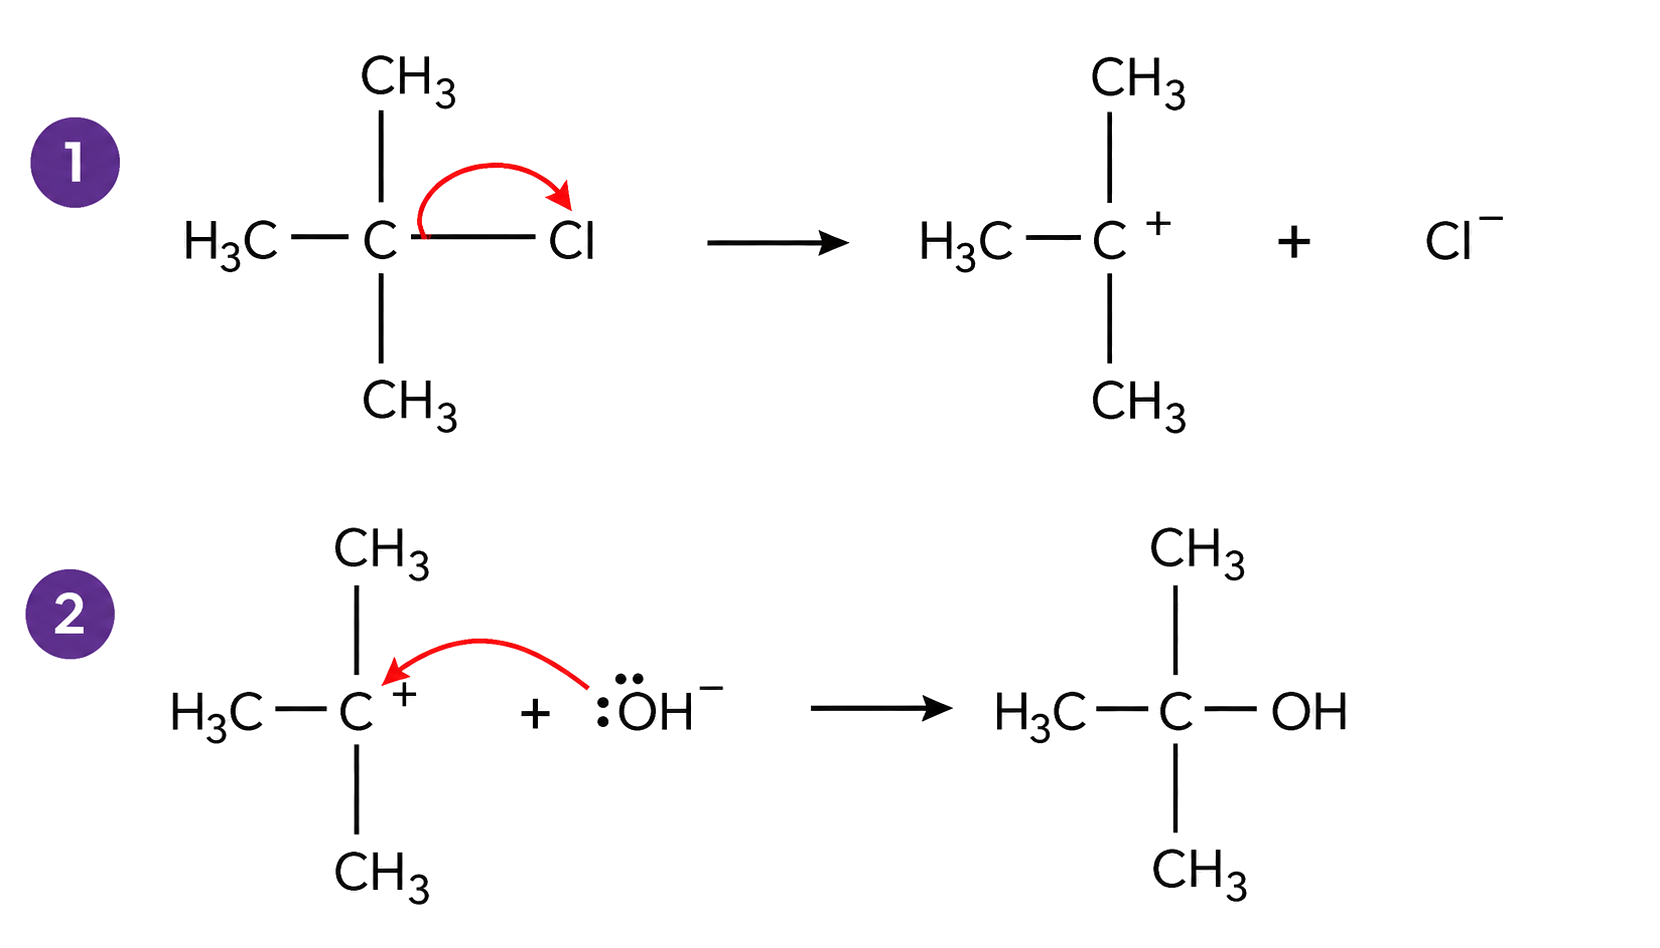
C-Cl heterolysis occurs before hydroxide attack; the tertiary carbocation has three CH3 substituents and an explicit positive charge; hydroxide donates its electron pair to C+ in the second step; Cl- appears once as the leaving-group product.

SN2 is a concerted backside attack: the nucleophile bonds as the carbon–halogen bond breaks. SN1 first forms a carbocation; the nucleophile then attacks the planar intermediate. Primary substrates generally favour SN2 and tertiary substrates generally favour SN1 because steric access and carbocation stability differ. Secondary substrates can support either mechanism, so use the conditions as well as the substrate class.

Do not treat SN1 and SN2 as labels that follow substrate class with no exceptions, or confuse inversion in SN2 with the mixture possible after planar SN1 attack.

Comparing SN1 and SN2

HL only

[ 4 ]

Sketch the mechanism of the reaction for step 1 in part (b), using curly arrows to show the movement of electron pairs.

Leaving Groups and Rate

HL only

Leaving-group identity affects substitution rate through carbon–halogen bond enthalpy and polarity: a weaker, more easily broken bond generally permits faster departure.

For comparable halogenoalkanes, the weaker C–I bond usually makes iodide a better leaving group and substitution faster than for chloride. Explain the trend with bond enthalpy and the actual rate-determining bond change, not polarity alone.

Explaining Leaving-Group Effects

HL only

[ 2 ]

Explain why CH3CHIC(CH3)3\mathrm{CH}_{3} \mathrm{CHIC}\left(\mathrm{CH}_{3}\right)_{3} reacts faster than CH3CHBrC(CH3)3\mathrm{CH}_{3} \mathrm{CHBrC}\left(\mathrm{CH}_{3}\right)_{3}.

Electrophilic-Addition Mechanisms

HL only

Use curly arrows from the alkene π bond to the electrophile, then from the intermediate to the nucleophile. Halogens may pass through a halonium intermediate; HX or water may pass through a carbocation.
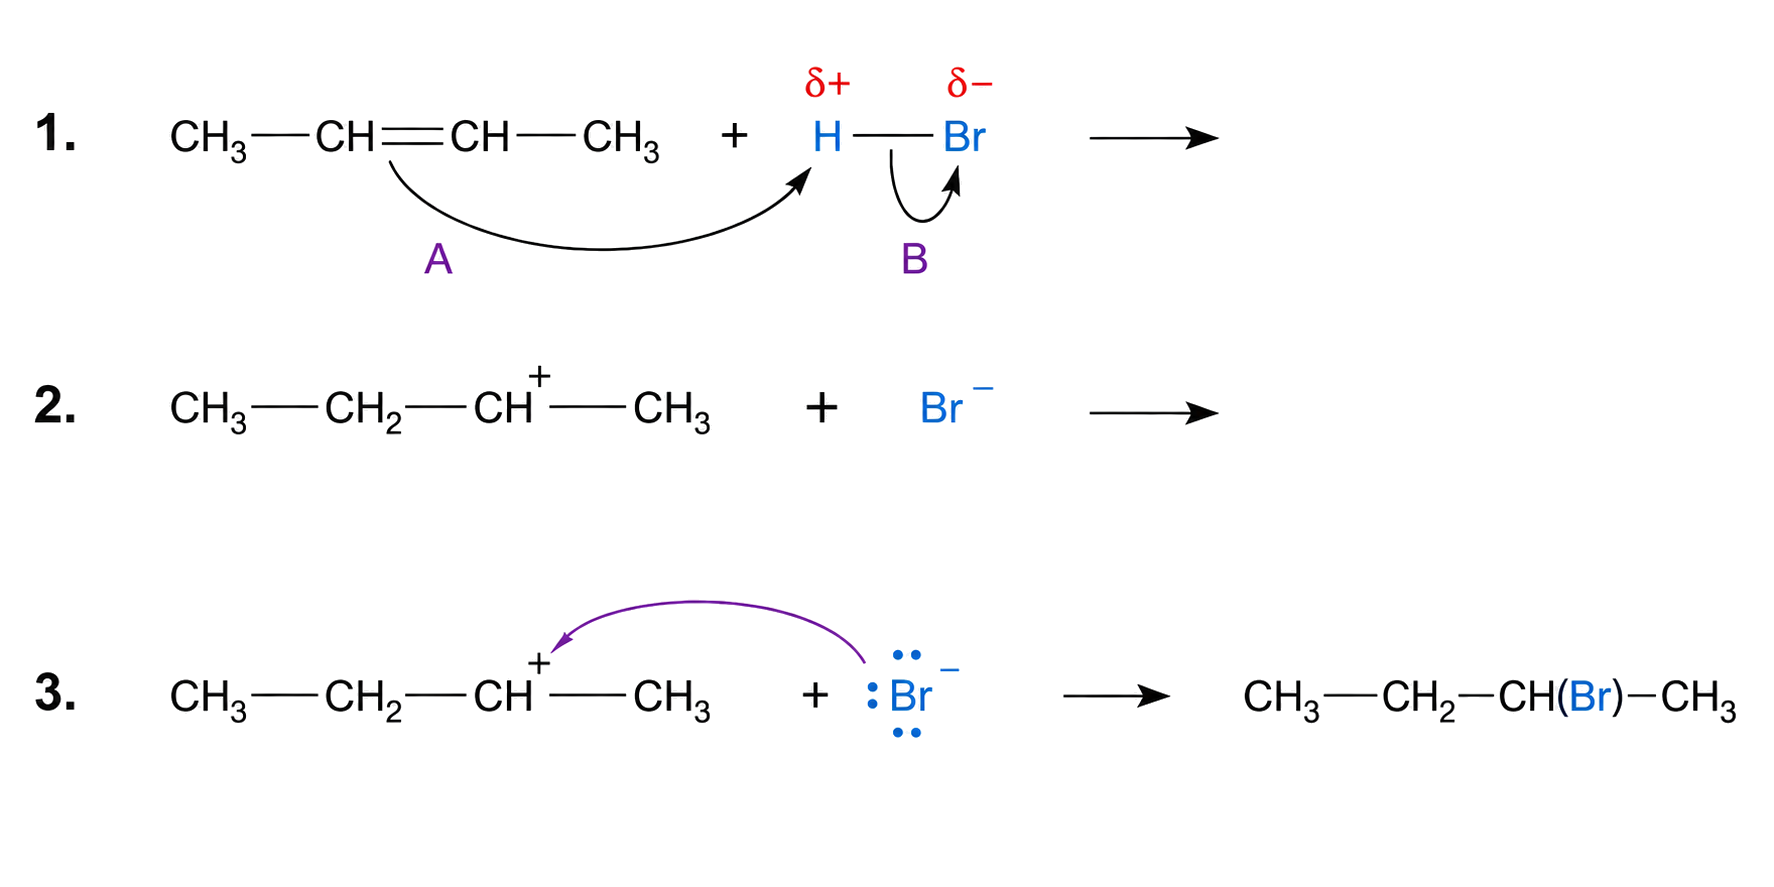
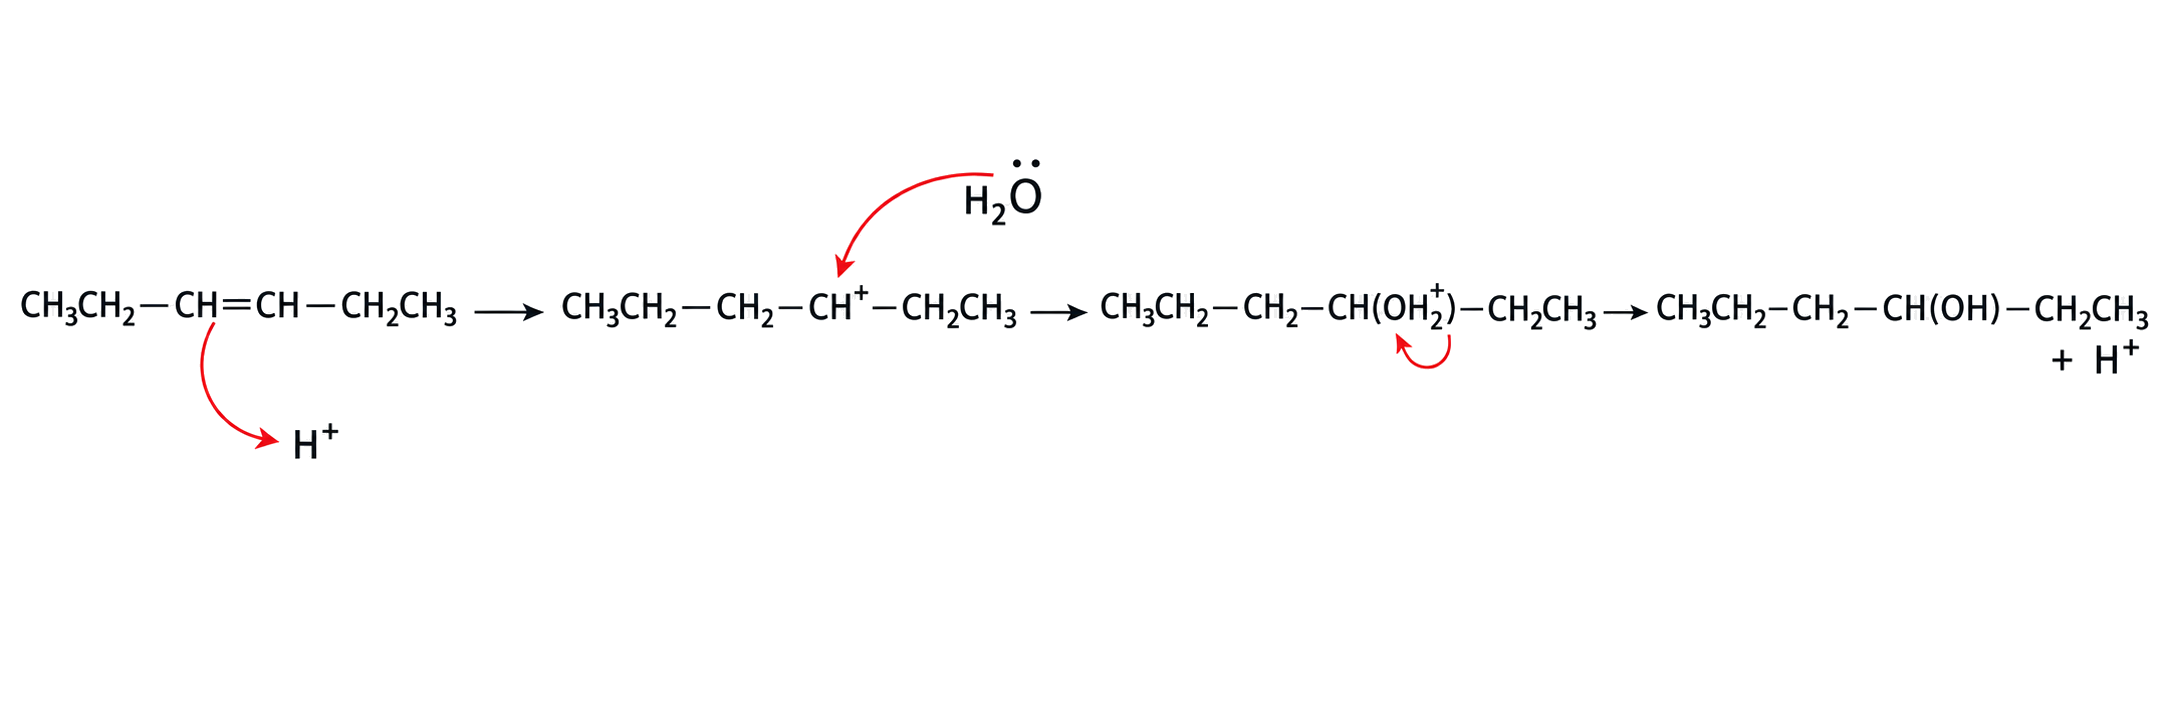
the C=C pi pair attacks Br delta positive and the Br-Br pair moves to Br delta negative; the textbook bromoethyl carbocation plus Br- intermediate is preserved; Br- attacks C+ and forms neutral 1,2-dibromoethane; two carbons, four hydrogens, and two bromines are conserved.
the pi bond attacks H delta positive and the H-Br bond pair moves to Br; the intermediate is the valid secondary butyl carbocation plus Br-; the bromide lone pair attacks C+; the final condensed formula is 2-bromobutane with no duplicated hydrogens.
hex-3-ene protonation forms the central secondary carbocation; water attacks from an oxygen lone pair to C+; the oxonium intermediate is positively charged and deprotonates to neutral hexan-3-ol; H+ is regenerated and six carbons are conserved without duplicate hydrogens.

Begin with an arrow from the alkene π bond to the electrophile. HX or acid-catalysed hydration can form a carbocation, whereas halogen addition uses a bridged halonium ion; the next arrow must start from the nucleophile's pair.

Drawing Addition Mechanisms

HL only

[ 3 ]

Describe the mechanism of this reaction, using curly arrows to represent the movement of electron pairs.

Major Products of Unsymmetrical Addition

HL only

For HX or water addition to an unsymmetrical alkene, compare the possible carbocations and select the pathway through the more stable carbocation as the major product.
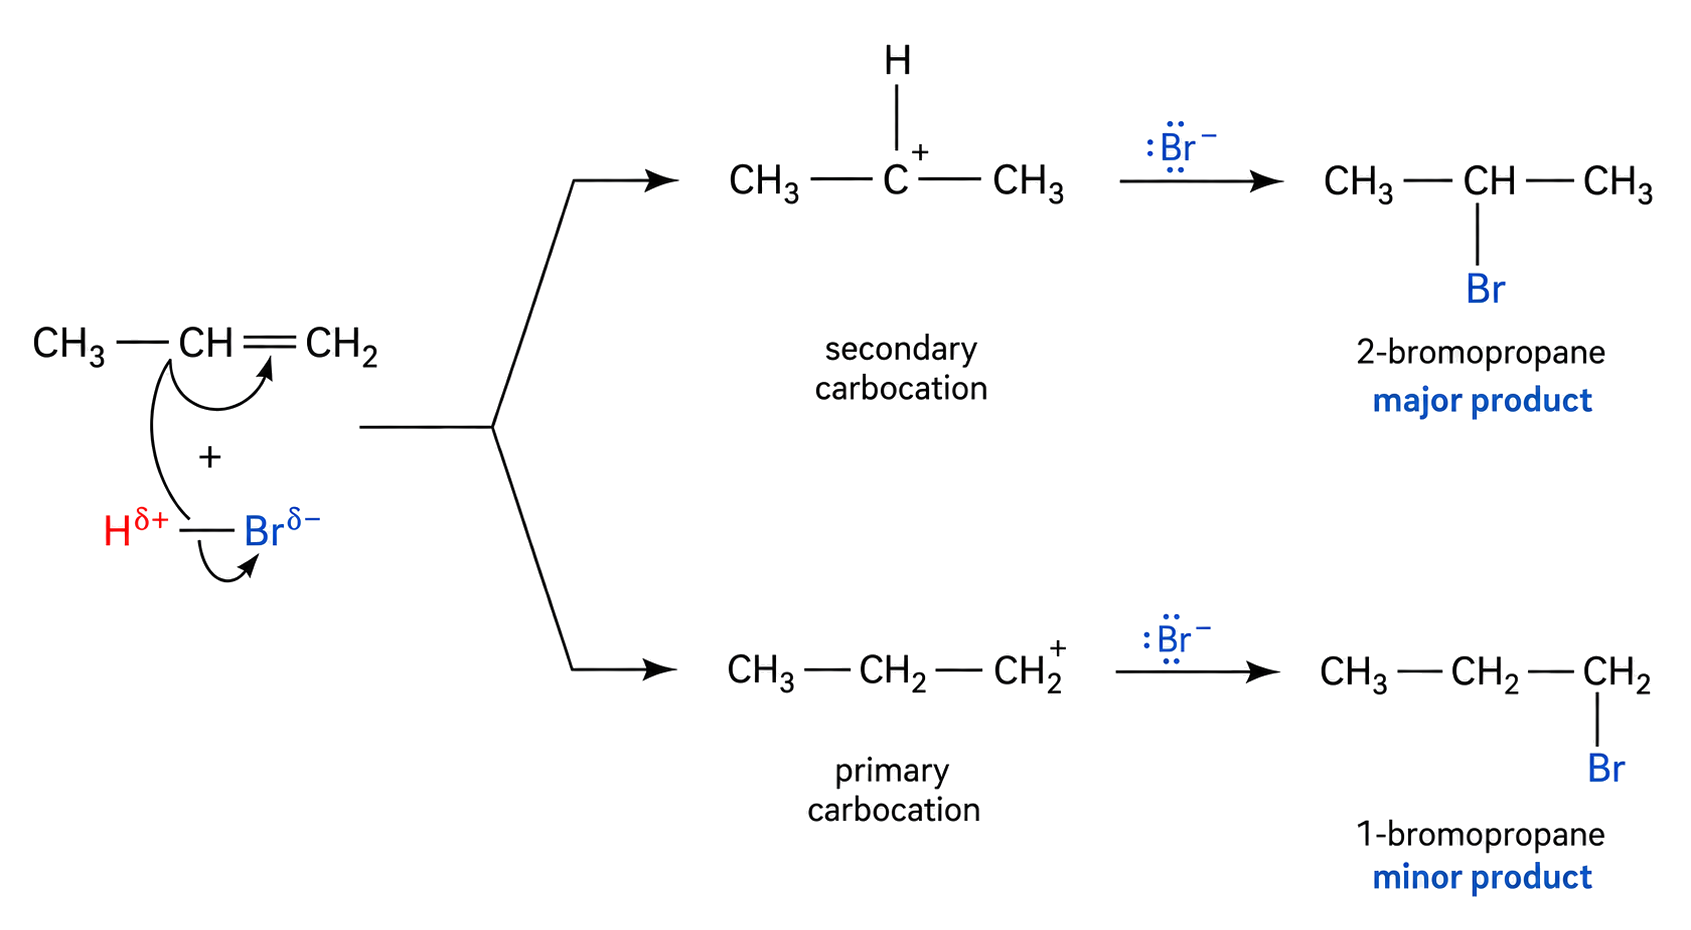
the upper route forms a valid secondary carbocation with one H on C+; the lower route forms a primary carbocation; 2-bromopropane is correctly labelled major and 1-bromopropane minor; three carbons are conserved in every structure.

For an unsymmetrical alkene, draw both protonation routes and compare the carbocations: tertiary is generally more stable than secondary, then primary. Choose the product from the more stable intermediate; do not apply this shortcut to a halonium pathway.

Predicting the Major Addition Product

HL only

[ 2 ]

Explain which of the two structural isomers is the major product.

Electrophilic Substitution of Benzene

HL only

Benzene attacks the charged electrophile E+, forms a charged intermediate, then loses H+ to restore aromaticity. The formation of E+ is outside this assessed mechanism boundary.
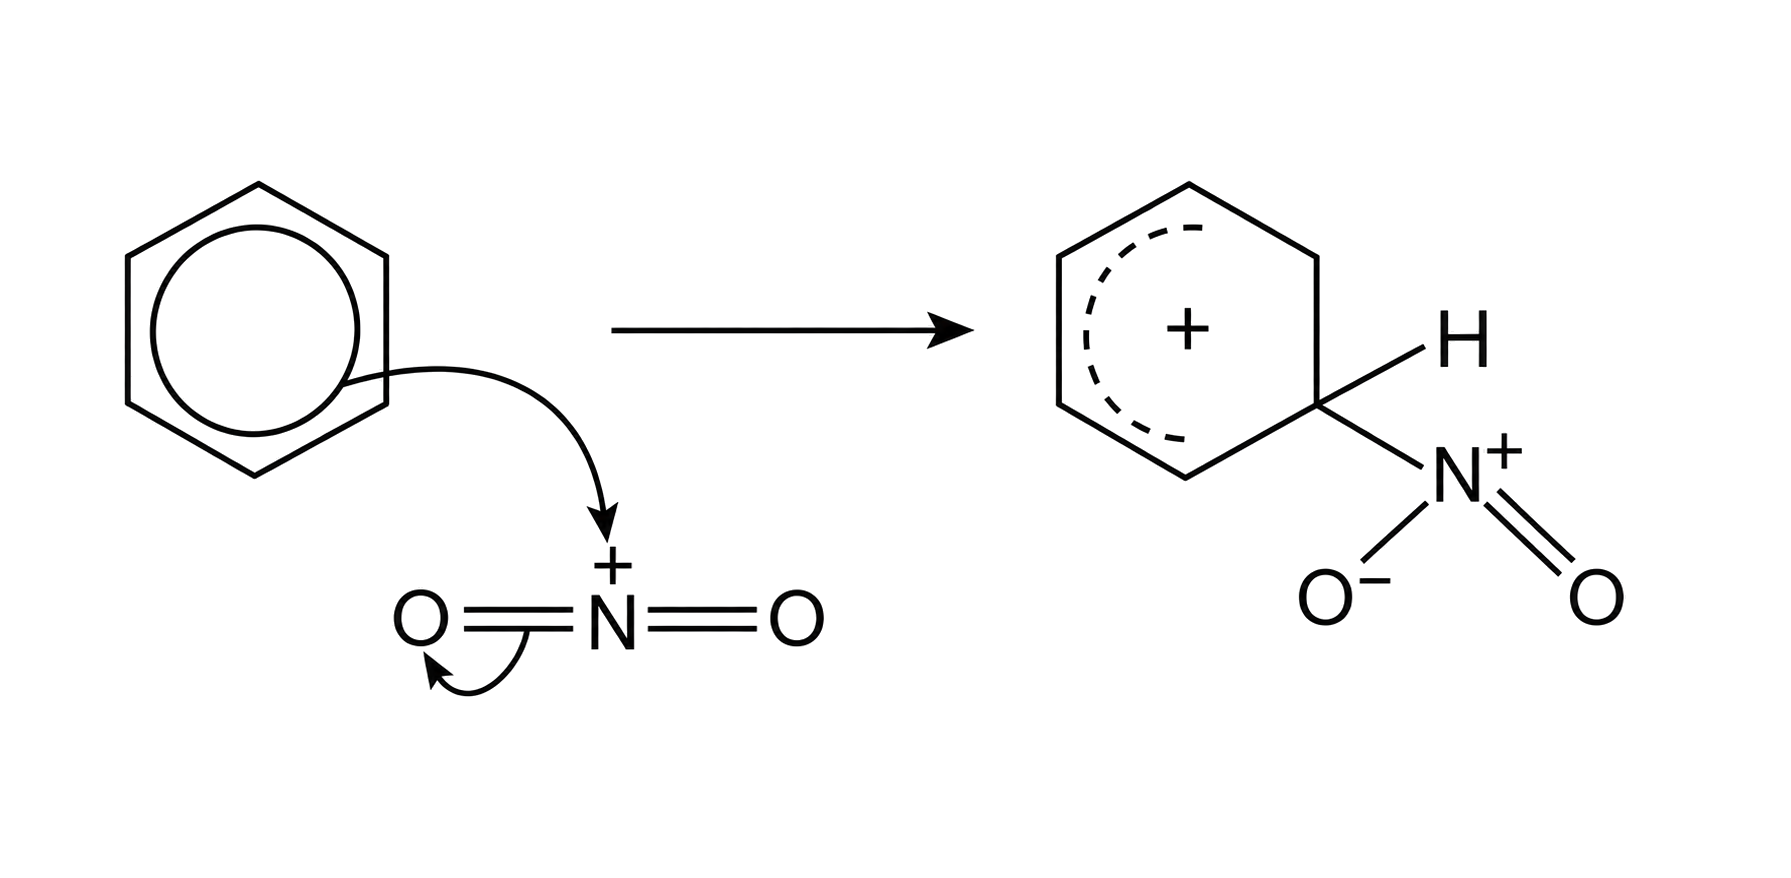
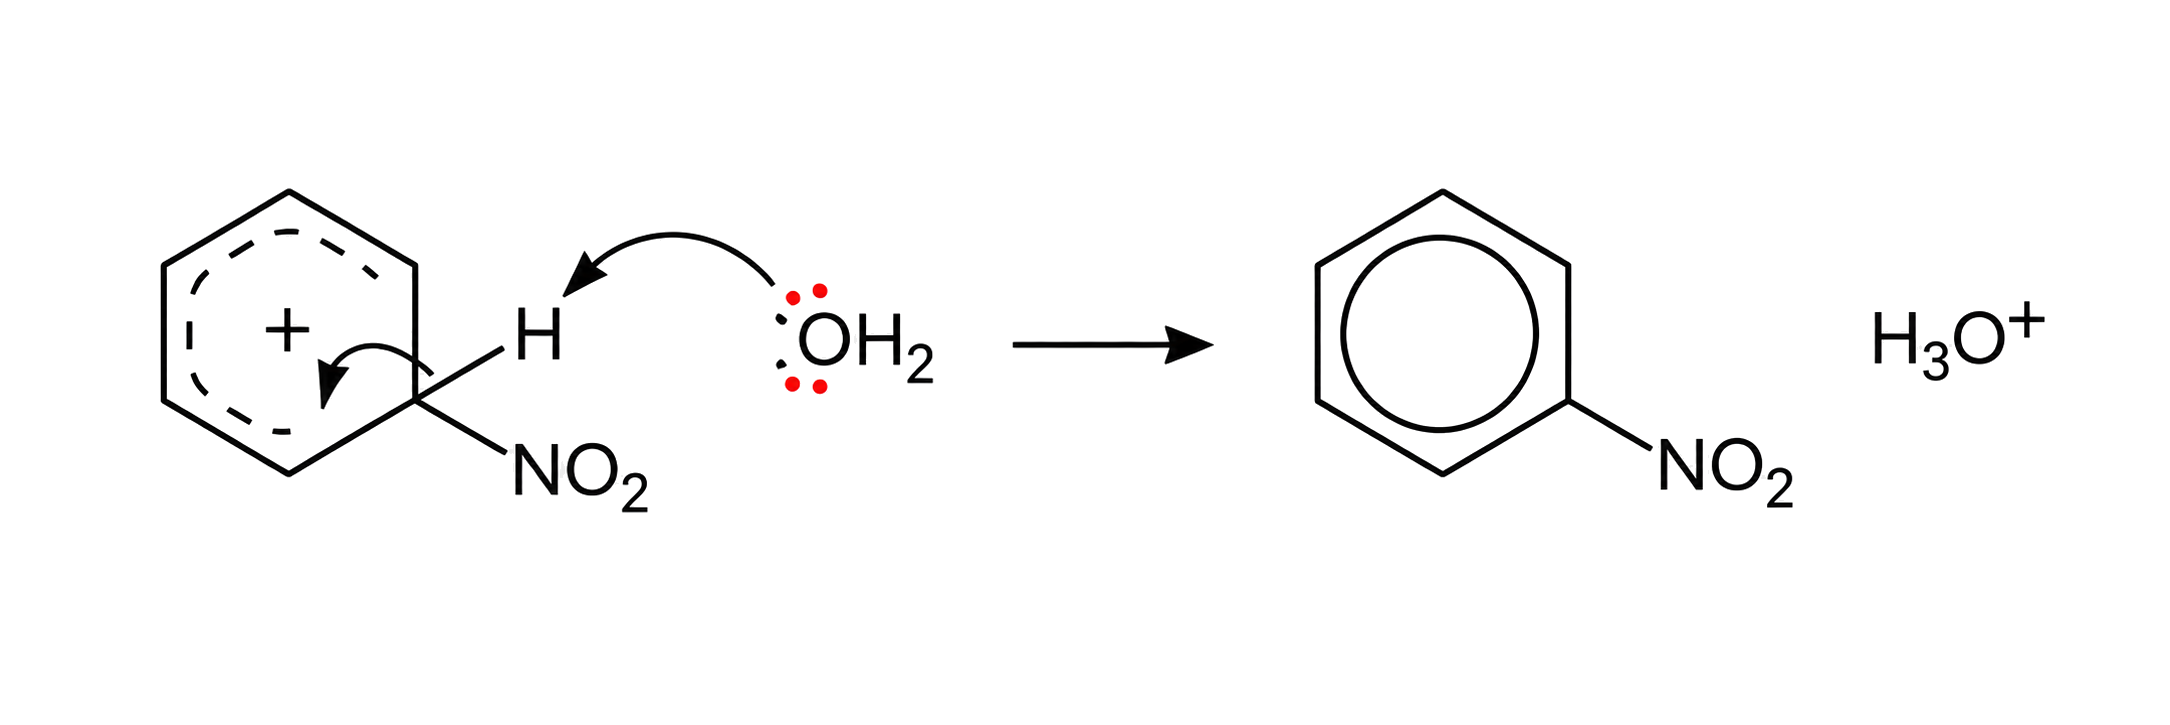
the attacking electron-pair arrow starts at the benzene pi system and ends at N+ of NO2+; the N=O pi-bond arrow ends on oxygen; the sigma-complex carbon bears both H and NO2; the incomplete dashed arc and ring plus sign show aromaticity loss and delocalized positive charge.
a water lone pair attacks the hydrogen on the sigma-complex carbon; the C–H bond electron pair returns to the ring; nitrobenzene with a restored aromatic circle and H3O+ are formed; NO2 remains attached to the same ring carbon.

The first step temporarily disrupts aromatic delocalization; loss of H⁺ then restores the ring and replaces H by E. An addition product would leave aromaticity broken, which explains why the assessed outcome is substitution.

Drawing Benzene Electrophilic Substitution

HL only

[ 4 ]

Explain the mechanism for the nitration of benzene, using curly arrows to indicate the movement of electron pairs.

Electron-Pair Sharing Summary

Retrieve the route: classify nucleophiles and electrophiles, show heterolysis, write substitution and addition mechanisms, map Lewis coordination, compare SN1/SN2, and restore aromaticity in benzene substitution.

Check electron-pair arrow origin and destination, leaving-group departure, intermediate identity, carbocation stability and the assessed mechanism boundary.