2.2 Rate of chemical change

Syllabus
First assessment 2025
Topic
2.2
Level
HL

Learning objectives

2.2.1Rate of reaction• Change in concentration per unit time• Determine rates from tangents of concentration, volume, or mass graphs2.2.2Collision theory• Sufficient energy + proper orientation required• Kinetic energy and temperature (Kelvin)• Explain successful collisions using activation energy and orientation2.2.3Factors affecting rate• Pressure, concentration, surface area, temperature, catalyst• Predict and explain rate changes when conditions change2.2.4Activation energy (Ea)• Minimum energy for successful collision• Maxwell-Boltzmann distribution curves• Use Maxwell-Boltzmann curves to show temperature effects on successful collisions2.2.5Catalysts• Provide alternative pathway with lower Ea• Energy profiles with/without catalysts• Explain catalyst effects on energy profiles and Maxwell-Boltzmann distributions2.2.6(HL)—Reaction mechanisms• Series of elementary steps• Slowest step = rate-determining• Intermediates vs. transition states• Evaluate proposed mechanisms against kinetic and stoichiometric data2.2.7(HL)—Energy profiles• Show Ea and transition state of rate-determining step• Construct and interpret multistep energy profiles from kinetic data2.2.8(HL)—Molecularity• Number of particles in elementary step• Unimolecular, bimolecular, termolecular2.2.9(HL)—Rate equations• Determined experimentally, depend on mechanism• Deduce rate equations from experimental data2.2.10(HL)—Reaction order• Exponent in rate equation• Zero, first, second order• Overall order = sum of individual orders• Analyse concentration-time and rate-concentration graphs2.2.11(HL)—Rate constant (k)• Temperature dependent• Units from overall order• Solve rate equation problems, including units of k2.2.12(HL)—Arrhenius equation• Temperature dependence of k determines Ea• Linear form: ln k vs. 1/T2.2.13(HL)—Arrhenius factor (A)• Frequency of proper collision orientations• Determine activation energy and Arrhenius factor from data

Reaction Rate

rate=changeinconcentration/timerate = change in concentration / time

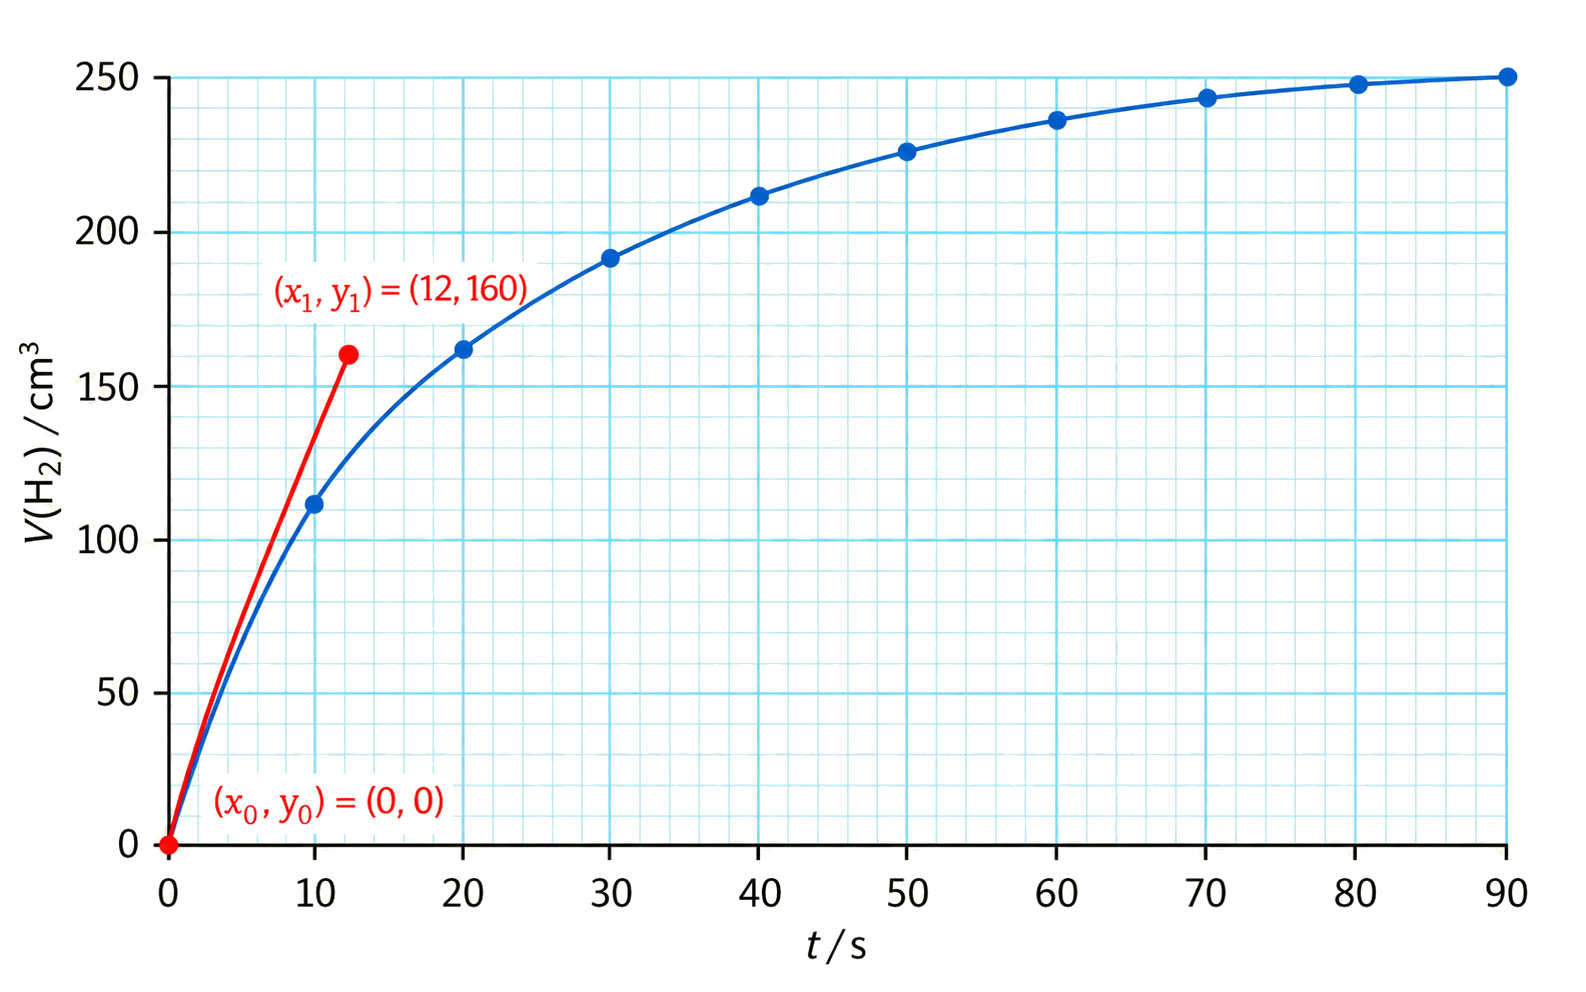
time in seconds is the horizontal variable and hydrogen volume in cubic centimetres is vertical; the product-volume curve rises rapidly then approaches a plateau; the initial tangent passes through (0,0) and (12,160); the tangent gradient represents initial rate without being confused with the curved data trace.

An instantaneous rate is the gradient of a tangent at the stated point on a concentration–time, volume–time or mass–time graph. Keep the units consistent.

Choose two well-separated points on the tangent, not on the curve, to calculate its gradient. A reactant concentration has a negative gradient, so report its disappearance rate as a positive magnitude unless a signed change is requested.

Requested rate Graph operation Evidence check
mean over an interval secant gradient between interval endpoints quote the interval and units
initial tangent gradient at t = 0 choose well-separated points on the tangent
instantaneous at time t tangent gradient at that time do not use two points on the curved trace

A measured mass, pressure or gas volume is a rate proxy only when its change is tied to reaction progress under the stated conditions. Preserve reactant/product slope sign or report a positive disappearance/formation magnitude as requested.

Worked tangent example: on a concentration–time graph for HCl\ce{HCl}, two points on the tangent at t=0t=0 are (0 s,0.250 mol dm−3)(0\,\mathrm{s},0.250\,\mathrm{mol\,dm^{-3}}) and (14 s,0.100 mol dm−3)(14\,\mathrm{s},0.100\,\mathrm{mol\,dm^{-3}}). The tangent gradient is (0.100−0.250)/(14−0)=−0.0107 mol dm−3 s−1(0.100-0.250)/(14-0)=-0.0107\,\mathrm{mol\,dm^{-3}\,s^{-1}}. For Mg+2 HCl→MgClX2+HX2\ce{Mg + 2HCl -> MgCl2 + H2}, divide the positive disappearance-rate magnitude by the HCl coefficient: v=0.0107/2=0.0054 mol dm−3 s−1v=0.0107/2=0.0054\,\mathrm{mol\,dm^{-3}\,s^{-1}}.

Finding an Instantaneous Rate

3 marks

Determine the instantaneous rate of reaction to two significant figures when [Br2]=0.0080 moldm−3\left[\mathrm{Br}_{2}\right]=0.0080 \mathrm{~mol} \mathrm{dm}^{-3}.

Collision Theory

A successful collision needs sufficient kinetic energy to overcome activation energy and a suitable orientation of the reacting particles.
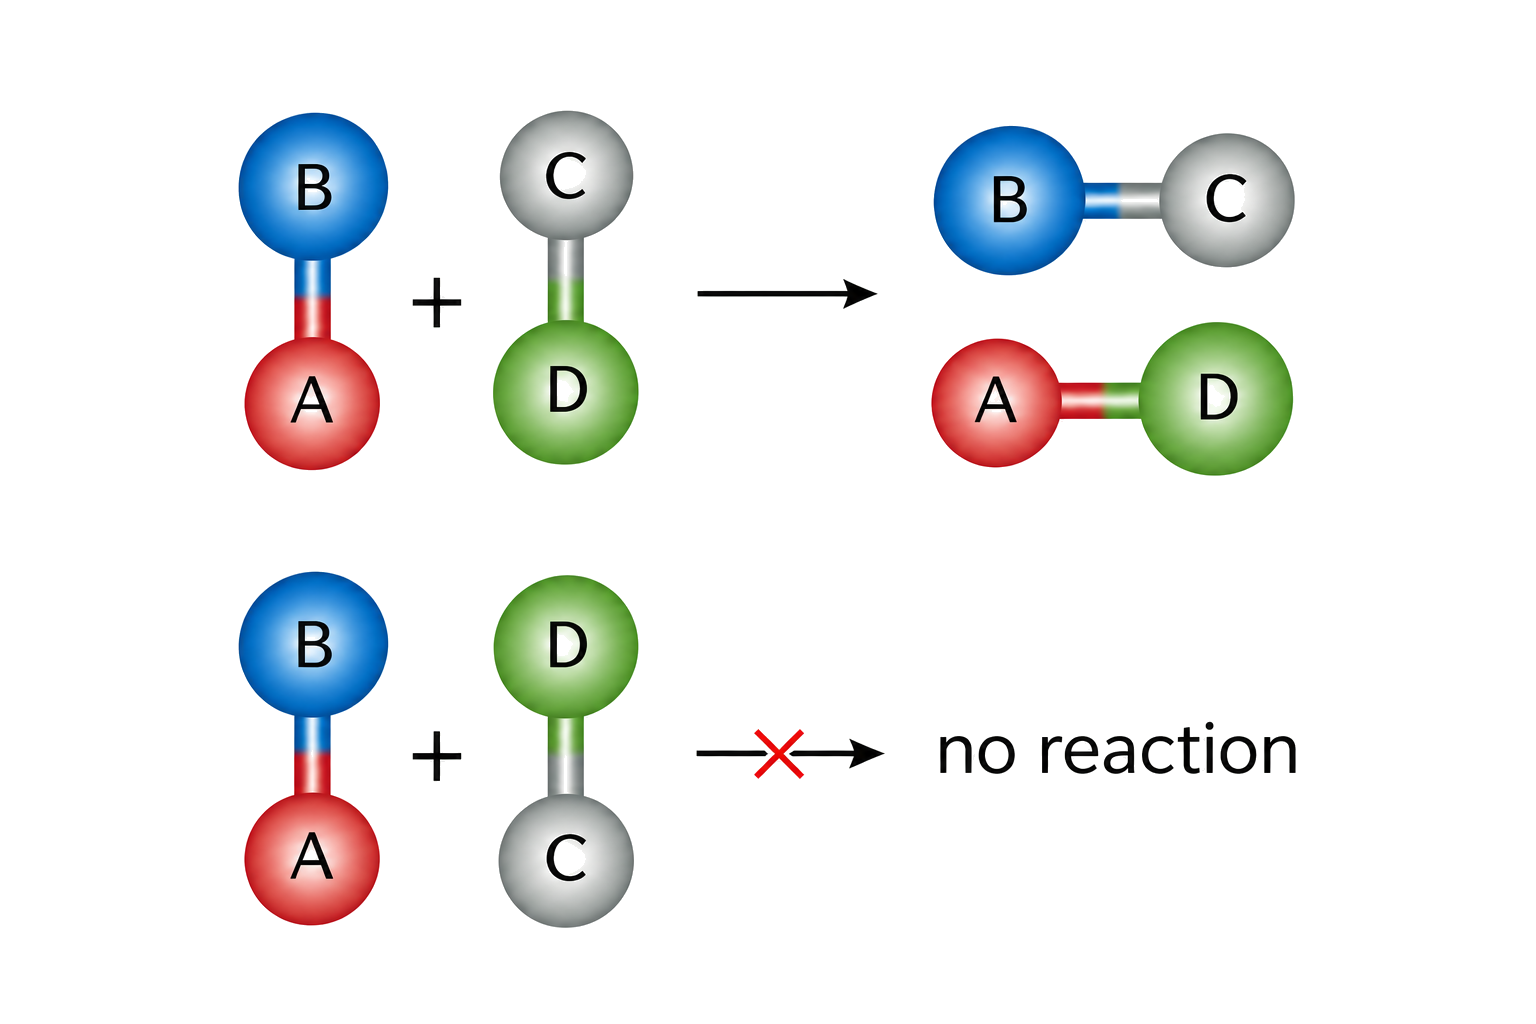
favourable AB plus CD orientation places A facing D and forms AD plus BC; unfavourable AB plus DC orientation places A facing C and gives no reaction; A, B, C and D are each conserved exactly once; the contrast teaches orientation rather than a change in collision energy.

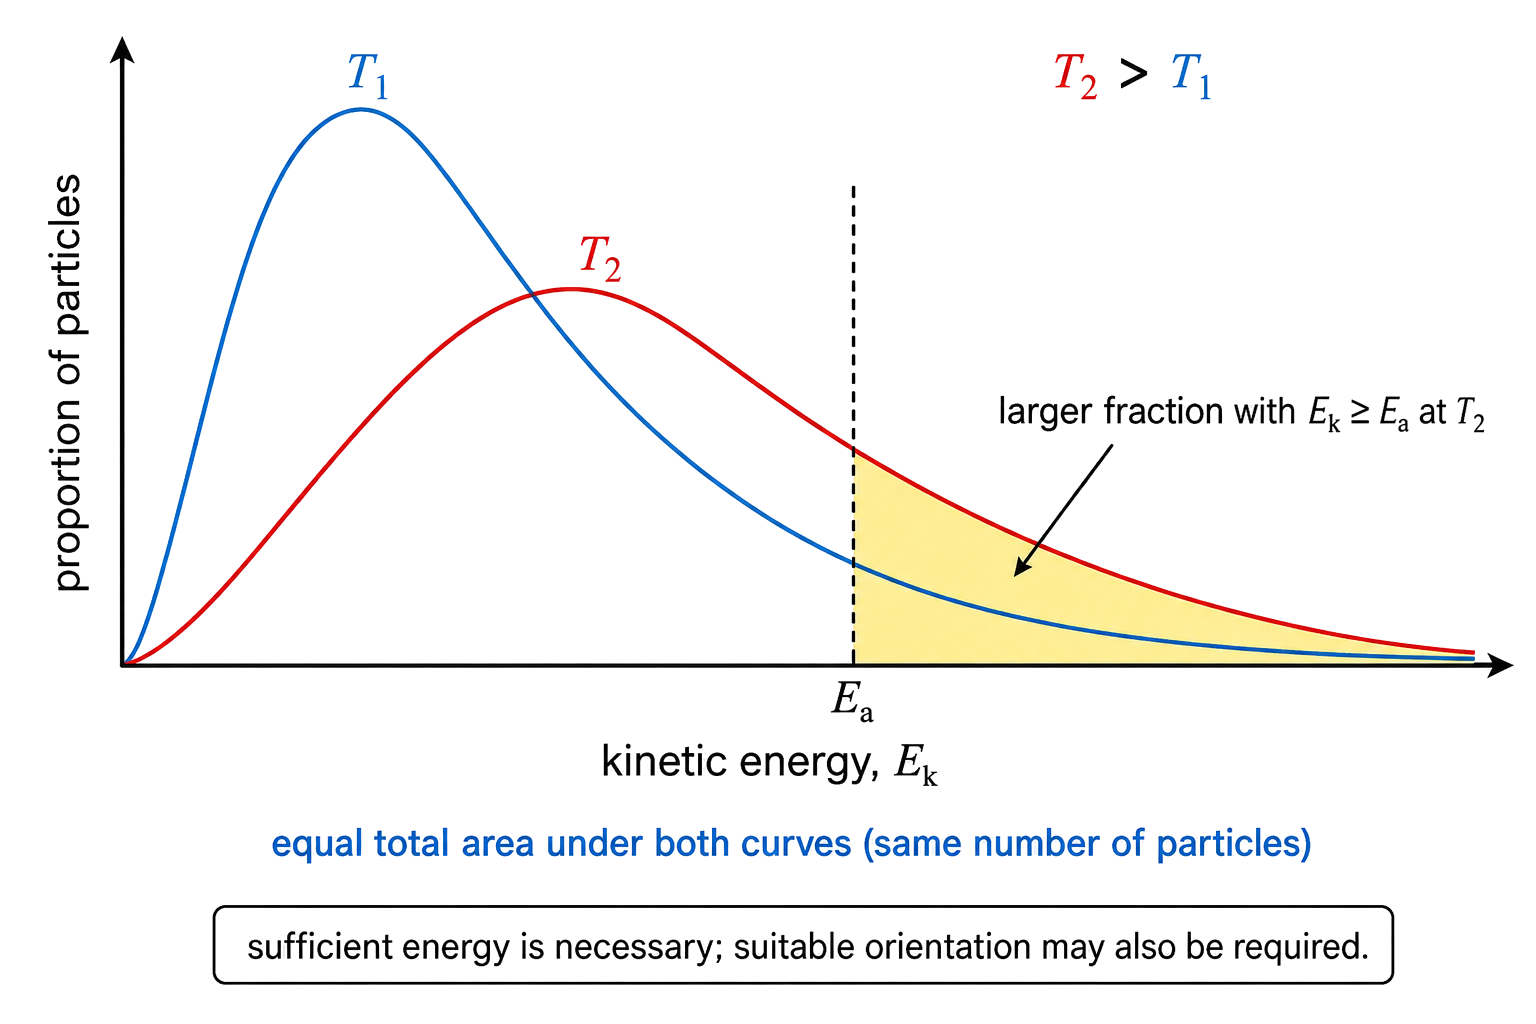
Temperature raises average kinetic energy and changes collision frequency and the fraction of particles with energy at least Ea. Collision frequency alone does not guarantee reaction.

At one temperature, only the fraction of collisions above Ea and with a productive orientation can react. Raising temperature increases that fraction, not the energy of every particle by the same amount. Use collision frequency to explain concentration or pressure effects and the energy distribution to explain the stronger temperature effect.
Explaining Rate with Collision Theory

3 marks

Explain, using collision theory, how an increase in temperature increases the reaction rate.

Factors Affecting Rate

Change Main collision consequence
concentration or pressure up more frequent collisions
surface area up more collisions at a solid surface
temperature up more frequent and more energetic collisions
catalyst alternative lower-Ea pathway

Explain a predicted rate change through collision frequency or the fraction of effective collisions, not just by saying particles move faster.

Powdered CaCO₃ reacts faster than equal-mass chips because more surface sites are exposed, not because its particles have higher kinetic energy. For each changed condition, identify exactly what changes—collision frequency, energy distribution or pathway—and hold other variables constant in a fair comparison.

Predicting Rate Changes

2 marks

The student then carried out the experiment at other acid concentrations with all other conditions remaining unchanged.

[H+]/ mol dm−3\left[\mathbf{H}^{+}\right] / \mathbf{~ m o l ~ d m}^{-\mathbf{3}}Relative rate of reaction
0.050.0025
0.100.0051
0.200.0100

State and explain the relationship between the rate of reaction and the concentration of acid.

Activation Energy and Maxwell–Boltzmann Curves

Ea=minimumkineticenergyforaneffectivecollisionEa = minimum kinetic energy for an effective collision

T2 is lower, broader, and peaks to the right of T1; both curves are stated to have equal total area for the same particle count; Ea is unchanged by temperature; the fraction with Ek at least Ea is larger at T2 and orientation remains a separate requirement.

A Maxwell–Boltzmann curve shows the distribution of particle kinetic energies. At higher temperature the peak is lower and shifts right; the area beyond Ea is larger, so more particles can react.

Two Maxwell–Boltzmann curves for the same number of particles have equal total area. At higher T the curve is broader with a lower peak and a larger area to the right of a fixed Ea line; the peak does not move to Ea and no particle count is lost.

Interpreting Maxwell–Boltzmann Distributions

3 marks

Explain why the reaction rate increases with temperature, adding annotations to the following Maxwell-Boltzmann graph to assist your explanation.

Explanation:

Catalysts and Activation Energy

A catalyst provides an alternative reaction pathway with lower activation energy. It does not change the energy levels of the reactants or products.
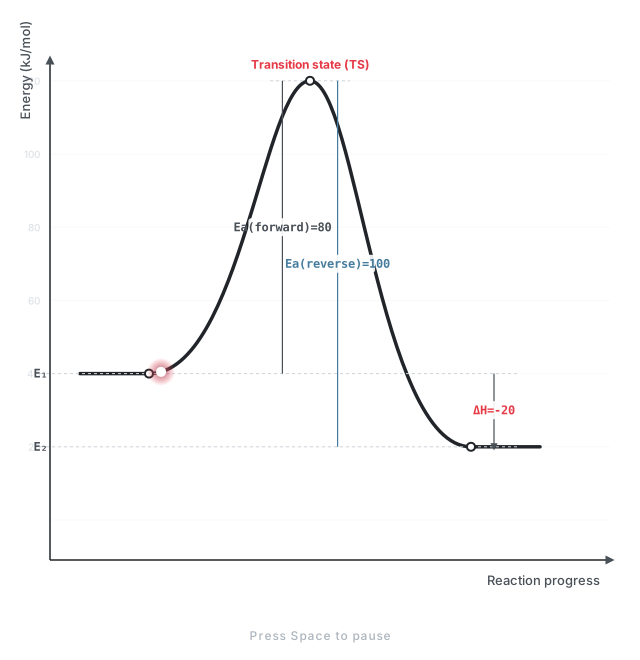
Catalysed and uncatalysed profiles have identical reactant and product energies, but the catalysed pathway has a lower activation barrier.

Because the catalysed Ea is lower, a larger fraction of the same distribution lies beyond the threshold. The reaction therefore has more effective collisions at the same temperature.

At fixed temperature a catalyst does not change the Maxwell–Boltzmann distribution; it moves the threshold to a lower Ea, increasing the area beyond it. On an energy profile it changes the pathway and peak, not ΔH, reactant/product energies or the equilibrium constant.

Showing a Catalyst on Energy Diagrams

2 marks

Sketch the Maxwell-Boltzmann energy distribution curve for this reaction. Label the activation energy with and without a catalyst on the diagram.

Reaction Mechanisms

HL only

A mechanism is a sequence of elementary steps. The slowest step is the rate-determining step; an intermediate is formed in one step and consumed in a later step, whereas a transition state is the high-energy configuration at a barrier.
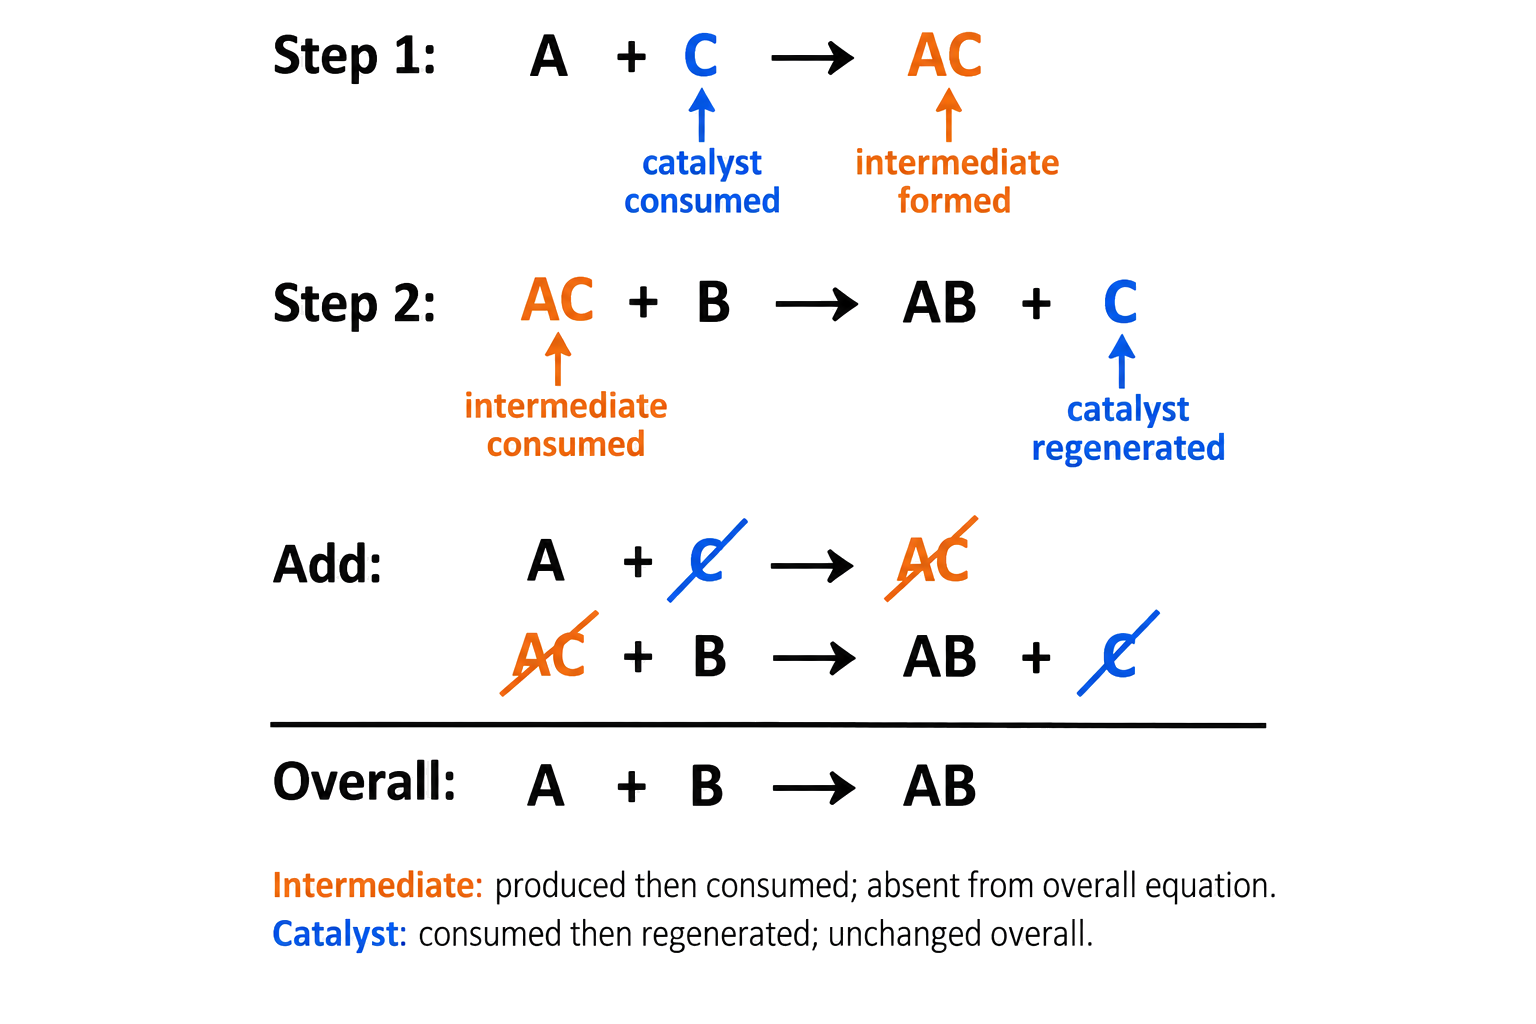
AC is produced in step 1 and consumed in step 2, so it is an intermediate; C is consumed in step 1 and regenerated in step 2, so it is a catalyst; AC and C cancel when elementary steps are added; overall equation is A + B to AB.

A proposed mechanism must be compared with the experimental rate equation and stoichiometry. Matching a rate equation can support a mechanism but does not prove it, because different mechanisms may give the same expression.

Add all elementary steps and cancel intermediates to recover the overall equation. Then derive the rate dependence expected from the slow step, eliminating an intermediate when necessary, and compare with experiment. A catalyst consumed early and regenerated later cancels from the overall equation but is not an intermediate.

Evaluating a Proposed Mechanism

HL only

1 mark

Suggest why experimental confirmation of the rate equation would not prove that the mechanism is correct.

Multistep Energy Profiles

HL only

In a multistep profile, peaks are transition states and valleys between peaks are intermediates. Each step has its own Ea; the largest barrier from its preceding intermediate identifies the rate-determining step.
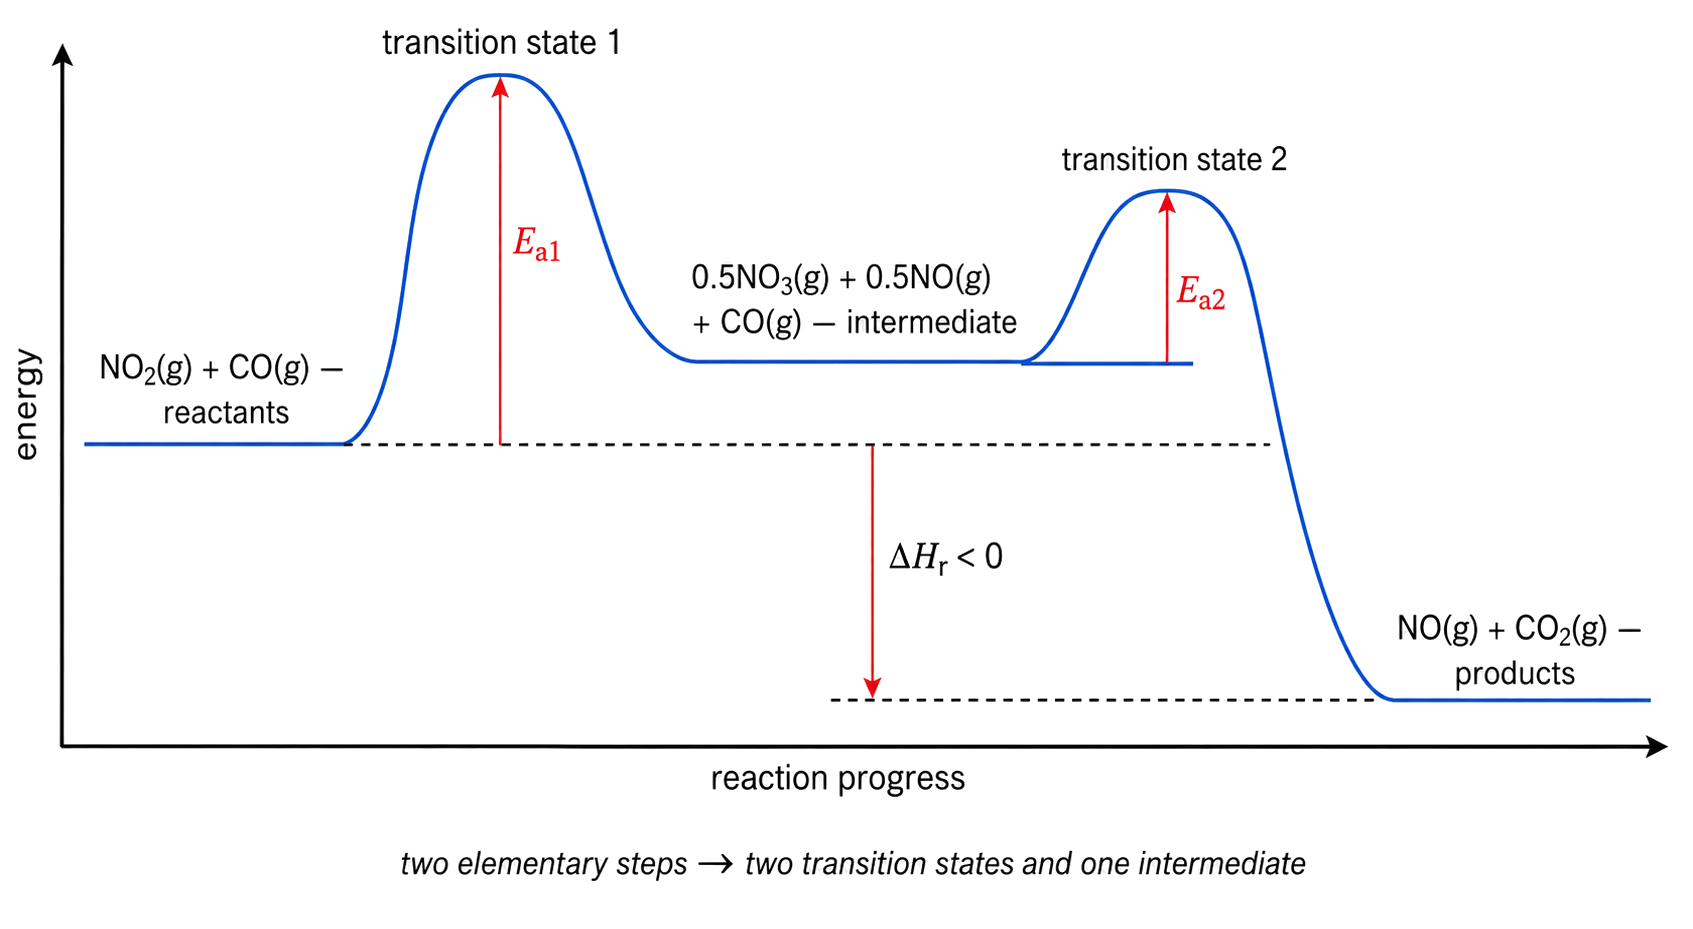
the specified reactants, intermediate, and products are preserved; two elementary steps correspond to two transition-state maxima and one intermediate minimum; Ea1 starts at reactants while Ea2 starts at the intermediate; Ea1 exceeds Ea2 and products are lower with ΔHr negative.

ΔH=energy(products)−energy(reactants)ΔH = energy(products) − energy(reactants)

For each step, measure Ea upward from its own preceding reactant or intermediate valley to the next peak. The rate-determining barrier is the largest of those step barriers, which need not be the peak with the greatest absolute height above the page baseline.
Constructing a Two-Step Energy Profile

HL only

4 marks

Sketch an energy profile for the two-step reaction, labelling reactants, intermediate and products, activation energies, EaE_{\mathrm{a}}, and overall enthalpy change, ΔH\Delta H. Assume that the reaction is exothermic.

Molecularity of an Elementary Step

HL only

Molecularity is the number of reacting particles in one elementary step: one is unimolecular, two is bimolecular and three is termolecular.

Count particles in the individual elementary step, not coefficients in the overall reaction equation.

A step A + 2B → products is termolecular because three reacting particles meet in that elementary event. Molecularity is always a positive whole-number description of one proposed step; reaction order is experimental and can be zero, fractional or unrelated to overall stoichiometric coefficients.

Identifying Molecularity

HL only

1 mark

Identify the molecularity of the rate-determining step in this reaction.

Deducing Rate Equations

HL only

rate=k[A]x[B]yrate = k[A]^x[B]^y

Use experimental trials that vary one concentration independently. Compare the rate factor with the concentration factor to infer each exponent; the balanced overall equation alone cannot determine the rate equation.

If doubling [A] while holding [B] constant quadruples rate, the order in A is 2; if doubling [B] leaves rate unchanged, the order in B is 0, giving rate = k[A]². Choose trial pairs with only one changed concentration before combining exponents.

Worked initial-rate deduction: doubling [FeX3+][\ce{Fe^{3+}}] from 1.00×10−21.00\times10^{-2} to 2.00×10⁻² mol dm−32.00\times10^{-2}\,\mathrm{mol\,dm^{-3}} at constant [IX−][\ce{I^-}] doubles rate, so the order in FeX3+\ce{Fe^{3+}} is 1. Doubling [IX−][\ce{I^-}] at constant [FeX3+][\ce{Fe^{3+}}] increases rate from 3.24×10−53.24\times10^{-5} to 1.30×10⁻⁴ mol dm−3 s−11.30\times10^{-4}\,\mathrm{mol\,dm^{-3}\,s^{-1}}, approximately fourfold, so its order is 2. Therefore v=k[FeX3+][IX−]2v=k[\ce{Fe^{3+}}][\ce{I^-}]^2, third order overall.

Using Experimental Rate Data

HL only

2 marks

Two more trials ( 2 and 3 ) were carried out. The results are given below.

TrialsVolume of 0.20 mol dm−3KI(aq)/cm3\mathbf{0 . 2 0 ~ m o l ~ d m}{ }^{\mathbf{- 3}} \mathbf{K I} \boldsymbol{(} \mathbf{a q} \boldsymbol{)} \boldsymbol{/} \mathbf{c m}^{\mathbf{3}}
Volume of
0.20 mol dm−3\mathbf{0 . 2 0 ~ m o l ~ d m}{ }^{\mathbf{- 3}}KI(aq)/cm3\mathbf{K I} \boldsymbol{(} \mathbf{a q} \boldsymbol{)} \boldsymbol{/} \mathbf{c m}^{\mathbf{3}}
Volume of
deionised
water / cm3\mathbf{c m}^{\mathbf{3}}
Volume of 3\% H2O2(aq)/cm3\mathrm{H}_{2} \mathrm{O}_{2}(\mathrm{aq}) / \mathbf{c m}^{3}
Volume of 3\%
H2O2(aq)\mathrm{H}_{2} \mathrm{O}_{2}(\mathrm{aq})/cm3/ \mathbf{c m}^{3}
Average rate of
reaction
/cm3O2(g)s−1/ \mathrm{cm}^{3} \mathbf{O}_{\mathbf{2}}(\mathrm{g}) \mathbf{s}^{-1}
110.015.05.0
210.010.010.00.0429
320.05.05.00.0451

Determine the rate equation for the reaction and its overall order, using your answer from (b)(i).

Rate equation:

Overall order:

Reaction Order

HL only

overallorder=x+yinrate=k[A]x[B]yoverall order = x + y in rate = k[A]^x[B]^y

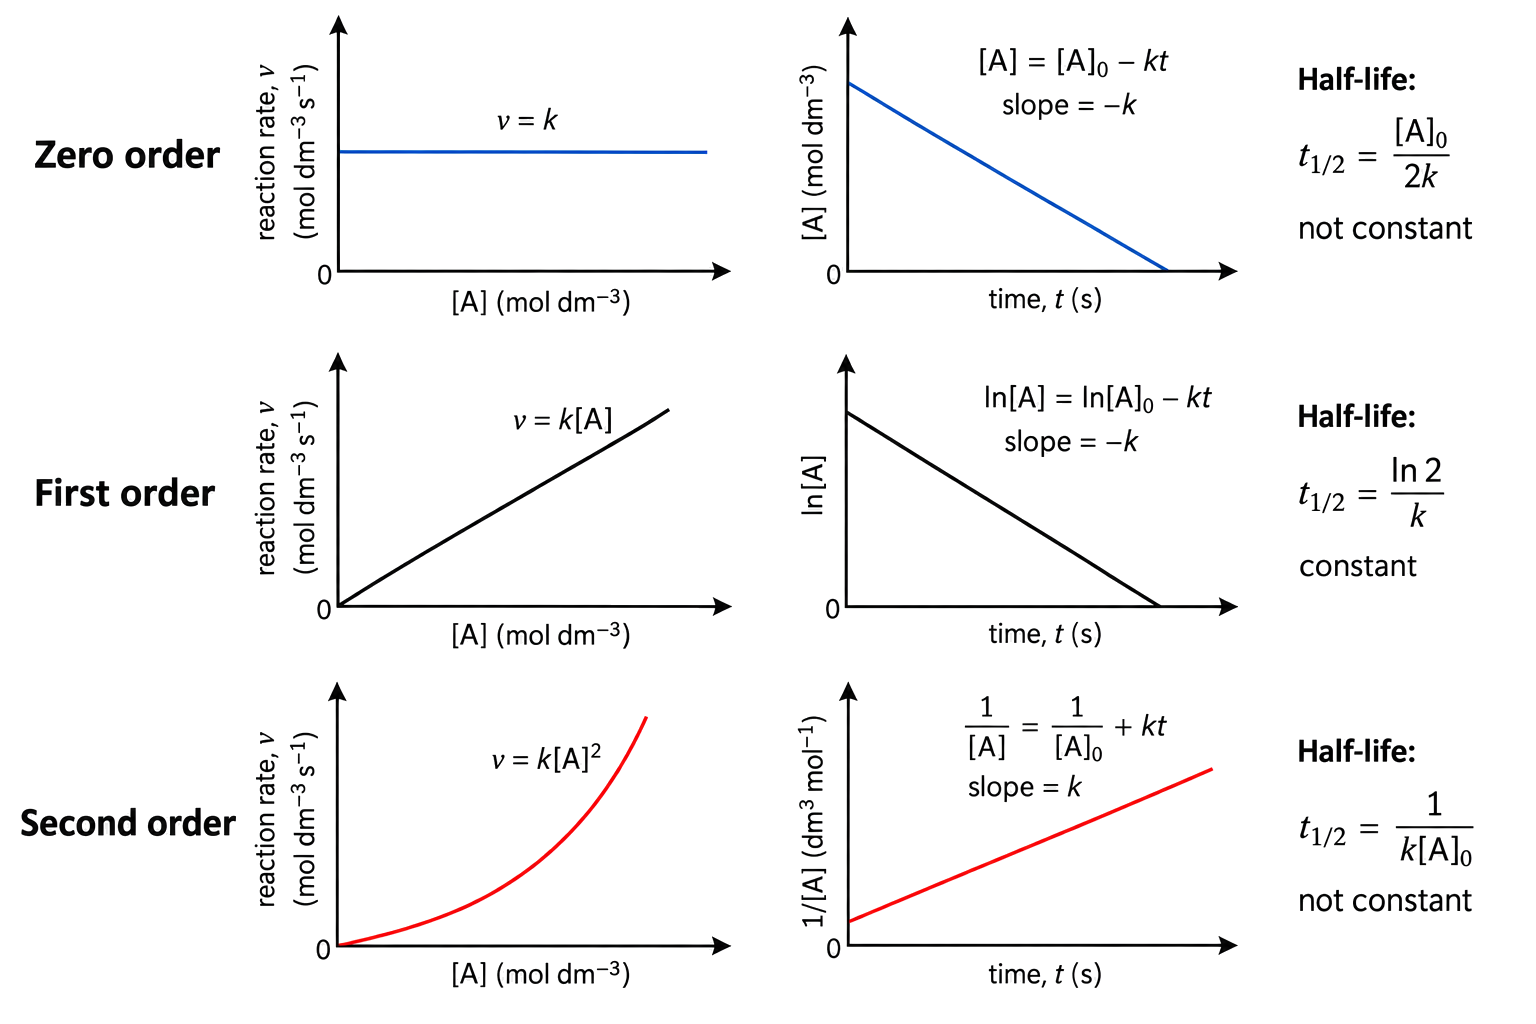
rate-concentration plots are horizontal, linear through origin and parabolic for zero, first and second order; diagnostic linear plots use [A], ln[A] and 1/[A] respectively; slopes are -k, -k and +k respectively; half-life expressions and concentration dependence are correct.

The exponent gives order with respect to that reactant. Use rate factors to identify zero, first or second order, then match the order to the concentration–time or rate–concentration graph.

Use multiple representations as cross-checks: zero-order rate is independent of concentration and [A] falls linearly; first-order rate is proportional to [A] and gives exponential decay; second-order rate curves upward on a rate-versus-concentration plot. Do not infer order from one balanced equation.
Determining Overall Order

HL only

1 mark

Compounds P and Q were mixed together at various concentrations and the initial rate of each reaction was measured.

Experiment[P] mol dm −3{ }^{-3}[Q] mol dm −3{ }^{-3}Initial rate of reaction
(mol dm −3 s−1{ }^{-3} \mathrm{~s}^{-1} )
10.200.150.50
20.100.150.25
30.200.301.00

What are the orders of reaction with respect to P and Q ?

Order with respect to P

Order with respect to Q

1

1

1

2

2

1

1

0

The Rate Constant k

HL only

k=rate/([A]x[B]y)k = rate / ([A]^x[B]^y)

The units of k depend on the overall order and must cancel the concentration and time units in the rate equation. For a particular reaction, k changes with temperature.

Derive rather than memorize the units: first-order k has units s⁻¹, while an overall second-order law written with mol dm⁻³ and seconds gives dm³ mol⁻¹ s⁻¹. Concentration changes rate but does not change k at fixed temperature.

Worked kk example: for v=k[FeX3+][IX−]2v=k[\ce{Fe^{3+}}][\ce{I^-}]^2, use v=1.62×10⁻⁵ mol dm−3 s−1v=1.62\times10^{-5}\,\mathrm{mol\,dm^{-3}\,s^{-1}} and both concentrations 1.00×10⁻² mol dm−31.00\times10^{-2}\,\mathrm{mol\,dm^{-3}}. k=1.62×10⁻⁵(1.00×10⁻²)(1.00×10⁻²)2=16.2 dm6 mol−2 s−1k=\frac{1.62\times10^{-5}}{(1.00\times10^{-2})(1.00\times10^{-2})^2}=16.2\,\mathrm{dm^6\,mol^{-2}\,s^{-1}}. Substitution returns the measured rate, and the units cancel those of a third-order concentration term.

Calculating k and Its Units

HL only

2 marks

Calculate the value of the rate constant stating its units.

The Arrhenius Equation

HL only

lnk=(−Ea/R)(1/T)+lnAln k = (−Ea/R)(1/T) + ln A

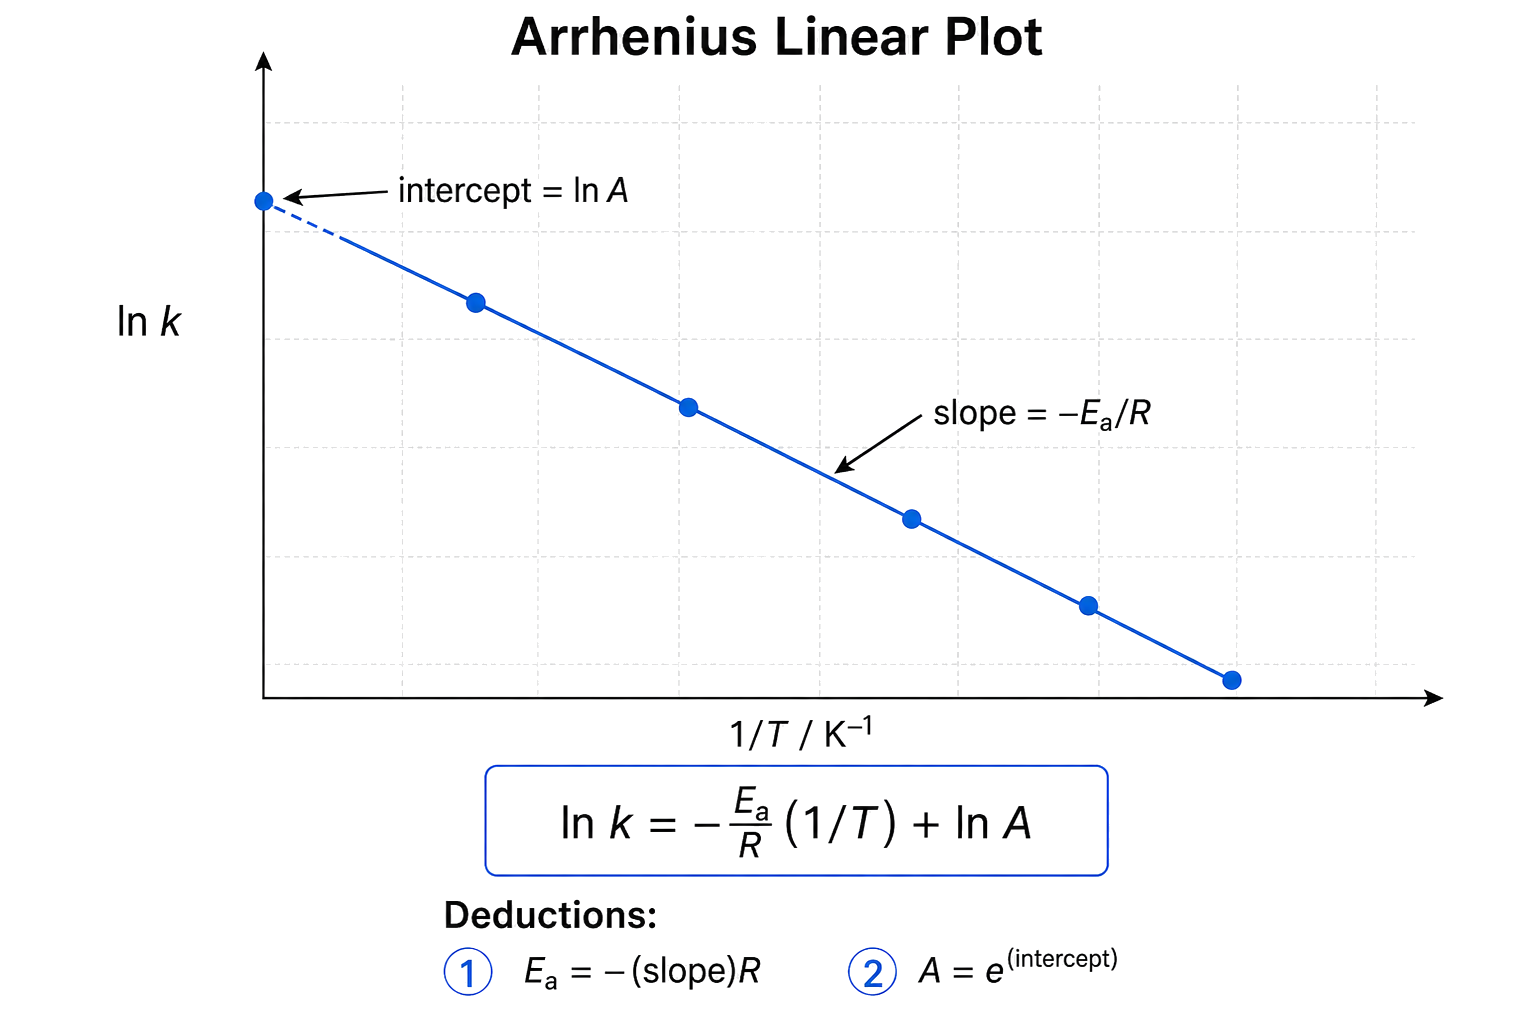
ln k is plotted against reciprocal temperature 1/T in K-1; the best-fit line has negative slope -Ea/R; the intercept is ln A; Ea and A rearrangements preserve the negative sign and exponential relationship.

On a plot of ln k against 1/T, gradient = −Ea/R and intercept = ln A. Use kelvin, preserve the negative gradient and convert J mol−1 to kJ mol−1 when required.

A steeper negative ln k versus 1/T gradient means a larger activation energy. When two temperatures are given, subtracting the two linear equations removes ln A and lets Ea be found without knowing the frequency factor.

ln(k2/k1)=−(Ea/R)(1/T2−1/T1)withTinKandEainJmol−1whenR=8.31Jmol−1K−1ln(k₂/k₁) = −(Eₐ/R)(1/T₂ − 1/T₁) with T in K and Eₐ in J mol⁻¹ when R = 8.31 J mol⁻¹ K⁻¹

Worked Arrhenius-plot example: using line points (0.00302 K−1,−6.8)(0.00302\,\mathrm{K^{-1}},-6.8) and (0.00334 K−1,−10.8)(0.00334\,\mathrm{K^{-1}},-10.8), the gradient is [−6.8−(−10.8)]/(0.00302−0.00334)=−1.25×104 K[-6.8-(-10.8)]/(0.00302-0.00334)=-1.25\times10^4\,\mathrm{K}. Since gradient =−Ea/R=-E_a/R, Ea=−(−1.25×10⁴)(8.31)=1.04×10⁵ J mol−1=104 kJ mol−1E_a=-(-1.25\times10^4)(8.31)=1.04\times10^5\,\mathrm{J\,mol^{-1}}=104\,\mathrm{kJ\,mol^{-1}}. The negative plot gradient therefore produces a positive activation energy.

Finding Ea from an Arrhenius Plot

HL only

2 marks

Determine the activation energy of the reaction, EaE_{\mathrm{a}}, in kJmol−1\mathrm{kJ} \mathrm{mol}^{-1}, from the Arrhenius plot. Use sections 1 and 2 of the data booklet.

The Arrhenius Factor A

HL only

A is the frequency factor: it represents the frequency of collisions with proper orientation. In the linear Arrhenius form, the y-intercept is ln A.

Read the intercept, find A by exponentiating ln A, and keep A distinct from Ea: Ea describes the energy barrier, while A describes collision frequency and orientation.

From a fitted line, intercept b gives A = eᵇ, while gradient m gives Ea = −mR. Check the pair against k = Ae^(−Ea/RT) at one data temperature. A has the same units as k for the stated rate law; it is not an activation energy or a universal constant.

Worked intercept example: for the same first-order Arrhenius line, use ln⁡k=−6.02\ln k=-6.02, 1/T=0.00296 K−11/T=0.00296\,\mathrm{K^{-1}} and gradient −Ea/R=−12500 K-E_a/R=-12500\,\mathrm{K}. From ln⁡k=(−Ea/R)(1/T)+ln⁡A\ln k=(-E_a/R)(1/T)+\ln A, −6.02=(−12500)(0.00296)+ln⁡A-6.02=(-12500)(0.00296)+\ln A, so ln⁡A=30.98\ln A=30.98 and A=e30.98=2.85×10¹³ s−1A=e^{30.98}=2.85\times10^{13}\,\mathrm{s^{-1}}. The unit is s−1\mathrm{s^{-1}} because this reaction is first order and AA has the same units as kk.

Finding A from the Intercept

HL only

2 marks

Calculate the numerical value of A.

Rate and Mechanisms Summary

Retrieve the route: measure a tangent rate, explain effective collisions, map rate factors, read Ea and energy profiles, evaluate mechanisms, determine molecularity and orders, calculate k, then use Arrhenius gradient and intercept for Ea and A.

Check tangent versus average slope, energy versus orientation, barrier labels, intermediate versus transition state, one-variable trial comparisons, order-dependent units, kelvin temperature and the signs of gradient and Ea.