Classification and cladistics use hierarchy, molecular evidence, clades, cladograms, molecular clocks, and domains to organize evolutionary relationships across biodiversity accurately.
Classification is a testable claim about relationships
HL only
Classification does more than file organisms under names. It proposes which organisms belong together, so it should organize observations and support predictions about shared structures, functions and evolutionary history.
A useful classification lets biologists:
use the same names and groups worldwide
compare unfamiliar organisms with known relatives
connect new evidence to an organized body of knowledge
Traditional hierarchy
Evolutionary classification
Places organisms in fixed ranks such as kingdom, phylum, class, order, family, genus and species
Groups organisms by their inferred common ancestry
Rank boundaries can be arbitrary because evolution is continuous
Branches can represent different amounts of evolutionary divergence
Similar appearance may dominate the decision
Morphology is tested against molecular and other evidence
A named group is therefore a biological hypothesis. If new evidence shows that its members do not share the expected ancestry, the classification should change.
Shared function is not always shared ancestry
HL only
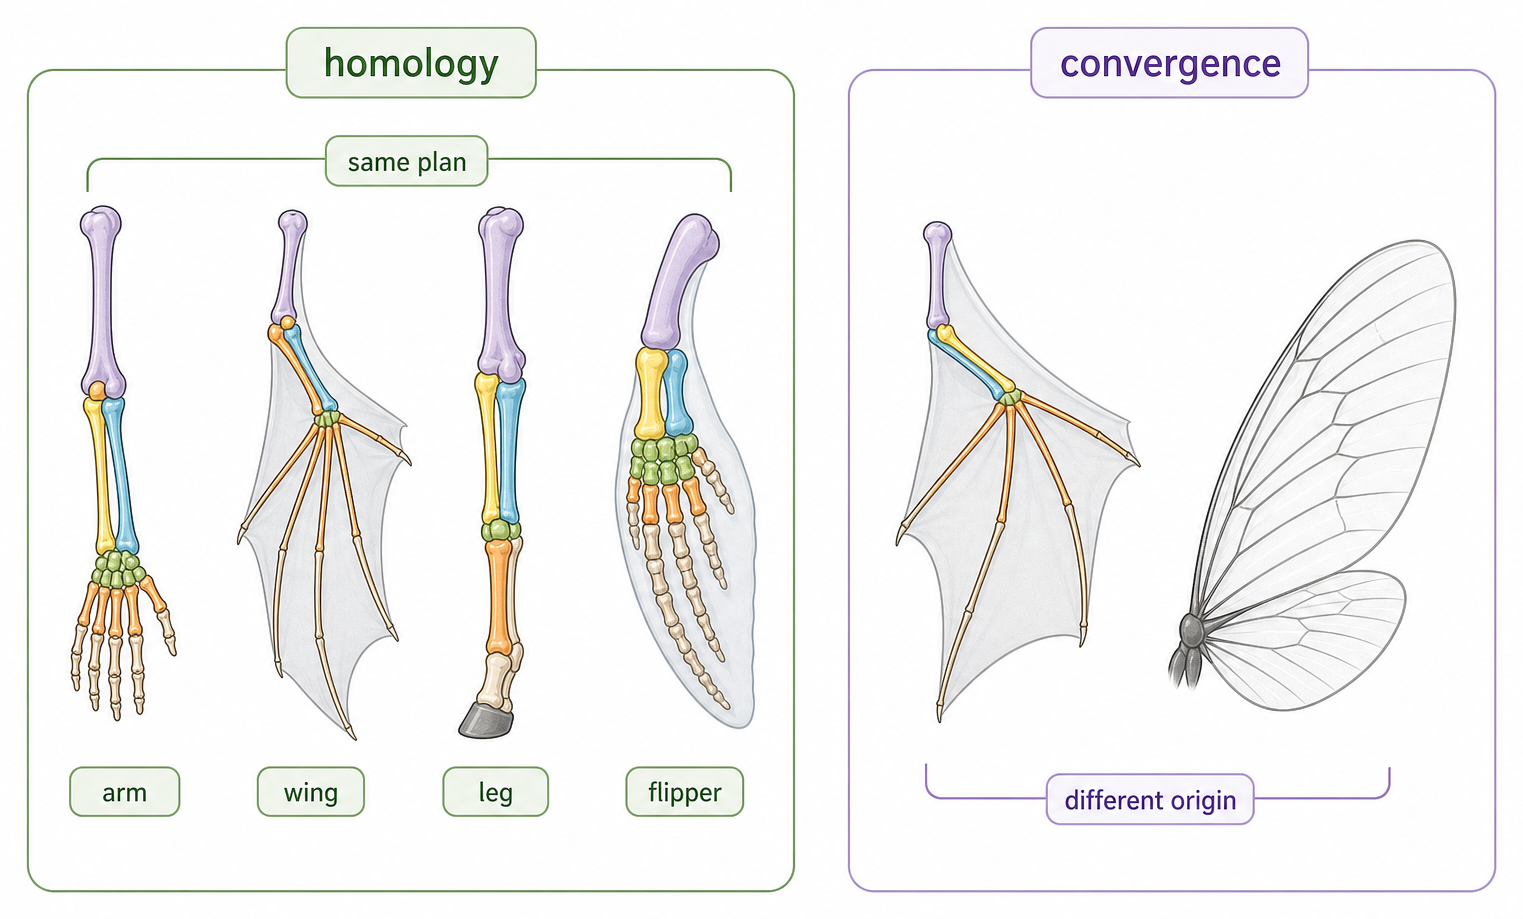
Homologous structures inherit the same underlying feature from a common ancestor, even when later evolution modifies that feature for different functions. Their shared origin can support placement in the same clade.
Analogous structures have similar functions but evolved independently under similar selection pressures. Treating resemblance alone as ancestry can therefore join unrelated lineages.
A clade is monophyletic: it contains one common ancestor and all of its descendants. A shared derived characteristic that helps identify a clade is a synapomorphy.
Possible synapomorphies include:
anatomical or physiological features
behaviour
homologous DNA, RNA or amino-acid sequences
Read relatedness by tracing backwards to a node
HL only
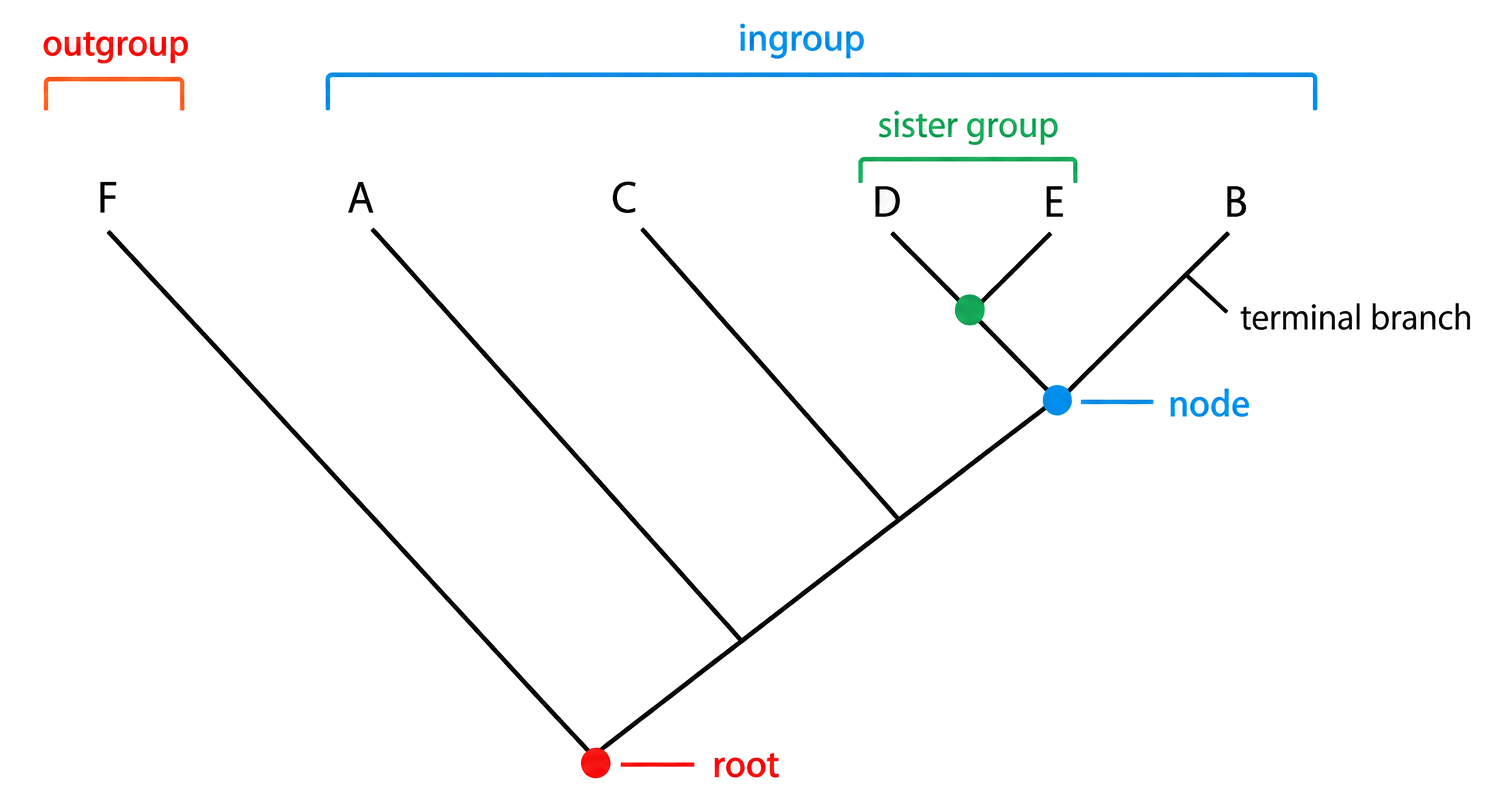
To compare two taxa, trace both branches backwards. The first node at which they meet represents their most recent common ancestor. Taxa that meet at a more recent node are more closely related, regardless of how near their tips appear on the page.
Root: common ancestor of every taxon shown
Node: hypothetical common ancestor and a divergence event
Terminal branch: lineage leading to the sampled taxon
Sister groups: lineages sharing an immediate common ancestor
Outgroup: a more distantly related comparison lineage outside the ingroup
Rotating branches around a node changes the drawing but not the relationships. The branching pattern carries the evidence; left-to-right tip order does not.
A cladogram is inferred from comparable characters
HL only
1
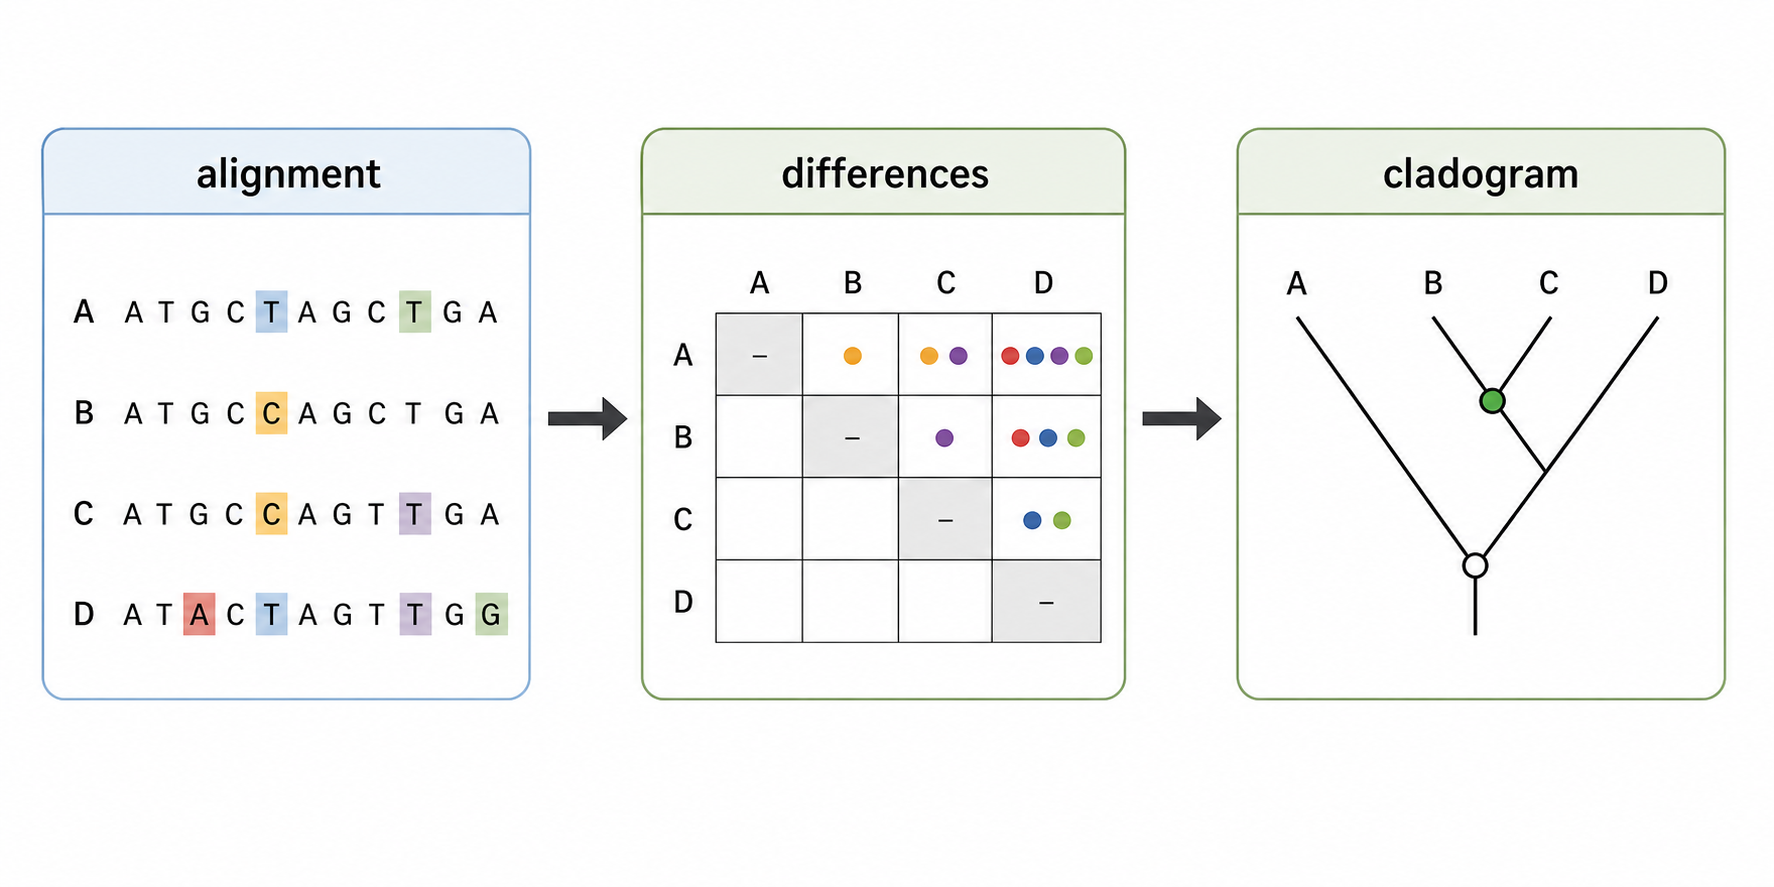
Choose the same homologous gene, RNA region or protein from every taxon and align the sequences so equivalent positions are compared.
2
Record shared characters and sequence differences. Similarity is informative only when the compared positions have the same evolutionary origin.
3
Computer analysis evaluates alternative branching patterns that could account for the observed distribution of characters.
4
Under parsimony, prefer the cladogram that explains the evidence with the fewest evolutionary changes. It is the best-supported hypothesis under that rule, not absolute proof.
Sequence difference becomes a clock only after calibration
HL only
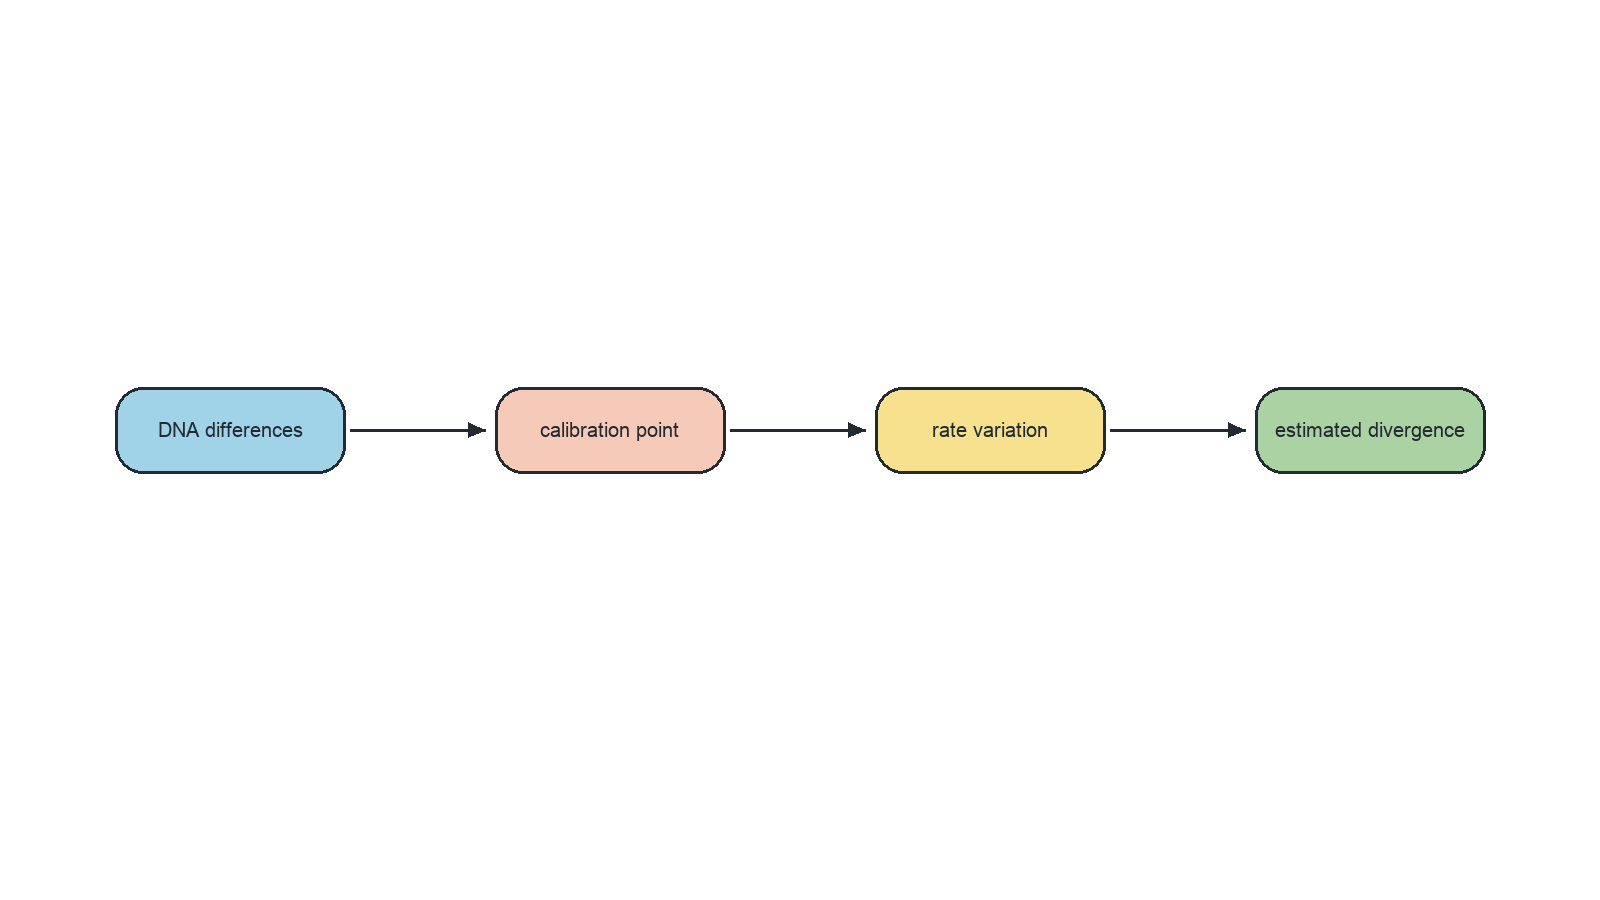
After two lineages diverge, mutations can accumulate independently in homologous sequences. On average, a greater sequence difference is consistent with a longer time since their common ancestor.
Using a molecular clock requires:
a homologous sequence compared across the taxa
an independently dated divergence, such as reliable fossil evidence, to calibrate the rate
the calibrated rate applied to the sequence difference for the unknown split
The result is an estimate. Mutation rates can differ among genes and lineages and can be affected by generation time, population history and selection. There is no single universal clock rate.
From a character to an evolutionary hypothesis
HL only
Move
Question to ask
What it supports
Choose a character
Is it homologous and comparable across taxa?
A defensible data set
Find shared derived states
Which taxa share a synapomorphy?
A candidate clade
Compare trees
Which branching pattern explains the characters with the fewest changes?
A parsimonious cladogram
Read the tree
Where is the most recent shared node?
A claim about relative relatedness
Calibrate sequence change
Is there an independent date and an appropriate rate?
An estimated divergence time
At every stage, distinguish the observation from the inference. A cladogram is a testable model of relationships, and a molecular-clock date is an estimate rather than direct observation of the past.
New evidence can split an old taxonomic group
HL only
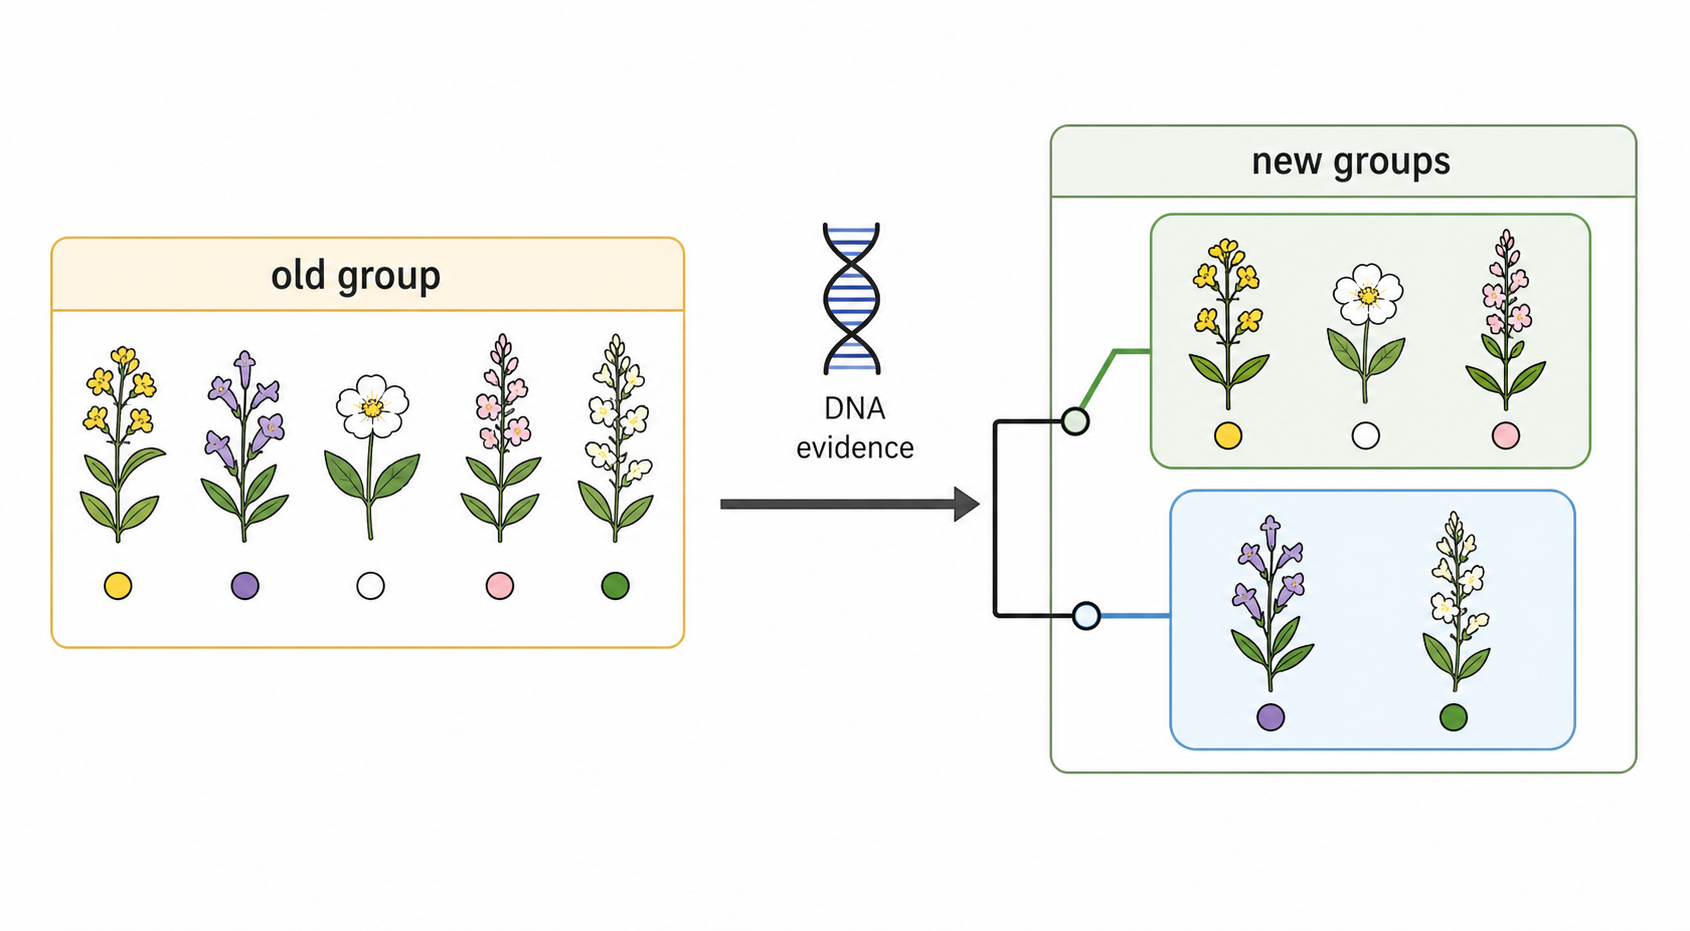
The traditional figwort family grouped many plants by similar flower morphology. That resemblance was a classification hypothesis, not proof of one recent common ancestor.
Comparisons of conserved chloroplast-gene sequences produced clades that did not preserve the old family boundary. Several genera were moved to other families, while some outsiders joined the redefined group.
The case shows how cladistics tests classification: molecular evidence can reveal that a shared appearance arose through convergence and falsify the original grouping. The specific genera are less important than the evidence → tree → reclassification reasoning.
rRNA evidence separated cellular life into three domains
HL only
Ribosomes occur in all cellular organisms, so homologous rRNA sequences can be compared across very distant groups. Their differences showed that prokaryotes contain two deeply distinct lineages, not one natural group.
The rRNA tree supports three deep lineages:
Bacteria (Eubacteria)
Archaea
Eukarya
“Prokaryote” still describes cells without a nucleus, but it does not name one monophyletic group. Similar cell organization can persist across lineages that diverged very early.
Independent cell features corroborate the three domains
HL only
Feature
Bacteria
Archaea
Eukarya
Chromosomes
Usually circular
Usually circular
Linear
Histone proteins
No true histones
Present
Present
Introns
Uncommon
Present in some genes
Common in many genes
Cell wall
Peptidoglycan when present
No peptidoglycan
No peptidoglycan; walls, if present, use cellulose or chitin
Membrane lipids
Ester-linked, unbranched fatty acids
Ether-linked, branched hydrocarbon chains
Ester-linked, unbranched fatty acids
These independent features agree with the rRNA evidence. Archaea share a prokaryotic cell organization with Bacteria but differ in fundamental molecular structures; some information-processing features resemble those of Eukarya.
Extreme habitats led scientists to study many archaea, but habitat does not define the domain. Archaea also occur in ordinary soils, oceans and animal digestive systems.
Match the claim to the evidence
HL only
Claim
Strongest relevant evidence
Boundary to remember
Two traits share an evolutionary origin
Homologous anatomy or homologous sequence data
Similar function alone may be convergence
Taxa form a clade
Shared derived characters and a tree containing an ancestor plus all descendants
A named traditional taxon may not be monophyletic
Two taxa are close relatives
A recent shared node
Tip position and branch rotation do not determine relatedness
One split is older than another
More accumulated difference in a calibrated homologous sequence
Rates vary, so dates are estimates
A classification should change
Independent evidence consistently contradicts its old boundary
A cladogram is a supported hypothesis, not direct observation
Cellular life has three domains
rRNA sequence patterns plus corroborating chromosome, histone, intron, wall and lipid evidence
“Prokaryote” describes cell organization but not one natural clade
Classification becomes scientifically useful when every group can be traced to evidence and revised when better evidence changes the inferred evolutionary relationships.
Need for classification
HL only
4 marks
Outline the principles used by scientists to classify organisms.
Analysing cladograms
HL only
2 marks
Outline what is meant by the "root" and "node" with reference to a cladogram.
Clades exam focus
HL only
3 marks
Outline the types of evidence that can be used to place a species in a particular clade.
Molecular clock
HL only
1 mark
State the relationship between divergence time and number of substitutions.
Constructing cladograms
HL only
3 marks
Discuss the use of amino acid sequences of proteins as the basis for constructing cladograms.
Difficulties with traditional hierarchy
HL only
1 mark
Based on the taxa shown, deduce a difficulty in gathering data to study turtle ancestry.
Advantages of evolutionary classification
HL only
8 marks
Explain how evolutionary relationships and cladograms can be used for classification.
Testing classification correspondence
HL only
1 mark
The figwort family is a large one consisting of many flowering plants that look similar. For what reason have some members of the family been reclassified into a new family?
Three domains classification
HL only
2 marks
Distinguish between the two domains of prokaryotes.